Jump to
S1C2
S1C3
Vibrational Frequencies calculated at B2PLYP/TZVP
Geometric Data calculated at B2PLYP/TZVP
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Vibrational Frequencies calculated at B2PLYP/TZVP
Geometric Data calculated at B2PLYP/TZVP
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B2PLYP/TZVP
| | hartrees |
|---|
| Energy at 0K | -210.261645 |
| Energy at 298.15K | -210.270801 |
| HF Energy | -210.034355 |
| Nuclear repulsion energy | 134.523305 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/TZVP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3741 |
3570 |
64.31 |
|
|
|
| 2 |
A |
3620 |
3455 |
3.77 |
|
|
|
| 3 |
A |
3529 |
3368 |
0.70 |
|
|
|
| 4 |
A |
3118 |
2976 |
41.88 |
|
|
|
| 5 |
A |
3108 |
2966 |
27.80 |
|
|
|
| 6 |
A |
3017 |
2880 |
75.94 |
|
|
|
| 7 |
A |
2992 |
2856 |
64.13 |
|
|
|
| 8 |
A |
1670 |
1594 |
34.48 |
|
|
|
| 9 |
A |
1530 |
1460 |
1.21 |
|
|
|
| 10 |
A |
1513 |
1444 |
4.83 |
|
|
|
| 11 |
A |
1460 |
1394 |
56.75 |
|
|
|
| 12 |
A |
1425 |
1360 |
11.02 |
|
|
|
| 13 |
A |
1388 |
1325 |
7.82 |
|
|
|
| 14 |
A |
1340 |
1279 |
2.66 |
|
|
|
| 15 |
A |
1265 |
1207 |
25.26 |
|
|
|
| 16 |
A |
1202 |
1147 |
11.43 |
|
|
|
| 17 |
A |
1113 |
1062 |
40.84 |
|
|
|
| 18 |
A |
1069 |
1021 |
42.58 |
|
|
|
| 19 |
A |
1015 |
968 |
8.43 |
|
|
|
| 20 |
A |
935 |
892 |
63.32 |
|
|
|
| 21 |
A |
887 |
847 |
18.00 |
|
|
|
| 22 |
A |
828 |
790 |
98.43 |
|
|
|
| 23 |
A |
574 |
548 |
128.12 |
|
|
|
| 24 |
A |
535 |
510 |
21.69 |
|
|
|
| 25 |
A |
329 |
314 |
0.58 |
|
|
|
| 26 |
A |
266 |
254 |
13.15 |
|
|
|
| 27 |
A |
182 |
174 |
6.35 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21824.3 cm
-1
Scaled (by 0.9544) Zero Point Vibrational Energy (zpe) 20829.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/TZVP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.366 |
-0.559 |
0.122 |
| C2 |
-0.633 |
0.650 |
-0.274 |
| C3 |
0.785 |
0.557 |
0.270 |
| O4 |
1.416 |
-0.633 |
-0.168 |
| H5 |
-1.668 |
-0.502 |
1.087 |
| H6 |
-2.189 |
-0.700 |
-0.448 |
| H7 |
-1.100 |
1.583 |
0.062 |
| H8 |
-0.588 |
0.668 |
-1.364 |
| H9 |
1.384 |
1.396 |
-0.084 |
| H10 |
0.758 |
0.605 |
1.367 |
| H11 |
0.729 |
-1.312 |
-0.101 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| N1 | | 1.4684 | 2.4278 | 2.7979 | 1.0128 | 1.0107 | 2.1600 | 2.0780 | 3.3806 | 2.7238 | 2.2371 |
C2 | 1.4684 | | 1.5215 | 2.4200 | 2.0621 | 2.0671 | 1.0966 | 1.0904 | 2.1592 | 2.1525 | 2.3948 | C3 | 2.4278 | 1.5215 | | 1.4166 | 2.7942 | 3.3073 | 2.1562 | 2.1367 | 1.0897 | 1.0989 | 1.9067 | O4 | 2.7979 | 2.4200 | 1.4166 | | 3.3320 | 3.6164 | 3.3610 | 2.6722 | 2.0309 | 2.0787 | 0.9683 | H5 | 1.0128 | 2.0621 | 2.7942 | 3.3320 | | 1.6326 | 2.3926 | 2.9226 | 3.7804 | 2.6819 | 2.7951 | H6 | 1.0107 | 2.0671 | 3.3073 | 3.6164 | 1.6326 | | 2.5805 | 2.2961 | 4.1583 | 3.6991 | 3.0014 | H7 | 2.1600 | 1.0966 | 2.1562 | 3.3610 | 2.3926 | 2.5805 | | 1.7697 | 2.4957 | 2.4730 | 3.4287 | H8 | 2.0780 | 1.0904 | 2.1367 | 2.6722 | 2.9226 | 2.2961 | 1.7697 | | 2.4612 | 3.0456 | 2.6926 | H9 | 3.3806 | 2.1592 | 1.0897 | 2.0309 | 3.7804 | 4.1583 | 2.4957 | 2.4612 | | 1.7675 | 2.7861 | H10 | 2.7238 | 2.1525 | 1.0989 | 2.0787 | 2.6819 | 3.6991 | 2.4730 | 3.0456 | 1.7675 | | 2.4149 | H11 | 2.2371 | 2.3948 | 1.9067 | 0.9683 | 2.7951 | 3.0014 | 3.4287 | 2.6926 | 2.7861 | 2.4149 | |
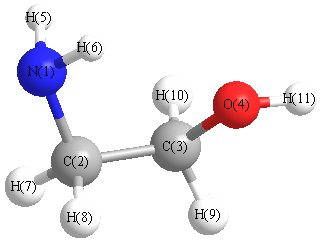 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
108.569 |
|
N1 |
C2 |
H7 |
113.943 |
| N1 |
C2 |
H8 |
107.680 |
|
C2 |
N1 |
H5 |
111.085 |
| C2 |
N1 |
H6 |
111.646 |
|
C2 |
C3 |
O4 |
110.856 |
| C2 |
C3 |
H9 |
110.465 |
|
C2 |
C3 |
H10 |
109.391 |
| C3 |
C2 |
H7 |
109.821 |
|
C3 |
C2 |
H8 |
108.655 |
| C3 |
O4 |
H11 |
104.592 |
|
O4 |
C3 |
H9 |
107.534 |
| O4 |
C3 |
H10 |
110.817 |
|
H5 |
N1 |
H6 |
107.571 |
| H7 |
C2 |
H8 |
108.039 |
|
H9 |
C3 |
H10 |
107.717 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/TZVP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.479 |
|
|
|
| 2 |
C |
-0.204 |
|
|
|
| 3 |
C |
-0.021 |
|
|
|
| 4 |
O |
-0.385 |
|
|
|
| 5 |
H |
0.203 |
|
|
|
| 6 |
H |
0.212 |
|
|
|
| 7 |
H |
0.099 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
| 9 |
H |
0.104 |
|
|
|
| 10 |
H |
0.080 |
|
|
|
| 11 |
H |
0.272 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-2.996 |
1.187 |
0.593 |
3.277 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-27.104 |
2.142 |
0.483 |
| y |
2.142 |
-25.917 |
-0.272 |
| z |
0.483 |
-0.272 |
-24.929 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.681 |
2.142 |
0.483 |
| y |
2.142 |
0.099 |
-0.272 |
| z |
0.483 |
-0.272 |
1.581 |
|
| Polar |
|---|
| 3z2-r2 | 3.162 |
|---|
| x2-y2 | -1.187 |
|---|
| xy | 2.142 |
|---|
| xz | 0.483 |
|---|
| yz | -0.272 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.850 |
0.168 |
0.012 |
| y |
0.168 |
5.654 |
0.056 |
| z |
0.012 |
0.056 |
5.167 |
<r2> (average value of r
2) Å
2
| <r2> |
87.458 |
| (<r2>)1/2 |
9.352 |
