Jump to
S1C2
S1C3
Energy calculated at MP3/TZVP
| | hartrees |
|---|
| Energy at 0K | -691.800208 |
| Energy at 298.15K | -691.799584 |
| HF Energy | -691.508636 |
| Nuclear repulsion energy | 59.275974 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP3/TZVP
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.364 |
| C2 |
0.000 |
0.000 |
-1.368 |
| N3 |
0.000 |
0.000 |
-2.530 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.7315 | 3.8941 |
C2 | 2.7315 | | 1.1626 | N3 | 3.8941 | 1.1626 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
180.000 |
|
K1 |
N3 |
C2 |
0.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP3/TZVP
| | hartrees |
|---|
| Energy at 0K | -691.802155 |
| Energy at 298.15K | -691.801477 |
| HF Energy | -691.515803 |
| Nuclear repulsion energy | 62.339170 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP3/TZVP
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
0.000 |
1.269 |
| C2 |
0.000 |
0.000 |
-2.483 |
| N3 |
0.000 |
0.000 |
-1.316 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 3.7526 | 2.5852 |
C2 | 3.7526 | | 1.1674 | N3 | 2.5852 | 1.1674 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
0.000 |
|
K1 |
N3 |
C2 |
180.000 |
| C2 |
K1 |
N3 |
0.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP3/TZVP
| | hartrees |
|---|
| Energy at 0K | -691.803489 |
| Energy at 298.15K | -691.803284 |
| HF Energy | -691.514138 |
| Nuclear repulsion energy | 65.941914 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at MP3/TZVP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| K1 |
0.000 |
1.109 |
0.000 |
| C2 |
0.624 |
-1.697 |
0.000 |
| N3 |
-0.534 |
-1.555 |
0.000 |
Atom - Atom Distances (Å)
| |
K1 |
C2 |
N3 |
| K1 | | 2.8746 | 2.7168 |
C2 | 2.8746 | | 1.1667 | N3 | 2.7168 | 1.1667 | |
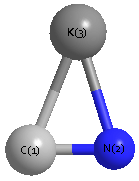 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| K1 |
C2 |
N3 |
70.461 |
|
K1 |
N3 |
C2 |
85.666 |
| C2 |
K1 |
N3 |
23.873 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
