Jump to
S1C2
Energy calculated at HSEh1PBE/TZVP
| | hartrees |
|---|
| Energy at 0K | -418.214814 |
| Energy at 298.15K | -418.218912 |
| HF Energy | -418.214814 |
| Nuclear repulsion energy | 61.165696 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/TZVP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3873 |
3718 |
57.54 |
|
|
|
| 2 |
A' |
2323 |
2230 |
114.14 |
|
|
|
| 3 |
A' |
1136 |
1091 |
15.43 |
|
|
|
| 4 |
A' |
1085 |
1041 |
43.31 |
|
|
|
| 5 |
A' |
905 |
869 |
29.80 |
|
|
|
| 6 |
A' |
800 |
768 |
160.67 |
|
|
|
| 7 |
A" |
2331 |
2238 |
150.55 |
|
|
|
| 8 |
A" |
915 |
878 |
17.77 |
|
|
|
| 9 |
A" |
406 |
390 |
118.12 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6886.2 cm
-1
Scaled (by 0.9601) Zero Point Vibrational Energy (zpe) 6611.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/TZVP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| P1 |
-0.108 |
-0.573 |
0.000 |
| O2 |
-0.108 |
1.099 |
0.000 |
| H3 |
0.768 |
1.493 |
0.000 |
| H4 |
0.855 |
-0.842 |
1.032 |
| H5 |
0.855 |
-0.842 |
-1.032 |
Atom - Atom Distances (Å)
| |
P1 |
O2 |
H3 |
H4 |
H5 |
| P1 | | 1.6725 | 2.2445 | 1.4363 | 1.4363 |
O2 | 1.6725 | | 0.9603 | 2.3996 | 2.3996 | H3 | 2.2445 | 0.9603 | | 2.5542 | 2.5542 | H4 | 1.4363 | 2.3996 | 2.5542 | | 2.0631 | H5 | 1.4363 | 2.3996 | 2.5542 | 2.0631 | |
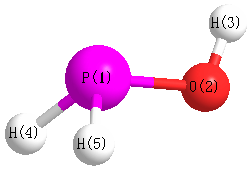 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| P1 |
O2 |
H3 |
114.231 |
|
O2 |
P1 |
H4 |
100.771 |
| O2 |
P1 |
H5 |
100.771 |
|
H4 |
P1 |
H5 |
91.808 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/TZVP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
P |
0.238 |
|
|
|
| 2 |
O |
-0.506 |
|
|
|
| 3 |
H |
0.296 |
|
|
|
| 4 |
H |
-0.014 |
|
|
|
| 5 |
H |
-0.014 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.309 |
-0.147 |
0.000 |
2.314 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.332 |
1.750 |
0.000 |
| y |
1.750 |
-19.911 |
0.000 |
| z |
0.000 |
0.000 |
-20.136 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.308 |
1.750 |
0.000 |
| y |
1.750 |
0.323 |
0.000 |
| z |
0.000 |
0.000 |
-0.014 |
|
| Polar |
|---|
| 3z2-r2 | -0.029 |
|---|
| x2-y2 | -0.421 |
|---|
| xy | 1.750 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.869 |
-0.312 |
0.000 |
| y |
-0.312 |
3.999 |
0.000 |
| z |
0.000 |
0.000 |
4.026 |
<r2> (average value of r
2) Å
2
| <r2> |
35.260 |
| (<r2>)1/2 |
5.938 |
Jump to
S1C1
Energy calculated at HSEh1PBE/TZVP
| | hartrees |
|---|
| Energy at 0K | -418.215428 |
| Energy at 298.15K | -418.219362 |
| HF Energy | -418.215428 |
| Nuclear repulsion energy | 61.073119 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HSEh1PBE/TZVP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3897 |
3742 |
102.78 |
|
|
|
| 2 |
A' |
2376 |
2281 |
87.68 |
|
|
|
| 3 |
A' |
1140 |
1095 |
34.23 |
|
|
|
| 4 |
A' |
1125 |
1080 |
81.10 |
|
|
|
| 5 |
A' |
898 |
862 |
29.58 |
|
|
|
| 6 |
A' |
793 |
762 |
114.37 |
|
|
|
| 7 |
A" |
2379 |
2284 |
112.79 |
|
|
|
| 8 |
A" |
927 |
890 |
1.99 |
|
|
|
| 9 |
A" |
257 |
247 |
111.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6895.4 cm
-1
Scaled (by 0.9601) Zero Point Vibrational Energy (zpe) 6620.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HSEh1PBE/TZVP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| P1 |
0.040 |
-0.579 |
0.000 |
| O2 |
0.040 |
1.104 |
0.000 |
| H3 |
0.948 |
1.413 |
0.000 |
| H4 |
-0.928 |
-0.781 |
1.032 |
| H5 |
-0.928 |
-0.781 |
-1.032 |
Atom - Atom Distances (Å)
| |
P1 |
O2 |
H3 |
H4 |
H5 |
| P1 | | 1.6829 | 2.1893 | 1.4293 | 1.4293 |
O2 | 1.6829 | | 0.9594 | 2.3567 | 2.3567 | H3 | 2.1893 | 0.9594 | | 3.0656 | 3.0656 | H4 | 1.4293 | 2.3567 | 3.0656 | | 2.0642 | H5 | 1.4293 | 2.3567 | 3.0656 | 2.0642 | |
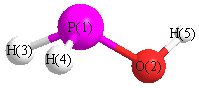 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| P1 |
O2 |
H3 |
108.796 |
|
O2 |
P1 |
H4 |
98.117 |
| O2 |
P1 |
H5 |
98.117 |
|
H4 |
P1 |
H5 |
92.459 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HSEh1PBE/TZVP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
P |
0.187 |
|
|
|
| 2 |
O |
-0.500 |
|
|
|
| 3 |
H |
0.293 |
|
|
|
| 4 |
H |
0.010 |
|
|
|
| 5 |
H |
0.010 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.770 |
-0.321 |
0.000 |
0.834 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.379 |
4.133 |
0.000 |
| y |
4.133 |
-20.510 |
0.000 |
| z |
0.000 |
0.000 |
-20.001 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.876 |
4.133 |
0.000 |
| y |
4.133 |
-0.819 |
0.000 |
| z |
0.000 |
0.000 |
-0.057 |
|
| Polar |
|---|
| 3z2-r2 | -0.114 |
|---|
| x2-y2 | 1.130 |
|---|
| xy | 4.133 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.967 |
0.569 |
0.000 |
| y |
0.569 |
4.022 |
0.000 |
| z |
0.000 |
0.000 |
4.020 |
<r2> (average value of r
2) Å
2
| <r2> |
35.250 |
| (<r2>)1/2 |
5.937 |
