Jump to
S1C2
Energy calculated at ROMP2/TZVP
| | hartrees |
|---|
| Energy at 0K | -151.565996 |
| Energy at 298.15K | |
| HF Energy | -151.123710 |
| Nuclear repulsion energy | 52.395439 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at ROMP2/TZVP
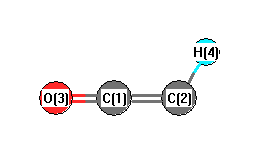
Geometric Data calculated at ROMP2/TZVP
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.017 |
| C2 |
0.000 |
0.000 |
-1.243 |
| O3 |
0.000 |
0.000 |
1.207 |
| H4 |
0.000 |
0.000 |
-2.303 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2600 | 1.1904 | 2.3198 |
C2 | 1.2600 | | 2.4504 | 1.0598 | O3 | 1.1904 | 2.4504 | | 3.5102 | H4 | 2.3198 | 1.0598 | 3.5102 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROMP2/TZVP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.385 |
|
|
|
| 2 |
C |
-0.264 |
|
|
|
| 3 |
O |
-0.364 |
|
|
|
| 4 |
H |
0.244 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
36.698 |
| (<r2>)1/2 |
6.058 |
Jump to
S1C1
Energy calculated at ROMP2/TZVP
| | hartrees |
|---|
| Energy at 0K | -151.569318 |
| Energy at 298.15K | |
| HF Energy | -151.123901 |
| Nuclear repulsion energy | 52.215926 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at ROMP2/TZVP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.057 |
0.000 |
| C2 |
1.139 |
-0.562 |
0.000 |
| O3 |
-1.125 |
0.411 |
0.000 |
| H4 |
2.166 |
-0.261 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2964 | 1.1794 | 2.1891 |
C2 | 1.2964 | | 2.4644 | 1.0701 | O3 | 1.1794 | 2.4644 | | 3.3588 | H4 | 2.1891 | 1.0701 | 3.3588 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
135.127 |
|
C2 |
C1 |
O3 |
168.944 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at ROMP2/TZVP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.466 |
|
|
|
| 2 |
C |
-0.299 |
|
|
|
| 3 |
O |
-0.335 |
|
|
|
| 4 |
H |
0.168 |
|
|
|
Electric dipole moments
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
36.811 |
| (<r2>)1/2 |
6.067 |
