return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CSNH2 (Ethanethioamide)

using model chemistry: MP3/TZVP

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP3/TZVP
 hartrees
Energy at 0K-531.308139
Energy at 298.15K 
HF Energy-530.677255
Nuclear repulsion energy155.229189
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/TZVP
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3787 3541 40.50      
2 A 3658 3421 75.13      
3 A 3228 3019 3.06      
4 A 3158 2954 13.29      
5 A 3092 2892 16.81      
6 A 1686 1577 172.58      
7 A 1513 1415 17.97      
8 A 1509 1411 11.91      
9 A 1446 1353 20.38      
10 A 1415 1324 357.46      
11 A 1368 1279 17.57      
12 A 1065 996 6.02      
13 A 1039 972 21.28      
14 A 1026 959 34.36      
15 A 760 711 6.25      
16 A 605 566 17.21      
17 A 490 458 12.12      
18 A 440 411 2.54      
19 A 387 362 4.09      
20 A 321 300 223.55      
21 A 77 72 3.63      

Unscaled Zero Point Vibrational Energy (zpe) 16034.3 cm-1
Scaled (by 0.9352) Zero Point Vibrational Energy (zpe) 14995.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/TZVP
ABC
0.32598 0.16555 0.11214

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/TZVP

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.275 0.060 -0.011
S2 -1.364 -0.114 0.003
C3 1.232 -1.106 0.001
N4 0.882 1.264 -0.032
H5 0.734 -2.001 -0.354
H6 2.103 -0.902 -0.625
H7 1.576 -1.282 1.022
H8 1.873 1.342 0.108
H9 0.319 2.085 0.091

Atom - Atom Distances (Å)
  C1 S2 C3 N4 H5 H6 H7 H8 H9
C11.64901.50851.34832.13942.15492.13522.05182.0282
S21.64902.77982.63562.84483.61073.32413.55112.7706
C31.50852.77982.39581.08401.09111.09172.53283.3207
N41.34832.63562.39583.28422.55622.84161.00351.0037
H52.13942.84481.08403.28421.77601.76563.56204.1315
H62.15493.61071.09112.55621.77601.77002.37263.5528
H72.13523.32411.09172.84161.76561.77002.79453.7131
H82.05183.55112.53281.00353.56202.37262.79451.7227
H92.02822.77063.32071.00374.13153.55283.71311.7227

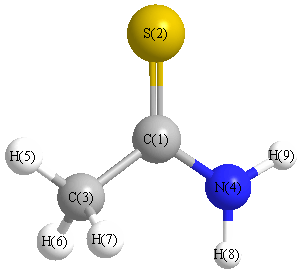
picture of Ethanethioamide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 H5 110.145 C1 C3 H6 110.960
C1 C3 H7 109.355 C1 N4 H8 120.775
C1 N4 H9 118.417 S2 C1 C3 123.316
S2 C1 N4 122.810 C3 C1 N4 113.874
H5 C3 H6 109.469 H5 C3 H7 108.488
H6 C3 H7 108.368 H8 N4 H9 118.245
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability