Jump to
S1C2
Vibrational Frequencies calculated at MP3/TZVP
Geometric Data calculated at MP3/TZVP
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP3/TZVP
| | hartrees |
|---|
| Energy at 0K | -553.232437 |
| Energy at 298.15K | -553.237902 |
| HF Energy | -552.492435 |
| Nuclear repulsion energy | 201.941019 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/TZVP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3308 |
3094 |
7.82 |
|
|
|
| 2 |
A |
3304 |
3090 |
5.95 |
|
|
|
| 3 |
A |
3248 |
3037 |
5.42 |
|
|
|
| 4 |
A |
3235 |
3025 |
5.68 |
|
|
|
| 5 |
A |
3216 |
3008 |
1.10 |
|
|
|
| 6 |
A |
3210 |
3002 |
2.02 |
|
|
|
| 7 |
A |
1697 |
1587 |
29.77 |
|
|
|
| 8 |
A |
1686 |
1577 |
26.72 |
|
|
|
| 9 |
A |
1460 |
1365 |
4.64 |
|
|
|
| 10 |
A |
1457 |
1362 |
6.28 |
|
|
|
| 11 |
A |
1335 |
1249 |
2.34 |
|
|
|
| 12 |
A |
1328 |
1242 |
10.50 |
|
|
|
| 13 |
A |
1081 |
1011 |
7.83 |
|
|
|
| 14 |
A |
1056 |
987 |
11.32 |
|
|
|
| 15 |
A |
1007 |
941 |
33.52 |
|
|
|
| 16 |
A |
996 |
932 |
30.72 |
|
|
|
| 17 |
A |
864 |
808 |
32.42 |
|
|
|
| 18 |
A |
847 |
792 |
37.93 |
|
|
|
| 19 |
A |
755 |
706 |
9.67 |
|
|
|
| 20 |
A |
707 |
661 |
4.52 |
|
|
|
| 21 |
A |
638 |
596 |
16.05 |
|
|
|
| 22 |
A |
598 |
559 |
13.39 |
|
|
|
| 23 |
A |
464 |
434 |
0.44 |
|
|
|
| 24 |
A |
376 |
352 |
0.56 |
|
|
|
| 25 |
A |
229 |
214 |
0.94 |
|
|
|
| 26 |
A |
134 |
125 |
2.85 |
|
|
|
| 27 |
A |
88 |
83 |
1.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 19161.7 cm
-1
Scaled (by 0.9352) Zero Point Vibrational Energy (zpe) 17920.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP3/TZVP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.013 |
-0.994 |
-0.000 |
| C2 |
1.080 |
0.372 |
0.383 |
| C3 |
-1.566 |
-0.186 |
-0.025 |
| C4 |
2.247 |
0.560 |
-0.225 |
| C5 |
-1.802 |
1.119 |
-0.128 |
| H6 |
0.765 |
0.985 |
1.217 |
| H7 |
-2.379 |
-0.893 |
0.066 |
| H8 |
2.920 |
1.337 |
0.108 |
| H9 |
2.559 |
-0.050 |
-1.060 |
| H10 |
-1.012 |
1.846 |
-0.245 |
| H11 |
-2.818 |
1.485 |
-0.110 |
Atom - Atom Distances (Å)
| |
S1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| S1 | | 1.7747 | 1.7733 | 2.7305 | 2.7885 | 2.4422 | 2.3955 | 3.7279 | 2.9145 | 3.0293 | 3.7646 |
C2 | 1.7747 | | 2.7342 | 1.3293 | 3.0208 | 1.0823 | 3.6968 | 2.0963 | 2.1089 | 2.6354 | 4.0838 | C3 | 1.7733 | 2.7342 | | 3.8903 | 1.3309 | 2.8892 | 1.0815 | 4.7395 | 4.2547 | 2.1181 | 2.0904 | C4 | 2.7305 | 1.3293 | 3.8903 | | 4.0890 | 2.1106 | 4.8581 | 1.0808 | 1.0806 | 3.5042 | 5.1506 | C5 | 2.7885 | 3.0208 | 1.3309 | 4.0890 | | 2.9011 | 2.1022 | 4.7335 | 4.6106 | 1.0800 | 1.0801 | H6 | 2.4422 | 1.0823 | 2.8892 | 2.1106 | 2.9011 | | 3.8394 | 2.4488 | 3.0780 | 2.4571 | 3.8538 | H7 | 2.3955 | 3.6968 | 1.0815 | 4.8581 | 2.1022 | 3.8394 | | 5.7500 | 5.1347 | 3.0771 | 2.4242 | H8 | 3.7279 | 2.0963 | 4.7395 | 1.0808 | 4.7335 | 2.4488 | 5.7500 | | 1.8497 | 3.9813 | 5.7448 | H9 | 2.9145 | 2.1089 | 4.2547 | 1.0806 | 4.6106 | 3.0780 | 5.1347 | 1.8497 | | 4.1250 | 5.6723 | H10 | 3.0293 | 2.6354 | 2.1181 | 3.5042 | 1.0800 | 2.4571 | 3.0771 | 3.9813 | 4.1250 | | 1.8468 | H11 | 3.7646 | 4.0838 | 2.0904 | 5.1506 | 1.0801 | 3.8538 | 2.4242 | 5.7448 | 5.6723 | 1.8468 | |
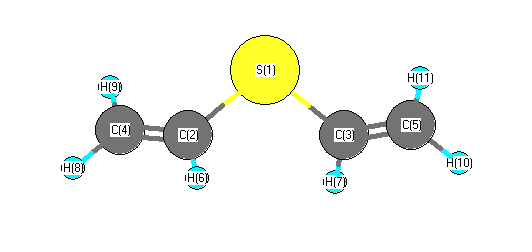 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
C2 |
C4 |
122.565 |
|
S1 |
C2 |
H6 |
115.334 |
| S1 |
C3 |
C5 |
127.298 |
|
S1 |
C3 |
H7 |
111.800 |
| C2 |
S1 |
C3 |
100.826 |
|
C2 |
C4 |
H8 |
120.531 |
| C2 |
C4 |
H9 |
121.774 |
|
C3 |
C5 |
H10 |
122.602 |
| C3 |
C5 |
H11 |
119.873 |
|
C4 |
C2 |
H6 |
121.810 |
| C5 |
C3 |
H7 |
120.900 |
|
H8 |
C4 |
H9 |
117.694 |
| H10 |
C5 |
H11 |
117.517 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
