return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C(CH3)3SH (2-Propanethiol, 2-methyl-)

using model chemistry: MP3/TZVP

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP3/TZVP
 hartrees
Energy at 0K-555.699254
Energy at 298.15K-555.710137
HF Energy-554.883393
Nuclear repulsion energy244.954687
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP3/TZVP
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3175 2969 27.71      
2 A' 3160 2956 32.77      
3 A' 3151 2947 40.88      
4 A' 3085 2885 23.43      
5 A' 3077 2878 31.69      
6 A' 2745 2567 9.98      
7 A' 1542 1442 6.73      
8 A' 1532 1433 8.33      
9 A' 1513 1415 0.16      
10 A' 1475 1379 0.69      
11 A' 1444 1350 7.65      
12 A' 1300 1215 0.19      
13 A' 1244 1163 32.24      
14 A' 1092 1021 1.92      
15 A' 977 913 0.25      
16 A' 904 846 4.96      
17 A' 861 805 1.55      
18 A' 616 576 3.78      
19 A' 403 377 1.50      
20 A' 376 352 0.04      
21 A' 310 290 0.18      
22 A' 295 276 0.77      
23 A" 3174 2968 29.22      
24 A" 3169 2964 4.40      
25 A" 3146 2942 1.12      
26 A" 3073 2874 17.28      
27 A" 1532 1433 6.98      
28 A" 1508 1411 0.05      
29 A" 1503 1406 0.05      
30 A" 1444 1350 7.97      
31 A" 1285 1202 2.36      
32 A" 1082 1012 0.11      
33 A" 998 933 0.01      
34 A" 967 904 0.01      
35 A" 408 382 0.75      
36 A" 312 292 1.22      
37 A" 290 271 0.04      
38 A" 243 228 5.96      
39 A" 176 164 10.14      

Unscaled Zero Point Vibrational Energy (zpe) 29291.8 cm-1
Scaled (by 0.9352) Zero Point Vibrational Energy (zpe) 27393.7 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP3/TZVP
ABC
0.15051 0.09965 0.09868

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP3/TZVP

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.345 -0.005 0.000
S2 -1.504 0.085 0.000
C3 0.835 1.444 0.000
C4 0.835 -0.726 1.254
C5 0.835 -0.726 -1.254
H6 -1.714 -1.240 0.000
H7 1.927 1.457 0.000
H8 0.486 1.977 -0.885
H9 0.486 1.977 0.885
H10 1.928 -0.749 1.264
H11 1.928 -0.749 -1.264
H12 0.487 -0.221 2.154
H13 0.475 -1.756 1.280
H14 0.487 -0.221 -2.154
H15 0.475 -1.756 -1.280

Atom - Atom Distances (Å)
  C1 S2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
C11.85171.52881.52711.52712.40202.15342.17482.17482.15782.15782.16982.17292.16982.1729
S21.85172.70472.77502.77501.34223.69472.88492.88493.75123.75122.94962.99062.94962.9906
C31.52882.70472.50622.50623.70171.09181.09041.09042.75702.75702.74493.46482.74493.4648
C41.52712.77502.50622.50732.88692.74443.46452.75061.09322.74501.08951.09103.46262.7583
C51.52712.77502.50622.50732.88692.74442.75063.46452.74501.09323.46262.75831.08951.0910
H62.40201.34223.70172.88692.88694.53143.99703.99703.88653.88653.24442.58783.24442.5878
H72.15343.69471.09182.74442.74444.53141.76861.76862.54292.54293.08713.75103.08713.7510
H82.17482.88491.09043.46452.75063.99701.76861.76983.75883.10703.75094.31512.53863.7537
H92.17482.88491.09042.75063.46453.99701.76861.76983.10703.75882.53863.75373.75094.3151
H102.15783.75122.75701.09322.74503.88652.54293.75883.10702.52871.77351.76783.74703.0980
H112.15783.75122.75702.74501.09323.88652.54293.10703.75882.52873.74703.09801.77351.7678
H122.16982.94962.74491.08953.46263.24443.08713.75092.53861.77353.74701.76614.30863.7614
H132.17292.99063.46481.09102.75832.58783.75104.31513.75371.76783.09801.76613.76142.5597
H142.16982.94962.74493.46261.08953.24443.08712.53863.75093.74701.77354.30863.76141.7661
H152.17292.99063.46482.75831.09102.58783.75103.75374.31513.09801.76783.76142.55971.7661

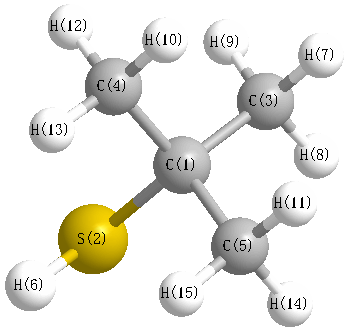
picture of 2-Propanethiol, 2-methyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 S2 H6 96.233 C1 C3 H7 109.380
C1 C3 H8 111.160 C1 C3 H9 111.160
C1 C4 H10 109.768 C1 C4 H12 110.940
C1 C4 H13 111.095 C1 C5 H11 109.768
C1 C5 H14 110.940 C1 C5 H15 111.095
S2 C1 C3 105.883 S2 C1 C4 110.063
S2 C1 C5 110.063 C3 C1 C4 110.190
C3 C1 C5 110.190 C4 C1 C5 110.362
H7 C3 H8 108.281 H7 C3 H9 108.281
H8 C3 H9 108.489 H10 C4 H12 108.694
H10 C4 H13 108.067 H11 C5 H14 108.694
H11 C5 H15 108.067 H12 C4 H13 108.192
H14 C5 H15 108.192
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability