Jump to
S1C2
Energy calculated at CCSD(T)/aug-cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -721.550027 |
| Energy at 298.15K | |
| HF Energy | -721.319737 |
| Nuclear repulsion energy | 89.400153 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCSD(T)/aug-cc-pCVTZ
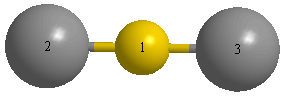
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.000 |
| Na2 |
0.000 |
0.000 |
2.442 |
| Na3 |
0.000 |
0.000 |
-2.442 |
Atom - Atom Distances (Å)
| |
S1 |
Na2 |
Na3 |
| S1 | | 2.4415 | 2.4415 |
Na2 | 2.4415 | | 4.8831 | Na3 | 2.4415 | 4.8831 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
S1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/aug-cc-pCVTZ
| | hartrees |
|---|
| Energy at 0K | -721.550739 |
| Energy at 298.15K | |
| HF Energy | -721.319030 |
| Nuclear repulsion energy | 90.162471 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at CCSD(T)/aug-cc-pCVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.611 |
| Na2 |
0.000 |
2.221 |
-0.445 |
| Na3 |
0.000 |
-2.221 |
-0.445 |
Atom - Atom Distances (Å)
| |
S1 |
Na2 |
Na3 |
| S1 | | 2.4591 | 2.4591 |
Na2 | 2.4591 | | 4.4415 | Na3 | 2.4591 | 4.4415 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
S1 |
Na3 |
129.131 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
