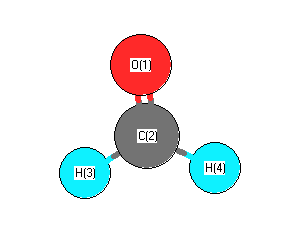
Vibrational Frequencies calculated at LSDA/aug-cc-pCVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
2784 |
2784 |
73.25 |
218.59 |
0.10 |
0.19 |
| 2 |
A1 |
1810 |
1810 |
108.81 |
7.93 |
0.40 |
0.57 |
| 3 |
A1 |
1461 |
1461 |
9.02 |
8.17 |
0.43 |
0.61 |
| 4 |
B1 |
1142 |
1142 |
7.12 |
0.61 |
0.75 |
0.86 |
| 5 |
B2 |
2837 |
2837 |
119.49 |
135.43 |
0.75 |
0.86 |
| 6 |
B2 |
1213 |
1213 |
8.35 |
1.46 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 5623.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 5623.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at LSDA/aug-cc-pCVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.446 |
|
|
|
| 2 |
C |
-0.315 |
|
|
|
| 3 |
H |
0.381 |
|
|
|
| 4 |
H |
0.381 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-2.252 |
2.252 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-11.838 |
0.000 |
0.000 |
| y |
0.000 |
-11.737 |
0.000 |
| z |
0.000 |
0.000 |
-12.467 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.264 |
0.000 |
0.000 |
| y |
0.000 |
0.416 |
0.000 |
| z |
0.000 |
0.000 |
-0.680 |
|
| Polar |
|---|
| 3z2-r2 | -1.359 |
|---|
| x2-y2 | -0.101 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.988 |
0.000 |
0.000 |
| y |
0.000 |
2.913 |
0.000 |
| z |
0.000 |
0.000 |
3.479 |
<r2> (average value of r
2) Å
2
| <r2> |
17.083 |
| (<r2>)1/2 |
4.133 |