All results from a given calculation for CH3CH2CHO (Propanal)
using model chemistry: CCSD(T)=FULL/daug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at CCSD(T)=FULL/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -192.919157 |
| Energy at 298.15K | |
| HF Energy | -192.025845 |
| Nuclear repulsion energy | 118.816171 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)=FULL/daug-cc-pVTZ
Geometric Data calculated at CCSD(T)=FULL/daug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.445 |
0.450 |
0.000 |
| C2 |
0.000 |
0.914 |
0.000 |
| C3 |
-0.995 |
-0.211 |
0.000 |
| O4 |
-0.703 |
-1.385 |
0.000 |
| H5 |
2.121 |
1.304 |
0.000 |
| H6 |
1.653 |
-0.159 |
0.878 |
| H7 |
1.653 |
-0.159 |
-0.878 |
| H8 |
-0.224 |
1.538 |
0.871 |
| H9 |
-0.224 |
1.538 |
-0.871 |
| H10 |
-2.056 |
0.099 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.5178 | 2.5279 | 2.8244 | 1.0892 | 1.0889 | 1.0889 | 2.1751 | 2.1751 | 3.5188 |
C2 | 1.5178 | | 1.5023 | 2.4038 | 2.1567 | 2.1579 | 2.1579 | 1.0950 | 1.0950 | 2.2121 | C3 | 2.5279 | 1.5023 | | 1.2091 | 3.4649 | 2.7903 | 2.7903 | 2.1010 | 2.1010 | 1.1056 | O4 | 2.8244 | 2.4038 | 1.2091 | | 3.8988 | 2.7967 | 2.7967 | 3.0872 | 3.0872 | 2.0080 | H5 | 1.0892 | 2.1567 | 3.4649 | 3.8988 | | 1.7694 | 1.7694 | 2.5133 | 2.5133 | 4.3478 | H6 | 1.0889 | 2.1579 | 2.7903 | 2.7967 | 1.7694 | | 1.7567 | 2.5311 | 3.0770 | 3.8205 | H7 | 1.0889 | 2.1579 | 2.7903 | 2.7967 | 1.7694 | 1.7567 | | 3.0770 | 2.5311 | 3.8205 | H8 | 2.1751 | 1.0950 | 2.1010 | 3.0872 | 2.5133 | 2.5311 | 3.0770 | | 1.7428 | 2.4874 | H9 | 2.1751 | 1.0950 | 2.1010 | 3.0872 | 2.5133 | 3.0770 | 2.5311 | 1.7428 | | 2.4874 | H10 | 3.5188 | 2.2121 | 1.1056 | 2.0080 | 4.3478 | 3.8205 | 3.8205 | 2.4874 | 2.4874 | |
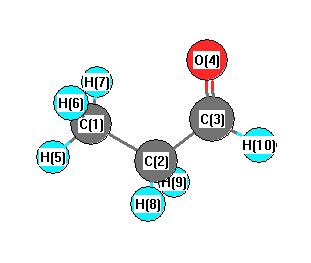 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.655 |
|
C1 |
C2 |
H8 |
111.686 |
| C1 |
C2 |
H9 |
111.686 |
|
C2 |
C1 |
H5 |
110.559 |
| C2 |
C1 |
H6 |
110.672 |
|
C2 |
C1 |
H7 |
110.672 |
| C2 |
C3 |
O4 |
124.536 |
|
C2 |
C3 |
H10 |
115.193 |
| C3 |
C2 |
H8 |
106.932 |
|
C3 |
C2 |
H9 |
106.932 |
| O4 |
C3 |
H10 |
120.271 |
|
H5 |
C1 |
H6 |
108.656 |
| H5 |
C1 |
H7 |
108.656 |
|
H6 |
C1 |
H7 |
107.536 |
| H8 |
C2 |
H9 |
105.455 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
