Jump to
S1C2
Energy calculated at mPW1PW91/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -722.860680 |
| Energy at 298.15K | |
| HF Energy | -722.860680 |
| Nuclear repulsion energy | 90.645651 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
281 |
281 |
0.00 |
170.81 |
0.07 |
0.13 |
| 2 |
Σu |
430 |
430 |
111.05 |
0.00 |
0.00 |
0.00 |
| 3 |
Πu |
39i |
39i |
70.26 |
0.00 |
0.00 |
0.00 |
| 3 |
Πu |
39i |
39i |
70.26 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 316.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 316.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/daug-cc-pVTZ
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.000 |
| Na2 |
0.000 |
0.000 |
2.408 |
| Na3 |
0.000 |
0.000 |
-2.408 |
Atom - Atom Distances (Å)
| |
S1 |
Na2 |
Na3 |
| S1 | | 2.4081 | 2.4081 |
Na2 | 2.4081 | | 4.8162 | Na3 | 2.4081 | 4.8162 | |
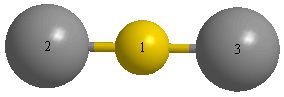 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
S1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.761 |
|
|
|
| 2 |
Na |
0.380 |
|
|
|
| 3 |
Na |
0.380 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.596 |
0.000 |
0.000 |
| y |
0.000 |
-30.596 |
0.000 |
| z |
0.000 |
0.000 |
18.679 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-24.637 |
0.000 |
0.000 |
| y |
0.000 |
-24.637 |
0.000 |
| z |
0.000 |
0.000 |
49.275 |
|
| Polar |
|---|
| 3z2-r2 | 98.549 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.153 |
0.000 |
0.000 |
| y |
0.000 |
13.153 |
0.000 |
| z |
0.000 |
0.000 |
20.346 |
<r2> (average value of r
2) Å
2
| <r2> |
136.430 |
| (<r2>)1/2 |
11.680 |
Jump to
S1C1
Energy calculated at mPW1PW91/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -722.861786 |
| Energy at 298.15K | |
| HF Energy | -722.861786 |
| Nuclear repulsion energy | 91.508486 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
305 |
305 |
16.77 |
255.40 |
0.06 |
0.12 |
| 2 |
A1 |
48 |
48 |
41.26 |
5.48 |
0.64 |
0.78 |
| 3 |
B2 |
399 |
399 |
88.01 |
10.92 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 375.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 375.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/daug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.634 |
| Na2 |
0.000 |
2.167 |
-0.461 |
| Na3 |
0.000 |
-2.167 |
-0.461 |
Atom - Atom Distances (Å)
| |
S1 |
Na2 |
Na3 |
| S1 | | 2.4275 | 2.4275 |
Na2 | 2.4275 | | 4.3334 | Na3 | 2.4275 | 4.3334 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
S1 |
Na3 |
126.390 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.636 |
|
|
|
| 2 |
Na |
0.318 |
|
|
|
| 3 |
Na |
0.318 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-6.071 |
6.071 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.745 |
0.000 |
0.000 |
| y |
0.000 |
8.701 |
0.000 |
| z |
0.000 |
0.000 |
-29.631 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-20.280 |
0.000 |
0.000 |
| y |
0.000 |
38.888 |
0.000 |
| z |
0.000 |
0.000 |
-18.608 |
|
| Polar |
|---|
| 3z2-r2 | -37.217 |
|---|
| x2-y2 | -39.445 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.308 |
0.000 |
0.000 |
| y |
0.000 |
20.518 |
0.000 |
| z |
0.000 |
0.000 |
14.985 |
<r2> (average value of r
2) Å
2
| <r2> |
125.139 |
| (<r2>)1/2 |
11.187 |
