Jump to
S1C2
Energy calculated at B2PLYP/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -722.498236 |
| Energy at 298.15K | |
| HF Energy | -722.429171 |
| Nuclear repulsion energy | 90.112470 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
277 |
277 |
0.00 |
255.43 |
0.08 |
0.14 |
| 2 |
Σu |
425 |
425 |
107.85 |
0.00 |
0.60 |
0.75 |
| 3 |
Πu |
45i |
45i |
66.83 |
0.00 |
0.00 |
0.00 |
| 3 |
Πu |
45i |
45i |
66.83 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 306.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 306.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/daug-cc-pVTZ
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.000 |
| Na2 |
0.000 |
0.000 |
2.422 |
| Na3 |
0.000 |
0.000 |
-2.422 |
Atom - Atom Distances (Å)
| |
S1 |
Na2 |
Na3 |
| S1 | | 2.4224 | 2.4224 |
Na2 | 2.4224 | | 4.8447 | Na3 | 2.4224 | 4.8447 | |
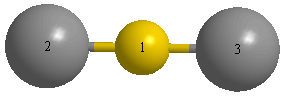 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
S1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.651 |
|
|
|
| 2 |
Na |
0.325 |
|
|
|
| 3 |
Na |
0.325 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.194 |
0.000 |
0.000 |
| y |
0.000 |
-31.194 |
0.000 |
| z |
0.000 |
0.000 |
18.382 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-24.788 |
0.000 |
0.000 |
| y |
0.000 |
-24.788 |
0.000 |
| z |
0.000 |
0.000 |
49.577 |
|
| Polar |
|---|
| 3z2-r2 | 99.153 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.069 |
0.000 |
0.000 |
| y |
0.000 |
14.069 |
0.000 |
| z |
0.000 |
0.000 |
23.026 |
<r2> (average value of r
2) Å
2
| <r2> |
138.254 |
| (<r2>)1/2 |
11.758 |
Jump to
S1C1
Energy calculated at B2PLYP/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -722.499748 |
| Energy at 298.15K | |
| HF Energy | -722.429616 |
| Nuclear repulsion energy | 91.040198 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
301 |
301 |
14.62 |
405.04 |
0.07 |
0.13 |
| 2 |
A1 |
52 |
52 |
35.36 |
7.91 |
0.67 |
0.80 |
| 3 |
B2 |
388 |
388 |
81.20 |
19.40 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 370.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 370.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/daug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.676 |
| Na2 |
0.000 |
2.150 |
-0.492 |
| Na3 |
0.000 |
-2.150 |
-0.492 |
Atom - Atom Distances (Å)
| |
S1 |
Na2 |
Na3 |
| S1 | | 2.4462 | 2.4462 |
Na2 | 2.4462 | | 4.2992 | Na3 | 2.4462 | 4.2992 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
S1 |
Na3 |
122.981 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.501 |
|
|
|
| 2 |
Na |
0.251 |
|
|
|
| 3 |
Na |
0.251 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-6.266 |
6.266 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.406 |
0.000 |
0.000 |
| y |
0.000 |
6.874 |
0.000 |
| z |
0.000 |
0.000 |
-30.113 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-19.786 |
0.000 |
0.000 |
| y |
0.000 |
37.633 |
0.000 |
| z |
0.000 |
0.000 |
-17.848 |
|
| Polar |
|---|
| 3z2-r2 | -35.695 |
|---|
| x2-y2 | -38.279 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
14.313 |
0.000 |
0.000 |
| y |
0.000 |
23.769 |
0.000 |
| z |
0.000 |
0.000 |
16.751 |
<r2> (average value of r
2) Å
2
| <r2> |
125.643 |
| (<r2>)1/2 |
11.209 |
