Jump to
S1C2
S1C3
Energy calculated at MP2/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1590.846894 |
| Energy at 298.15K | |
| HF Energy | -1590.151044 |
| Nuclear repulsion energy | 351.392716 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
533 |
533 |
0.00 |
38.21 |
0.04 |
0.07 |
| 2 |
A1 |
213 |
213 |
0.00 |
1.18 |
0.71 |
0.83 |
| 3 |
B1 |
474 |
474 |
0.00 |
13.22 |
0.75 |
0.86 |
| 4 |
B2 |
303 |
303 |
0.04 |
5.82 |
0.75 |
0.86 |
| 5 |
E |
446 |
446 |
0.01 |
3.02 |
0.75 |
0.86 |
| 5 |
E |
446 |
446 |
0.01 |
3.02 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 1207.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1207.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/daug-cc-pVTZ
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.423 |
0.327 |
| S2 |
0.000 |
-1.423 |
0.327 |
| S3 |
-1.423 |
0.000 |
-0.327 |
| S4 |
1.423 |
0.000 |
-0.327 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8452 | 2.1153 | 2.1153 |
S2 | 2.8452 | | 2.1153 | 2.1153 | S3 | 2.1153 | 2.1153 | | 2.8452 | S4 | 2.1153 | 2.1153 | 2.8452 | |
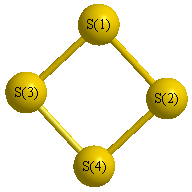 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.737 |
|
S1 |
S3 |
S4 |
47.737 |
| S2 |
S1 |
S3 |
47.737 |
|
S2 |
S4 |
S3 |
47.737 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1590.916037 |
| Energy at 298.15K | |
| HF Energy | -1590.081790 |
| Nuclear repulsion energy | 327.418483 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
649 |
649 |
0.00 |
0.00 |
0.33 |
0.50 |
| 2 |
A1 |
458 |
458 |
0.83 |
23743.41 |
0.33 |
0.50 |
| 3 |
A1 |
228 |
228 |
0.00 |
0.00 |
0.33 |
0.50 |
| 4 |
A2 |
225 |
225 |
0.00 |
0.03 |
0.75 |
0.86 |
| 5 |
B2 |
3953 |
3953 |
7481508.00 |
68.64 |
0.75 |
0.86 |
| 6 |
B2 |
307 |
307 |
0.00 |
4391.32 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 2910.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 2910.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/daug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.304 |
0.967 |
| S2 |
0.000 |
-1.304 |
0.967 |
| S3 |
0.000 |
1.304 |
-0.967 |
| S4 |
0.000 |
-1.304 |
-0.967 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.6079 | 1.9342 | 3.2469 |
S2 | 2.6079 | | 3.2469 | 1.9342 | S3 | 1.9342 | 3.2469 | | 2.6078 | S4 | 3.2469 | 1.9342 | 2.6078 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
89.998 |
|
S1 |
S3 |
S4 |
90.002 |
| S2 |
S1 |
S3 |
89.998 |
|
S2 |
S4 |
S3 |
90.002 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1590.916037 |
| Energy at 298.15K | |
| HF Energy | -1590.081833 |
| Nuclear repulsion energy | 327.435879 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
650 |
650 |
0.00 |
0.00 |
0.33 |
0.50 |
| 2 |
Ag |
228 |
228 |
0.00 |
0.00 |
0.33 |
0.50 |
| 3 |
Au |
225 |
225 |
0.00 |
0.00 |
0.75 |
0.86 |
| 4 |
B1u |
3930 |
3930 |
7300007.00 |
0.00 |
0.74 |
0.85 |
| 5 |
B2u |
458 |
458 |
0.82 |
0.01 |
0.61 |
0.76 |
| 6 |
B3g |
307 |
307 |
0.00 |
4248.39 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 2898.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 2898.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2/daug-cc-pVTZ
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.967 |
1.304 |
| S2 |
0.000 |
0.967 |
-1.304 |
| S3 |
0.000 |
-0.967 |
1.304 |
| S4 |
0.000 |
-0.967 |
-1.304 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.6080 | 1.9339 | 3.2468 |
S2 | 2.6080 | | 3.2468 | 1.9339 | S3 | 1.9339 | 3.2468 | | 2.6080 | S4 | 3.2468 | 1.9339 | 2.6080 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
