Jump to
S1C2
S1C3
Energy calculated at CCSD=FULL/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1591.041155 |
| Energy at 298.15K | |
| HF Energy | -1590.151491 |
| Nuclear repulsion energy | 351.592122 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
539 |
539 |
0.00 |
|
|
|
| 2 |
A1 |
210 |
210 |
0.00 |
|
|
|
| 3 |
B1 |
488 |
488 |
0.00 |
|
|
|
| 4 |
B2 |
322 |
322 |
0.00 |
|
|
|
| 5 |
E |
463 |
463 |
0.05 |
|
|
|
| 5 |
E |
463 |
463 |
0.05 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1242.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1242.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/daug-cc-pVTZ
Point Group is D2d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.425 |
0.315 |
| S2 |
0.000 |
-1.425 |
0.315 |
| S3 |
-1.425 |
0.000 |
-0.315 |
| S4 |
1.425 |
0.000 |
-0.315 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.8510 | 2.1121 | 2.1121 |
S2 | 2.8510 | | 2.1121 | 2.1121 | S3 | 2.1121 | 2.1121 | | 2.8510 | S4 | 2.1121 | 2.1121 | 2.8510 | |
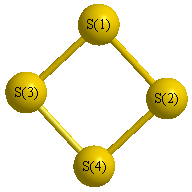 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
47.552 |
|
S1 |
S3 |
S4 |
47.552 |
| S2 |
S1 |
S3 |
47.552 |
|
S2 |
S4 |
S3 |
47.552 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCSD=FULL/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1591.046638 |
| Energy at 298.15K | |
| HF Energy | -1590.126295 |
| Nuclear repulsion energy | 334.667048 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
698 |
698 |
6.74 |
|
|
|
| 2 |
A1 |
448 |
448 |
5.22 |
|
|
|
| 3 |
A1 |
139 |
139 |
0.29 |
|
|
|
| 4 |
A2 |
218 |
218 |
|
|
|
|
| 5 |
B2 |
668 |
668 |
134.82 |
|
|
|
| 6 |
B2 |
343 |
343 |
0.24 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1256.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1256.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/daug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
1.031 |
0.914 |
| S2 |
0.000 |
-1.031 |
0.914 |
| S3 |
0.000 |
1.592 |
-0.914 |
| S4 |
0.000 |
-1.592 |
-0.914 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.0618 | 1.9123 | 3.1973 |
S2 | 2.0618 | | 3.1973 | 1.9123 | S3 | 1.9123 | 3.1973 | | 3.1845 | S4 | 3.1973 | 1.9123 | 3.1845 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
107.071 |
|
S1 |
S3 |
S4 |
72.929 |
| S2 |
S1 |
S3 |
107.071 |
|
S2 |
S4 |
S3 |
72.929 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCSD=FULL/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1591.035501 |
| Energy at 298.15K | |
| HF Energy | -1590.091229 |
| Nuclear repulsion energy | 338.509724 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD=FULL/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
762 |
762 |
0.00 |
|
|
|
| 2 |
Ag |
315 |
315 |
0.00 |
|
|
|
| 3 |
Au |
253 |
253 |
0.00 |
|
|
|
| 4 |
B1u |
697 |
697 |
151.34 |
|
|
|
| 5 |
B2u |
216i |
216i |
19.87 |
|
|
|
| 6 |
B3g |
347 |
347 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 1079.3 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 1079.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD=FULL/daug-cc-pVTZ
Point Group is D2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.944 |
1.249 |
| S2 |
0.000 |
0.944 |
-1.249 |
| S3 |
0.000 |
-0.944 |
1.249 |
| S4 |
0.000 |
-0.944 |
-1.249 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
S4 |
| S1 | | 2.4976 | 1.8881 | 3.1310 |
S2 | 2.4976 | | 3.1310 | 1.8881 | S3 | 1.8881 | 3.1310 | | 2.4976 | S4 | 3.1310 | 1.8881 | 2.4976 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
S4 |
90.000 |
|
S1 |
S3 |
S4 |
90.000 |
| S2 |
S1 |
S3 |
90.000 |
|
S2 |
S4 |
S3 |
90.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
