Jump to
S1C2
Energy calculated at B97D3/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -148.752368 |
| Energy at 298.15K | |
| HF Energy | -148.752368 |
| Nuclear repulsion energy | 59.177164 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3480 |
3480 |
27.66 |
160.66 |
0.09 |
0.16 |
| 2 |
A' |
2278 |
2278 |
101.13 |
73.59 |
0.16 |
0.28 |
| 3 |
A' |
1603 |
1603 |
35.27 |
9.65 |
0.52 |
0.68 |
| 4 |
A' |
1068 |
1068 |
9.85 |
11.40 |
0.06 |
0.11 |
| 5 |
A' |
570 |
570 |
167.89 |
1.55 |
0.62 |
0.76 |
| 6 |
A' |
483 |
483 |
37.73 |
1.05 |
0.41 |
0.58 |
| 7 |
A" |
3570 |
3570 |
52.94 |
49.94 |
0.75 |
0.86 |
| 8 |
A" |
1175 |
1175 |
0.00 |
0.13 |
0.75 |
0.86 |
| 9 |
A" |
396 |
396 |
0.03 |
1.35 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 7311.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7311.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/daug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.219 |
0.000 |
| N2 |
0.004 |
1.384 |
0.000 |
| N3 |
0.073 |
-1.123 |
0.000 |
| H4 |
-0.269 |
-1.568 |
0.843 |
| H5 |
-0.269 |
-1.568 |
-0.843 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1646 | 1.3441 | 1.9943 | 1.9943 |
N2 | 1.1646 | | 2.5077 | 3.0820 | 3.0820 | N3 | 1.3441 | 2.5077 | | 1.0126 | 1.0126 | H4 | 1.9943 | 3.0820 | 1.0126 | | 1.6854 | H5 | 1.9943 | 3.0820 | 1.0126 | 1.6854 | |
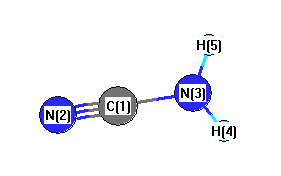 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
114.879 |
|
C1 |
N3 |
H5 |
114.879 |
| N2 |
C1 |
N3 |
176.688 |
|
H4 |
N3 |
H5 |
112.656 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.080 |
|
|
|
| 2 |
N |
-0.731 |
|
|
|
| 3 |
N |
0.130 |
|
|
|
| 4 |
H |
0.341 |
|
|
|
| 5 |
H |
0.341 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.975 |
-4.464 |
0.000 |
4.569 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.218 |
0.027 |
0.000 |
| y |
0.027 |
5.981 |
0.000 |
| z |
0.000 |
0.000 |
3.295 |
<r2> (average value of r
2) Å
2
| <r2> |
39.992 |
| (<r2>)1/2 |
6.324 |
Jump to
S1C1
Energy calculated at B97D3/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -148.751078 |
| Energy at 298.15K | |
| HF Energy | -148.751078 |
| Nuclear repulsion energy | 59.334350 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3554 |
3554 |
50.58 |
153.65 |
0.09 |
0.16 |
| 2 |
A1 |
2279 |
2279 |
130.67 |
73.97 |
0.17 |
0.29 |
| 3 |
A1 |
1586 |
1586 |
41.93 |
10.61 |
0.54 |
0.70 |
| 4 |
A1 |
1101 |
1101 |
14.51 |
12.59 |
0.06 |
0.11 |
| 5 |
B1 |
509 |
509 |
0.08 |
0.66 |
0.75 |
0.86 |
| 6 |
B1 |
417i |
417i |
219.31 |
0.05 |
0.75 |
0.86 |
| 7 |
B2 |
3665 |
3665 |
80.81 |
45.00 |
0.75 |
0.86 |
| 8 |
B2 |
1112 |
1112 |
1.24 |
0.01 |
0.75 |
0.86 |
| 9 |
B2 |
393 |
393 |
0.13 |
1.28 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 6891.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 6891.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/daug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.219 |
| N2 |
0.000 |
0.000 |
1.385 |
| N3 |
0.000 |
0.000 |
-1.110 |
| H4 |
0.000 |
0.866 |
-1.621 |
| H5 |
0.000 |
-0.866 |
-1.621 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
N3 |
H4 |
H5 |
| C1 | | 1.1660 | 1.3291 | 2.0343 | 2.0343 |
N2 | 1.1660 | | 2.4952 | 3.1290 | 3.1290 | N3 | 1.3291 | 2.4952 | | 1.0059 | 1.0059 | H4 | 2.0343 | 3.1290 | 1.0059 | | 1.7323 | H5 | 2.0343 | 3.1290 | 1.0059 | 1.7323 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H4 |
120.566 |
|
C1 |
N3 |
H5 |
120.566 |
| N2 |
C1 |
N3 |
180.000 |
|
H4 |
N3 |
H5 |
118.869 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.229 |
|
|
|
| 2 |
N |
-0.772 |
|
|
|
| 3 |
N |
0.301 |
|
|
|
| 4 |
H |
0.350 |
|
|
|
| 5 |
H |
0.350 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.810 |
4.810 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.211 |
0.000 |
0.000 |
| y |
0.000 |
3.294 |
0.000 |
| z |
0.000 |
0.000 |
6.052 |
<r2> (average value of r
2) Å
2
| <r2> |
39.926 |
| (<r2>)1/2 |
6.319 |
