return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C2H4S (Thiirane)

using model chemistry: QCISD(T)=FULL/daug-cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at QCISD(T)=FULL/daug-cc-pVTZ
 hartrees
Energy at 0K-476.273416
Energy at 298.15K 
HF Energy-475.612156
Nuclear repulsion energy101.180804
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/daug-cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3035 3035        
2 A1 1451 1451        
3 A1 1134 1134        
4 A1 640 640        
5 A1 556i 556i        
6 A2 3015 3015        
7 A2 855 855        
8 A2 859i 859i        
9 B1 3173 3173        
10 B1 414 414        
11 B1 503i 503i        
12 B2 2927 2927        
13 B2 1646 1646        
14 B2 667 667        
15 B2 826i 826i        

Unscaled Zero Point Vibrational Energy (zpe) 8106.1 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 8106.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)=FULL/daug-cc-pVTZ
ABC
0.73975 0.35889 0.26749

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)=FULL/daug-cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 0.000 0.000 0.866
C2 0.000 0.741 -0.798
C3 0.000 -0.741 -0.798
H4 -0.914 1.245 -1.071
H5 0.914 1.245 -1.071
H6 0.914 -1.245 -1.071
H7 -0.914 -1.245 -1.071

Atom - Atom Distances (Å)
  S1 C2 C3 H4 H5 H6 H7
S11.82181.82182.47732.47732.47732.4773
C21.82181.48151.07901.07902.20312.2031
C31.82181.48152.20312.20311.07901.0790
H42.47731.07902.20311.82833.08942.4903
H52.47731.07902.20311.82832.49033.0894
H62.47732.20311.07903.08942.49031.8283
H72.47732.20311.07902.49033.08941.8283

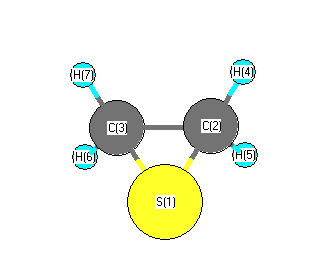
picture of Thiirane state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C3 66.008 S1 C2 H4 114.878
S1 C2 H5 114.878 S1 C3 C2 66.008
S1 C3 H6 114.878 S1 C3 H7 114.878
C2 S1 C3 47.984 C2 C3 H6 117.869
C2 C3 H7 117.869 C3 C2 H4 117.869
C3 C2 H5 117.869 H4 C2 H5 115.820
H6 C3 H7 115.820
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability