Jump to
S1C2
Energy calculated at CCSD/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -277.925742 |
| Energy at 298.15K | |
| HF Energy | -277.039911 |
| Nuclear repulsion energy | 125.939298 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/daug-cc-pVTZ
Geometric Data calculated at CCSD/daug-cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.425 |
0.627 |
0.000 |
| C2 |
-0.425 |
-0.627 |
0.000 |
| F3 |
-0.425 |
1.719 |
0.000 |
| F4 |
0.425 |
-1.719 |
0.000 |
| H5 |
1.051 |
0.672 |
0.891 |
| H6 |
1.051 |
0.672 |
-0.891 |
| H7 |
-1.051 |
-0.672 |
0.891 |
| H8 |
-1.051 |
-0.672 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5144 | 1.3841 | 2.3457 | 1.0897 | 1.0897 | 2.1584 | 2.1584 |
C2 | 1.5144 | | 2.3457 | 1.3841 | 2.1584 | 2.1584 | 1.0897 | 1.0897 | F3 | 1.3841 | 2.3457 | | 3.5415 | 2.0168 | 2.0168 | 2.6273 | 2.6273 | F4 | 2.3457 | 1.3841 | 3.5415 | | 2.6273 | 2.6273 | 2.0168 | 2.0168 | H5 | 1.0897 | 2.1584 | 2.0168 | 2.6273 | | 1.7820 | 2.4946 | 3.0657 | H6 | 1.0897 | 2.1584 | 2.0168 | 2.6273 | 1.7820 | | 3.0657 | 2.4946 | H7 | 2.1584 | 1.0897 | 2.6273 | 2.0168 | 2.4946 | 3.0657 | | 1.7820 | H8 | 2.1584 | 1.0897 | 2.6273 | 2.0168 | 3.0657 | 2.4946 | 1.7820 | |
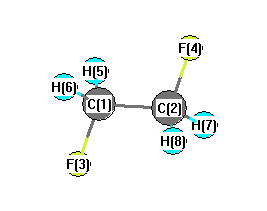 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.968 |
|
C1 |
C2 |
H7 |
110.905 |
| C1 |
C2 |
H8 |
110.905 |
|
C2 |
C1 |
F3 |
107.968 |
| C2 |
C1 |
H5 |
110.905 |
|
C2 |
C1 |
H6 |
110.905 |
| F3 |
C1 |
H5 |
108.645 |
|
F3 |
C1 |
H6 |
108.645 |
| F4 |
C2 |
H7 |
108.645 |
|
F4 |
C2 |
H8 |
108.645 |
| H5 |
C1 |
H6 |
109.700 |
|
H7 |
C2 |
H8 |
109.700 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -277.926927 |
| Energy at 298.15K | |
| HF Energy | -277.040518 |
| Nuclear repulsion energy | 128.066125 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/daug-cc-pVTZ
Geometric Data calculated at CCSD/daug-cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.266 |
0.703 |
0.511 |
| C2 |
-0.266 |
-0.703 |
0.511 |
| F3 |
-0.266 |
1.417 |
-0.546 |
| F4 |
0.266 |
-1.417 |
-0.546 |
| H5 |
-0.020 |
1.204 |
1.439 |
| H6 |
1.351 |
0.704 |
0.410 |
| H7 |
0.020 |
-1.204 |
1.439 |
| H8 |
-1.351 |
-0.704 |
0.410 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5033 | 1.3820 | 2.3692 | 1.0920 | 1.0903 | 2.1345 | 2.1457 |
C2 | 1.5033 | | 2.3692 | 1.3820 | 2.1345 | 2.1457 | 1.0920 | 1.0903 | F3 | 1.3820 | 2.3692 | | 2.8832 | 2.0115 | 2.0089 | 3.2999 | 2.5672 | F4 | 2.3692 | 1.3820 | 2.8832 | | 3.2999 | 2.5672 | 2.0115 | 2.0089 | H5 | 1.0920 | 2.1345 | 2.0115 | 3.2999 | | 1.7858 | 2.4076 | 2.5436 | H6 | 1.0903 | 2.1457 | 2.0089 | 2.5672 | 1.7858 | | 2.5436 | 3.0471 | H7 | 2.1345 | 1.0920 | 3.2999 | 2.0115 | 2.4076 | 2.5436 | | 1.7858 | H8 | 2.1457 | 1.0903 | 2.5672 | 2.0089 | 2.5436 | 3.0471 | 1.7858 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
110.326 |
|
C1 |
C2 |
H7 |
109.653 |
| C1 |
C2 |
H8 |
110.642 |
|
C2 |
C1 |
F3 |
110.326 |
| C2 |
C1 |
H5 |
109.653 |
|
C2 |
C1 |
H6 |
110.642 |
| F3 |
C1 |
H5 |
108.221 |
|
F3 |
C1 |
H6 |
108.116 |
| F4 |
C2 |
H7 |
108.221 |
|
F4 |
C2 |
H8 |
108.116 |
| H5 |
C1 |
H6 |
109.836 |
|
H7 |
C2 |
H8 |
109.836 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
