All results from a given calculation for C2H4F2 (1,2-difluoroethane)
using model chemistry: CCSD(T)=FULL/daug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C2H anti |
1Ag |
Energy calculated at CCSD(T)=FULL/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -278.026274 |
| Energy at 298.15K | |
| HF Energy | -277.039791 |
| Nuclear repulsion energy | 125.954175 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)=FULL/daug-cc-pVTZ
Geometric Data calculated at CCSD(T)=FULL/daug-cc-pVTZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.425 |
0.624 |
0.000 |
| C2 |
-0.425 |
-0.624 |
0.000 |
| F3 |
-0.425 |
1.719 |
0.000 |
| F4 |
0.425 |
-1.719 |
0.000 |
| H5 |
1.049 |
0.669 |
0.892 |
| H6 |
1.049 |
0.669 |
-0.892 |
| H7 |
-1.049 |
-0.669 |
0.892 |
| H8 |
-1.049 |
-0.669 |
-0.892 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5100 | 1.3865 | 2.3429 | 1.0892 | 1.0892 | 2.1537 | 2.1537 |
C2 | 1.5100 | | 2.3429 | 1.3865 | 2.1537 | 2.1537 | 1.0892 | 1.0892 | F3 | 1.3865 | 2.3429 | | 3.5417 | 2.0178 | 2.0178 | 2.6239 | 2.6239 | F4 | 2.3429 | 1.3865 | 3.5417 | | 2.6239 | 2.6239 | 2.0178 | 2.0178 | H5 | 1.0892 | 2.1537 | 2.0178 | 2.6239 | | 1.7837 | 2.4876 | 3.0610 | H6 | 1.0892 | 2.1537 | 2.0178 | 2.6239 | 1.7837 | | 3.0610 | 2.4876 | H7 | 2.1537 | 1.0892 | 2.6239 | 2.0178 | 2.4876 | 3.0610 | | 1.7837 | H8 | 2.1537 | 1.0892 | 2.6239 | 2.0178 | 3.0610 | 2.4876 | 1.7837 | |
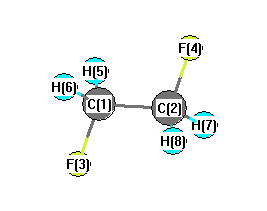 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
107.899 |
|
C1 |
C2 |
H7 |
110.874 |
| C1 |
C2 |
H8 |
110.874 |
|
C2 |
C1 |
F3 |
107.899 |
| C2 |
C1 |
H5 |
110.874 |
|
C2 |
C1 |
H6 |
110.874 |
| F3 |
C1 |
H5 |
108.590 |
|
F3 |
C1 |
H6 |
108.590 |
| F4 |
C2 |
H7 |
108.590 |
|
F4 |
C2 |
H8 |
108.590 |
| H5 |
C1 |
H6 |
109.933 |
|
H7 |
C2 |
H8 |
109.933 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
