Jump to
S1C2
Vibrational Frequencies calculated at CCSD/daug-cc-pVTZ
Geometric Data calculated at CCSD/daug-cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -189.484629 |
| Energy at 298.15K | |
| HF Energy | -188.837547 |
| Nuclear repulsion energy | 70.145967 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3859 |
3859 |
73.33 |
|
|
|
| 2 |
A' |
3036 |
3036 |
56.76 |
|
|
|
| 3 |
A' |
1889 |
1889 |
302.38 |
|
|
|
| 4 |
A' |
1438 |
1438 |
0.67 |
|
|
|
| 5 |
A' |
1311 |
1311 |
318.03 |
|
|
|
| 6 |
A' |
1138 |
1138 |
52.16 |
|
|
|
| 7 |
A' |
669 |
669 |
9.97 |
|
|
|
| 8 |
A" |
1052 |
1052 |
0.00 |
|
|
|
| 9 |
A" |
512 |
512 |
84.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7451.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 7451.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD/daug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.385 |
0.000 |
| O2 |
-0.895 |
-0.624 |
0.000 |
| O3 |
1.176 |
0.195 |
0.000 |
| H4 |
-0.458 |
1.384 |
0.000 |
| H5 |
-1.786 |
-0.261 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
O3 |
H4 |
H5 |
| C1 | | 1.3487 | 1.1912 | 1.0988 | 1.8989 |
O2 | 1.3487 | | 2.2273 | 2.0545 | 0.9614 | O3 | 1.1912 | 2.2273 | | 2.0206 | 2.9965 | H4 | 1.0988 | 2.0545 | 2.0206 | | 2.1136 | H5 | 1.8989 | 0.9614 | 2.9965 | 2.1136 | |
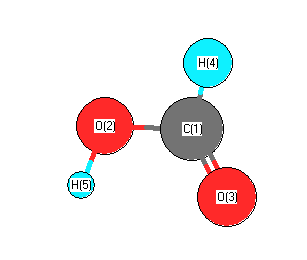 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.422 |
|
O2 |
C1 |
O3 |
122.419 |
| O2 |
C1 |
H4 |
113.772 |
|
O3 |
C1 |
H4 |
123.810 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
