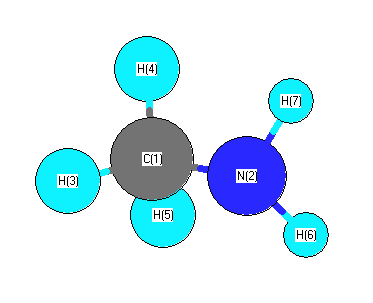
Vibrational Frequencies calculated at B97D3/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3431 |
3431 |
2.83 |
141.39 |
0.08 |
0.14 |
| 2 |
A' |
3006 |
3006 |
35.78 |
119.89 |
0.25 |
0.40 |
| 3 |
A' |
2907 |
2907 |
104.24 |
186.55 |
0.14 |
0.25 |
| 4 |
A' |
1634 |
1634 |
17.49 |
2.88 |
0.68 |
0.81 |
| 5 |
A' |
1467 |
1467 |
6.43 |
6.11 |
0.65 |
0.78 |
| 6 |
A' |
1426 |
1426 |
2.61 |
0.93 |
0.40 |
0.57 |
| 7 |
A' |
1146 |
1146 |
5.28 |
0.55 |
0.11 |
0.19 |
| 8 |
A' |
1024 |
1024 |
10.51 |
8.46 |
0.11 |
0.20 |
| 9 |
A' |
816 |
816 |
135.21 |
2.40 |
0.21 |
0.34 |
| 10 |
A" |
3510 |
3510 |
0.36 |
55.40 |
0.75 |
0.86 |
| 11 |
A" |
3044 |
3044 |
33.06 |
62.37 |
0.75 |
0.86 |
| 12 |
A" |
1488 |
1488 |
3.87 |
6.03 |
0.75 |
0.86 |
| 13 |
A" |
1322 |
1322 |
0.09 |
0.28 |
0.75 |
0.86 |
| 14 |
A" |
958 |
958 |
0.04 |
0.19 |
0.75 |
0.86 |
| 15 |
A" |
291 |
291 |
30.27 |
0.36 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13735.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13735.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.621 |
|
|
|
| 2 |
N |
-0.527 |
|
|
|
| 3 |
H |
0.309 |
|
|
|
| 4 |
H |
0.332 |
|
|
|
| 5 |
H |
0.332 |
|
|
|
| 6 |
H |
0.088 |
|
|
|
| 7 |
H |
0.088 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.225 |
0.351 |
0.000 |
1.275 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-15.225 |
2.127 |
0.000 |
| y |
2.127 |
-14.596 |
0.000 |
| z |
0.000 |
0.000 |
-12.780 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.537 |
2.127 |
0.000 |
| y |
2.127 |
-0.593 |
0.000 |
| z |
0.000 |
0.000 |
2.130 |
|
| Polar |
|---|
| 3z2-r2 | 4.260 |
|---|
| x2-y2 | -0.629 |
|---|
| xy | 2.127 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.931 |
-0.055 |
0.000 |
| y |
-0.055 |
4.509 |
0.000 |
| z |
0.000 |
0.000 |
3.750 |
<r2> (average value of r
2) Å
2
| <r2> |
26.985 |
| (<r2>)1/2 |
5.195 |