All results from a given calculation for CH2CHF (Ethene, fluoro-)
using model chemistry: CCSD/daug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at CCSD/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -177.562493 |
| Energy at 298.15K | |
| HF Energy | -176.956809 |
| Nuclear repulsion energy | 67.609173 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/daug-cc-pVTZ
Geometric Data calculated at CCSD/daug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.435 |
0.000 |
| C2 |
1.185 |
-0.153 |
0.000 |
| F3 |
-1.142 |
-0.270 |
0.000 |
| H4 |
-0.179 |
1.501 |
0.000 |
| H5 |
1.279 |
-1.229 |
0.000 |
| H6 |
2.072 |
0.460 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
F3 |
H4 |
H5 |
H6 |
| C1 | | 1.3226 | 1.3424 | 1.0803 | 2.0990 | 2.0723 |
C2 | 1.3226 | | 2.3297 | 2.1433 | 1.0798 | 1.0787 | F3 | 1.3424 | 2.3297 | | 2.0158 | 2.6045 | 3.2962 | H4 | 1.0803 | 2.1433 | 2.0158 | | 3.0945 | 2.4799 | H5 | 2.0990 | 1.0798 | 2.6045 | 3.0945 | | 1.8655 | H6 | 2.0723 | 1.0787 | 3.2962 | 2.4799 | 1.8655 | |
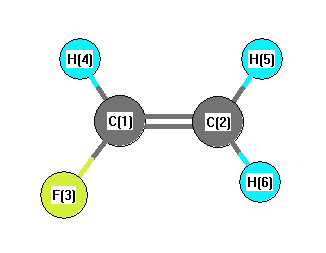 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.450 |
|
C1 |
C2 |
H6 |
118.958 |
| C2 |
C1 |
F3 |
121.894 |
|
C2 |
C1 |
H4 |
125.941 |
| F3 |
C1 |
H4 |
112.165 |
|
H5 |
C2 |
H6 |
119.592 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
