All results from a given calculation for C2H4O (Ethylene oxide)
using model chemistry: CCSD/daug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C2V |
1A1 |
Energy calculated at CCSD/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -153.532466 |
| Energy at 298.15K | |
| HF Energy | -152.928991 |
| Nuclear repulsion energy | 75.414281 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD/daug-cc-pVTZ
Geometric Data calculated at CCSD/daug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.852 |
| C2 |
0.000 |
0.732 |
-0.372 |
| C3 |
0.000 |
-0.732 |
-0.372 |
| H4 |
0.919 |
1.264 |
-0.587 |
| H5 |
-0.919 |
1.264 |
-0.587 |
| H6 |
-0.919 |
-1.264 |
-0.587 |
| H7 |
0.919 |
-1.264 |
-0.587 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
| O1 | | 1.4263 | 1.4263 | 2.1240 | 2.1240 | 2.1240 | 2.1240 |
C2 | 1.4263 | | 1.4640 | 1.0831 | 1.0831 | 2.2076 | 2.2076 | C3 | 1.4263 | 1.4640 | | 2.2076 | 2.2076 | 1.0831 | 1.0831 | H4 | 2.1240 | 1.0831 | 2.2076 | | 1.8376 | 3.1250 | 2.5276 | H5 | 2.1240 | 1.0831 | 2.2076 | 1.8376 | | 2.5276 | 3.1250 | H6 | 2.1240 | 2.2076 | 1.0831 | 3.1250 | 2.5276 | | 1.8376 | H7 | 2.1240 | 2.2076 | 1.0831 | 2.5276 | 3.1250 | 1.8376 | |
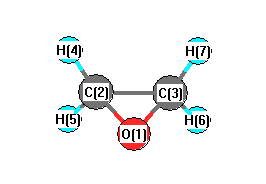 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
59.123 |
|
O1 |
C2 |
H4 |
114.958 |
| O1 |
C2 |
H5 |
114.958 |
|
O1 |
C3 |
C2 |
59.123 |
| O1 |
C3 |
H6 |
114.958 |
|
O1 |
C3 |
H7 |
114.958 |
| C2 |
O1 |
C3 |
61.754 |
|
C2 |
C3 |
H6 |
119.408 |
| C2 |
C3 |
H7 |
119.408 |
|
C3 |
C2 |
H4 |
119.408 |
| C3 |
C2 |
H5 |
119.408 |
|
H4 |
C2 |
H5 |
116.063 |
| H6 |
C3 |
H7 |
116.063 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
