Jump to
S1C2
Energy calculated at B97D3/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -244.984665 |
| Energy at 298.15K | |
| HF Energy | -244.984665 |
| Nuclear repulsion energy | 124.343237 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3121 |
3121 |
3.39 |
43.10 |
0.72 |
0.84 |
| 2 |
A' |
3029 |
3029 |
1.71 |
146.73 |
0.01 |
0.01 |
| 3 |
A' |
1446 |
1446 |
10.52 |
4.05 |
0.74 |
0.85 |
| 4 |
A' |
1381 |
1381 |
15.61 |
4.00 |
0.58 |
0.74 |
| 5 |
A' |
1343 |
1343 |
89.63 |
10.55 |
0.36 |
0.53 |
| 6 |
A' |
1101 |
1101 |
0.44 |
0.69 |
0.33 |
0.50 |
| 7 |
A' |
887 |
887 |
17.38 |
14.90 |
0.07 |
0.13 |
| 8 |
A' |
634 |
634 |
9.10 |
7.28 |
0.31 |
0.48 |
| 9 |
A' |
594 |
594 |
3.73 |
1.64 |
0.44 |
0.61 |
| 10 |
A" |
3151 |
3151 |
1.19 |
39.93 |
0.75 |
0.86 |
| 11 |
A" |
1559 |
1559 |
287.28 |
4.55 |
0.75 |
0.86 |
| 12 |
A" |
1433 |
1433 |
49.03 |
7.54 |
0.75 |
0.86 |
| 13 |
A" |
1080 |
1080 |
10.49 |
2.16 |
0.75 |
0.86 |
| 14 |
A" |
469 |
469 |
0.86 |
1.17 |
0.75 |
0.86 |
| 15 |
A" |
16i |
16i |
0.02 |
0.38 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 10605.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10605.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/daug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
-1.335 |
0.000 |
| N2 |
-0.010 |
0.178 |
0.000 |
| H3 |
1.050 |
-1.634 |
0.000 |
| H4 |
-0.493 |
-1.676 |
0.908 |
| H5 |
-0.493 |
-1.676 |
-0.908 |
| O6 |
0.000 |
0.734 |
-1.091 |
| O7 |
0.000 |
0.734 |
1.091 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
H5 |
O6 |
O7 |
| C1 | | 1.5127 | 1.0916 | 1.0879 | 1.0879 | 2.3390 | 2.3390 |
N2 | 1.5127 | | 2.0990 | 2.1195 | 2.1195 | 1.2245 | 1.2245 | H3 | 1.0916 | 2.0990 | | 1.7910 | 1.7910 | 2.8107 | 2.8107 | H4 | 1.0879 | 2.1195 | 1.7910 | | 1.8154 | 3.1693 | 2.4667 | H5 | 1.0879 | 2.1195 | 1.7910 | 1.8154 | | 2.4667 | 3.1693 | O6 | 2.3390 | 1.2245 | 2.8107 | 3.1693 | 2.4667 | | 2.1814 | O7 | 2.3390 | 1.2245 | 2.8107 | 2.4667 | 3.1693 | 2.1814 | |
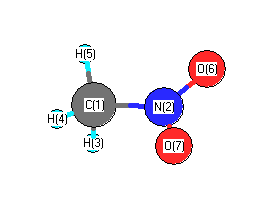 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
O6 |
117.028 |
|
C1 |
N2 |
O7 |
117.028 |
| N2 |
C1 |
H3 |
106.284 |
|
N2 |
C1 |
H4 |
108.066 |
| N2 |
C1 |
H5 |
108.066 |
|
H3 |
C1 |
H4 |
110.519 |
| H3 |
C1 |
H5 |
110.519 |
|
H4 |
C1 |
H5 |
113.092 |
| O6 |
N2 |
O7 |
125.927 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.493 |
|
|
|
| 2 |
N |
1.640 |
|
|
|
| 3 |
H |
0.377 |
|
|
|
| 4 |
H |
0.411 |
|
|
|
| 5 |
H |
0.411 |
|
|
|
| 6 |
O |
-1.173 |
|
|
|
| 7 |
O |
-1.173 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.024 |
-3.422 |
0.000 |
3.422 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.643 |
-0.036 |
0.000 |
| y |
-0.036 |
5.531 |
0.000 |
| z |
0.000 |
0.000 |
5.866 |
<r2> (average value of r
2) Å
2
| <r2> |
64.752 |
| (<r2>)1/2 |
8.047 |
Jump to
S1C1
Energy calculated at B97D3/daug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -244.984670 |
| Energy at 298.15K | |
| HF Energy | -244.984670 |
| Nuclear repulsion energy | 124.343724 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/daug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3150 |
3150 |
2.88 |
38.72 |
0.75 |
0.85 |
| 2 |
A' |
3029 |
3029 |
1.96 |
146.96 |
0.00 |
0.01 |
| 3 |
A' |
1559 |
1559 |
286.33 |
4.53 |
0.75 |
0.85 |
| 4 |
A' |
1433 |
1433 |
48.77 |
7.22 |
0.73 |
0.85 |
| 5 |
A' |
1381 |
1381 |
15.82 |
3.99 |
0.59 |
0.74 |
| 6 |
A' |
1343 |
1343 |
90.20 |
10.75 |
0.36 |
0.53 |
| 7 |
A' |
1079 |
1079 |
10.47 |
2.64 |
0.63 |
0.78 |
| 8 |
A' |
887 |
887 |
17.19 |
14.82 |
0.07 |
0.13 |
| 9 |
A' |
628 |
628 |
10.60 |
8.26 |
0.31 |
0.48 |
| 10 |
A' |
467 |
467 |
1.00 |
1.23 |
0.75 |
0.86 |
| 11 |
A" |
3120 |
3120 |
1.37 |
44.47 |
0.75 |
0.86 |
| 12 |
A" |
1447 |
1447 |
10.84 |
4.40 |
0.75 |
0.86 |
| 13 |
A" |
1103 |
1103 |
0.33 |
0.10 |
0.75 |
0.86 |
| 14 |
A" |
605 |
605 |
2.30 |
0.59 |
0.75 |
0.86 |
| 15 |
A" |
23 |
23 |
0.02 |
0.26 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 10626.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 10626.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/daug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.040 |
-1.334 |
0.000 |
| N2 |
0.000 |
0.178 |
0.000 |
| H3 |
1.080 |
-1.652 |
0.000 |
| H4 |
-0.492 |
-1.667 |
0.891 |
| H5 |
-0.492 |
-1.667 |
-0.891 |
| O6 |
1.069 |
0.775 |
0.000 |
| O7 |
-1.111 |
0.693 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
H5 |
O6 |
O7 |
| C1 | | 1.5123 | 1.0871 | 1.0903 | 1.0903 | 2.3464 | 2.3309 |
N2 | 1.5123 | | 2.1244 | 2.1070 | 2.1070 | 1.2245 | 1.2247 | H3 | 1.0871 | 2.1244 | | 1.8070 | 1.8070 | 2.4266 | 3.2088 | H4 | 1.0903 | 2.1070 | 1.8070 | | 1.7829 | 3.0321 | 2.5973 | H5 | 1.0903 | 2.1070 | 1.8070 | 1.7829 | | 3.0321 | 2.5973 | O6 | 2.3464 | 1.2245 | 2.4266 | 3.0321 | 3.0321 | | 2.1818 | O7 | 2.3309 | 1.2247 | 3.2088 | 2.5973 | 2.5973 | 2.1818 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
O6 |
117.661 |
|
C1 |
N2 |
O7 |
116.385 |
| N2 |
C1 |
H3 |
108.517 |
|
N2 |
C1 |
H4 |
106.994 |
| N2 |
C1 |
H5 |
106.994 |
|
H3 |
C1 |
H4 |
112.182 |
| H3 |
C1 |
H5 |
112.182 |
|
H4 |
C1 |
H5 |
109.694 |
| O6 |
N2 |
O7 |
125.954 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/daug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.552 |
|
|
|
| 2 |
N |
1.712 |
|
|
|
| 3 |
H |
0.408 |
|
|
|
| 4 |
H |
0.415 |
|
|
|
| 5 |
H |
0.415 |
|
|
|
| 6 |
O |
-1.093 |
|
|
|
| 7 |
O |
-1.305 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.096 |
-3.419 |
0.000 |
3.420 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.865 |
0.063 |
0.000 |
| y |
0.063 |
5.533 |
0.000 |
| z |
0.000 |
0.000 |
3.642 |
<r2> (average value of r
2) Å
2
| <r2> |
64.752 |
| (<r2>)1/2 |
8.047 |
