return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5CHO (benzaldehyde)

using model chemistry: B2PLYP=FULLultrafine/daug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
 hartrees
Energy at 0K-345.280742
Energy at 298.15K 
HF Energy-344.894757
Nuclear repulsion energy319.647326
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3226 3226 8.84 250.79 0.09 0.17
2 A' 3220 3220 9.69 83.41 0.15 0.25
3 A' 3211 3211 9.90 84.26 0.74 0.85
4 A' 3200 3200 1.06 81.42 0.69 0.82
5 A' 3189 3189 3.66 47.61 0.39 0.56
6 A' 2930 2930 106.67 145.35 0.28 0.43
7 A' 1735 1735 248.22 118.15 0.33 0.50
8 A' 1644 1644 28.45 94.24 0.55 0.71
9 A' 1626 1626 15.55 14.97 0.53 0.69
10 A' 1512 1512 1.32 1.23 0.48 0.65
11 A' 1475 1475 11.99 2.93 0.25 0.40
12 A' 1411 1411 5.06 3.15 0.37 0.54
13 A' 1385 1385 5.87 0.81 0.70 0.83
14 A' 1328 1328 15.21 1.36 0.31 0.48
15 A' 1229 1229 59.02 31.28 0.18 0.31
16 A' 1183 1183 15.15 8.06 0.12 0.22
17 A' 1176 1176 1.92 6.44 0.68 0.81
18 A' 1097 1097 4.62 0.90 0.09 0.17
19 A' 1041 1041 2.25 17.24 0.04 0.07
20 A' 1010 1010 0.47 37.57 0.04 0.08
21 A' 836 836 31.51 15.35 0.06 0.11
22 A' 655 655 21.80 3.11 0.29 0.44
23 A' 621 621 0.58 4.31 0.75 0.85
24 A' 440 440 0.28 6.20 0.19 0.32
25 A' 218 218 8.03 0.66 0.42 0.59
26 A" 1027 1027 1.21 1.70 0.75 0.86
27 A" 1012 1012 0.01 0.02 0.75 0.86
28 A" 999 999 0.00 0.06 0.75 0.86
29 A" 944 944 1.15 0.71 0.75 0.86
30 A" 870 870 0.01 0.44 0.75 0.86
31 A" 763 763 53.08 0.38 0.75 0.86
32 A" 697 697 23.08 0.00 0.75 0.85
33 A" 469 469 7.40 0.15 0.75 0.86
34 A" 417 417 0.07 0.02 0.75 0.86
35 A" 239 239 7.55 0.72 0.75 0.86
36 A" 119 119 4.70 1.35 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 24076.3 cm-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 24076.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULLultrafine/daug-cc-pVDZ
ABC
0.17347 0.05173 0.03985

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.575 0.000
C2 -1.049 -0.363 0.000
C3 -0.757 -1.728 0.000
C4 0.579 -2.162 0.000
C5 1.626 -1.231 0.000
C6 1.335 0.138 0.000
C7 -0.287 2.030 0.000
O8 -1.406 2.518 0.000
H9 0.615 2.683 0.000
H10 -2.076 -0.001 0.000
H11 -1.565 -2.458 0.000
H12 0.802 -3.228 0.000
H13 2.660 -1.572 0.000
H14 2.141 0.873 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 O8 H9 H10 H11 H12 H13 H14
C11.40692.42362.79702.42951.40421.48352.39882.19652.15443.41283.88653.41792.1614
C21.40691.39562.42622.81192.43562.51122.90303.47101.08892.15803.41123.90103.4207
C32.42361.39561.40492.43402.80293.78694.29504.61942.17271.08922.16403.42073.8937
C42.79702.42621.40491.40082.42084.28045.08354.84533.42312.16431.08952.16313.4130
C52.42952.81192.43401.40081.39953.78064.82154.04283.90063.41852.16051.08902.1660
C61.40422.43562.80292.42081.39952.49213.63002.64533.41353.89213.40812.16321.0908
C71.48352.51123.78694.28043.78062.49211.22061.11382.70644.66665.36994.65392.6897
O82.39882.90304.29505.08354.82153.63001.22062.02762.60664.97886.15605.76703.9098
H92.19653.47104.61944.84534.04282.64531.11382.02763.80085.58465.91464.72112.3678
H102.15441.08892.17273.42313.90063.41352.70642.60663.80082.50984.32414.98964.3063
H113.41282.15801.08922.16433.41853.89214.66664.97885.58462.50982.48914.31704.9829
H123.88653.41122.16401.08952.16053.40815.36996.15605.91464.32412.48912.48934.3141
H133.41793.90103.42072.16311.08902.16324.65395.76704.72114.98964.31702.48932.4991
H142.16143.42073.89373.41302.16601.09082.68973.90982.36784.30634.98294.31412.4991

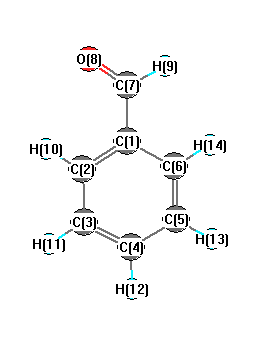
picture of benzaldehyde state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.718 C1 C2 H10 118.804
C1 C6 C5 120.118 C1 C6 H14 119.527
C1 C7 O8 124.737 C1 C7 H9 114.749
C2 C1 C6 120.097 C2 C1 C7 120.618
C2 C3 C4 120.071 C2 C3 H11 120.060
C3 C2 H10 121.477 C3 C4 C5 120.349
C3 C4 H12 119.818 C4 C3 H11 119.869
C4 C5 C6 119.647 C4 C5 H13 120.121
C5 C4 H12 119.833 C5 C6 H14 120.355
C6 C1 C7 119.286 C6 C5 H13 120.231
O8 C7 H9 120.515
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability