Jump to
S1C2
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -191.756791 |
| Energy at 298.15K | |
| HF Energy | -191.551328 |
| Nuclear repulsion energy | 102.559181 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3268 |
3268 |
4.21 |
65.61 |
0.54 |
0.70 |
| 2 |
A' |
3214 |
3214 |
1.11 |
100.68 |
0.27 |
0.42 |
| 3 |
A' |
3163 |
3163 |
4.46 |
91.01 |
0.08 |
0.14 |
| 4 |
A' |
2927 |
2927 |
85.04 |
114.91 |
0.27 |
0.42 |
| 5 |
A' |
1739 |
1739 |
236.53 |
102.55 |
0.33 |
0.50 |
| 6 |
A' |
1668 |
1668 |
4.93 |
29.16 |
0.08 |
0.14 |
| 7 |
A' |
1452 |
1452 |
10.23 |
6.42 |
0.46 |
0.63 |
| 8 |
A' |
1379 |
1379 |
4.66 |
21.58 |
0.27 |
0.43 |
| 9 |
A' |
1296 |
1296 |
2.84 |
11.83 |
0.14 |
0.25 |
| 10 |
A' |
1177 |
1177 |
38.31 |
12.13 |
0.64 |
0.78 |
| 11 |
A' |
923 |
923 |
17.35 |
1.78 |
0.00 |
0.00 |
| 12 |
A' |
569 |
569 |
4.27 |
6.67 |
0.16 |
0.28 |
| 13 |
A' |
321 |
321 |
10.43 |
1.05 |
0.31 |
0.48 |
| 14 |
A" |
1045 |
1045 |
9.68 |
0.83 |
0.75 |
0.86 |
| 15 |
A" |
1031 |
1031 |
8.15 |
2.03 |
0.75 |
0.86 |
| 16 |
A" |
1011 |
1011 |
39.55 |
1.76 |
0.75 |
0.86 |
| 17 |
A" |
623 |
623 |
9.82 |
0.70 |
0.75 |
0.86 |
| 18 |
A" |
171 |
171 |
4.51 |
0.47 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13488.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13488.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.151 |
-0.745 |
0.000 |
| C2 |
0.000 |
0.723 |
0.000 |
| C3 |
1.223 |
1.285 |
0.000 |
| O4 |
-1.225 |
-1.325 |
0.000 |
| H5 |
0.806 |
-1.316 |
0.000 |
| H6 |
-0.915 |
1.315 |
0.000 |
| H7 |
1.363 |
2.364 |
0.000 |
| H8 |
2.120 |
0.665 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4760 | 2.4510 | 1.2205 | 1.1147 | 2.1979 | 3.4582 | 2.6736 |
C2 | 1.4760 | | 1.3454 | 2.3862 | 2.1927 | 1.0904 | 2.1330 | 2.1211 | C3 | 2.4510 | 1.3454 | | 3.5777 | 2.6339 | 2.1381 | 1.0883 | 1.0910 | O4 | 1.2205 | 2.3862 | 3.5777 | | 2.0317 | 2.6582 | 4.5059 | 3.8925 | H5 | 1.1147 | 2.1927 | 2.6339 | 2.0317 | | 3.1447 | 3.7219 | 2.3772 | H6 | 2.1979 | 1.0904 | 2.1381 | 2.6582 | 3.1447 | | 2.5078 | 3.1046 | H7 | 3.4582 | 2.1330 | 1.0883 | 4.5059 | 3.7219 | 2.5078 | | 1.8603 | H8 | 2.6736 | 2.1211 | 1.0910 | 3.8925 | 2.3772 | 3.1046 | 1.8603 | |
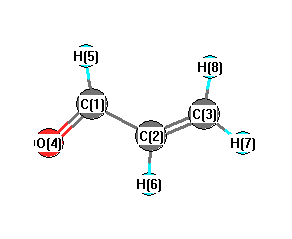 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.553 |
|
C1 |
C2 |
H6 |
117.040 |
| C2 |
C1 |
O4 |
124.210 |
|
C2 |
C1 |
H5 |
114.929 |
| C2 |
C3 |
H7 |
122.081 |
|
C2 |
C3 |
H8 |
120.704 |
| C3 |
C2 |
H6 |
122.406 |
|
O4 |
C1 |
H5 |
120.862 |
| H7 |
C3 |
H8 |
117.215 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -191.753164 |
| Energy at 298.15K | |
| HF Energy | -191.547558 |
| Nuclear repulsion energy | 104.039177 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3279 |
3279 |
1.49 |
54.25 |
0.62 |
0.76 |
| 2 |
A' |
3201 |
3201 |
10.56 |
127.95 |
0.22 |
0.36 |
| 3 |
A' |
3172 |
3172 |
4.49 |
72.58 |
0.10 |
0.18 |
| 4 |
A' |
2952 |
2952 |
129.24 |
184.74 |
0.28 |
0.44 |
| 5 |
A' |
1743 |
1743 |
94.71 |
16.11 |
0.66 |
0.79 |
| 6 |
A' |
1661 |
1661 |
78.18 |
75.30 |
0.14 |
0.25 |
| 7 |
A' |
1434 |
1434 |
40.21 |
6.86 |
0.52 |
0.68 |
| 8 |
A' |
1415 |
1415 |
1.06 |
11.04 |
0.11 |
0.20 |
| 9 |
A' |
1310 |
1310 |
2.64 |
21.70 |
0.25 |
0.40 |
| 10 |
A' |
1065 |
1065 |
3.37 |
3.08 |
0.72 |
0.83 |
| 11 |
A' |
940 |
940 |
59.42 |
7.17 |
0.06 |
0.12 |
| 12 |
A' |
678 |
678 |
12.62 |
0.70 |
0.75 |
0.86 |
| 13 |
A' |
283 |
283 |
6.89 |
3.48 |
0.31 |
0.47 |
| 14 |
A" |
1044 |
1044 |
5.09 |
3.99 |
0.75 |
0.86 |
| 15 |
A" |
1029 |
1029 |
43.39 |
1.43 |
0.75 |
0.86 |
| 16 |
A" |
1020 |
1020 |
0.80 |
0.10 |
0.75 |
0.86 |
| 17 |
A" |
564 |
564 |
10.17 |
0.80 |
0.75 |
0.86 |
| 18 |
A" |
156 |
156 |
6.35 |
0.27 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13473.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13473.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.893 |
-0.290 |
0.000 |
| C2 |
0.000 |
0.899 |
0.000 |
| C3 |
1.339 |
0.779 |
0.000 |
| O4 |
-0.501 |
-1.447 |
0.000 |
| H5 |
-1.983 |
-0.070 |
0.000 |
| H6 |
-0.482 |
1.878 |
0.000 |
| H7 |
1.991 |
1.650 |
0.000 |
| H8 |
1.799 |
-0.210 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4869 | 2.4747 | 1.2211 | 1.1121 | 2.2065 | 3.4756 | 2.6927 |
C2 | 1.4869 | | 1.3449 | 2.3986 | 2.2066 | 1.0910 | 2.1279 | 2.1132 | C3 | 2.4747 | 1.3449 | | 2.8876 | 3.4286 | 2.1273 | 1.0883 | 1.0899 | O4 | 1.2211 | 2.3986 | 2.8876 | | 2.0228 | 3.3246 | 3.9750 | 2.6113 | H5 | 1.1121 | 2.2066 | 3.4286 | 2.0228 | | 2.4587 | 4.3297 | 3.7841 | H6 | 2.2065 | 1.0910 | 2.1273 | 3.3246 | 2.4587 | | 2.4830 | 3.0919 | H7 | 3.4756 | 2.1279 | 1.0883 | 3.9750 | 4.3297 | 2.4830 | | 1.8700 | H8 | 2.6927 | 2.1132 | 1.0899 | 2.6113 | 3.7841 | 3.0919 | 1.8700 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.754 |
|
C1 |
C2 |
H6 |
116.900 |
| C2 |
C1 |
O4 |
124.394 |
|
C2 |
C1 |
H5 |
115.457 |
| C2 |
C3 |
H7 |
121.622 |
|
C2 |
C3 |
H8 |
120.073 |
| C3 |
C2 |
H6 |
121.345 |
|
O4 |
C1 |
H5 |
120.149 |
| H7 |
C3 |
H8 |
118.304 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
