Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -218.195169 |
| Energy at 298.15K | |
| HF Energy | -217.976483 |
| Nuclear repulsion energy | 129.633373 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3144 |
3144 |
23.65 |
|
|
|
| 2 |
A |
3132 |
3132 |
55.61 |
|
|
|
| 3 |
A |
3127 |
3127 |
27.16 |
|
|
|
| 4 |
A |
3097 |
3097 |
15.53 |
|
|
|
| 5 |
A |
3068 |
3068 |
33.99 |
|
|
|
| 6 |
A |
3053 |
3053 |
25.21 |
|
|
|
| 7 |
A |
3050 |
3050 |
18.16 |
|
|
|
| 8 |
A |
1501 |
1501 |
2.09 |
|
|
|
| 9 |
A |
1495 |
1495 |
8.16 |
|
|
|
| 10 |
A |
1483 |
1483 |
5.33 |
|
|
|
| 11 |
A |
1462 |
1462 |
2.56 |
|
|
|
| 12 |
A |
1409 |
1409 |
7.36 |
|
|
|
| 13 |
A |
1401 |
1401 |
4.96 |
|
|
|
| 14 |
A |
1367 |
1367 |
0.74 |
|
|
|
| 15 |
A |
1295 |
1295 |
1.20 |
|
|
|
| 16 |
A |
1262 |
1262 |
1.07 |
|
|
|
| 17 |
A |
1170 |
1170 |
1.44 |
|
|
|
| 18 |
A |
1118 |
1118 |
3.68 |
|
|
|
| 19 |
A |
1081 |
1081 |
43.44 |
|
|
|
| 20 |
A |
962 |
962 |
63.26 |
|
|
|
| 21 |
A |
913 |
913 |
4.49 |
|
|
|
| 22 |
A |
880 |
880 |
6.57 |
|
|
|
| 23 |
A |
760 |
760 |
1.02 |
|
|
|
| 24 |
A |
475 |
475 |
4.39 |
|
|
|
| 25 |
A |
315 |
315 |
1.30 |
|
|
|
| 26 |
A |
215 |
215 |
1.67 |
|
|
|
| 27 |
A |
139 |
139 |
2.42 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21187.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 21187.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.791 |
0.521 |
0.302 |
| C2 |
-0.598 |
0.651 |
-0.290 |
| C3 |
-1.537 |
-0.486 |
0.120 |
| F4 |
1.411 |
-0.657 |
-0.171 |
| H5 |
1.440 |
1.357 |
0.012 |
| H6 |
0.762 |
0.442 |
1.398 |
| H7 |
-0.511 |
0.703 |
-1.384 |
| H8 |
-1.003 |
1.619 |
0.045 |
| H9 |
-2.530 |
-0.353 |
-0.326 |
| H10 |
-1.142 |
-1.455 |
-0.209 |
| H11 |
-1.658 |
-0.521 |
1.211 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.5155 | 2.5428 | 1.4133 | 1.0974 | 1.0988 | 2.1381 | 2.1188 | 3.4916 | 2.8110 | 2.8122 |
C2 | 1.5155 | | 1.5301 | 2.4007 | 2.1781 | 2.1777 | 1.0990 | 1.1013 | 2.1780 | 2.1763 | 2.1792 | C3 | 2.5428 | 1.5301 | | 2.9673 | 3.5029 | 2.7891 | 2.1740 | 2.1730 | 1.0972 | 1.0964 | 1.0988 | F4 | 1.4133 | 2.4007 | 2.9673 | | 2.0227 | 2.0224 | 2.6486 | 3.3251 | 3.9565 | 2.6753 | 3.3690 | H5 | 1.0974 | 2.1781 | 3.5029 | 2.0227 | | 1.7938 | 2.4869 | 2.4572 | 4.3365 | 3.8239 | 3.8159 | H6 | 1.0988 | 2.1777 | 2.7891 | 2.0224 | 1.7938 | | 3.0706 | 2.5166 | 3.8007 | 3.1311 | 2.6111 | H7 | 2.1381 | 1.0990 | 2.1740 | 2.6486 | 2.4869 | 3.0706 | | 1.7674 | 2.5126 | 2.5363 | 3.0899 | H8 | 2.1188 | 1.1013 | 2.1730 | 3.3251 | 2.4572 | 2.5166 | 1.7674 | | 2.5221 | 3.0873 | 2.5235 | H9 | 3.4916 | 2.1780 | 1.0972 | 3.9565 | 4.3365 | 3.8007 | 2.5126 | 2.5221 | | 1.7761 | 1.7755 | H10 | 2.8110 | 2.1763 | 1.0964 | 2.6753 | 3.8239 | 3.1311 | 2.5363 | 3.0873 | 1.7761 | | 1.7764 | H11 | 2.8122 | 2.1792 | 1.0988 | 3.3690 | 3.8159 | 2.6111 | 3.0899 | 2.5235 | 1.7755 | 1.7764 | |
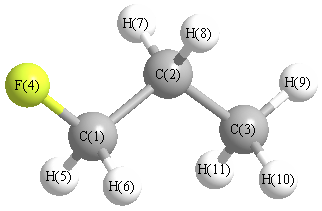 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.205 |
|
C1 |
C2 |
H7 |
108.679 |
| C1 |
C2 |
H8 |
107.067 |
|
C2 |
C1 |
F4 |
110.054 |
| C2 |
C1 |
H5 |
111.946 |
|
C2 |
C1 |
H6 |
111.826 |
| C2 |
C3 |
H9 |
110.917 |
|
C2 |
C3 |
H10 |
110.822 |
| C2 |
C3 |
H11 |
110.909 |
|
C3 |
C2 |
H7 |
110.485 |
| C3 |
C2 |
H8 |
110.272 |
|
F4 |
C1 |
H5 |
106.665 |
| F4 |
C1 |
H6 |
106.558 |
|
H5 |
C1 |
H6 |
109.523 |
| H7 |
C2 |
H8 |
106.880 |
|
H9 |
C3 |
H10 |
108.129 |
| H9 |
C3 |
H11 |
107.900 |
|
H10 |
C3 |
H11 |
108.042 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
