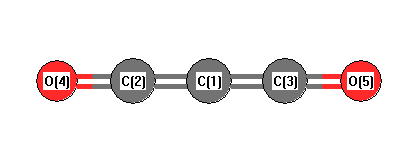
Vibrational Frequencies calculated at PBEPBE/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σg |
2189 |
2189 |
0.00 |
68.78 |
0.48 |
0.65 |
| 2 |
Σg |
759 |
759 |
0.00 |
45.20 |
0.16 |
0.28 |
| 3 |
Σu |
2372 |
2372 |
2346.58 |
0.00 |
0.00 |
0.00 |
| 4 |
Σu |
1582 |
1582 |
113.84 |
0.00 |
0.00 |
0.00 |
| 5 |
Πg |
547 |
547 |
0.00 |
0.95 |
0.75 |
0.86 |
| 5 |
Πg |
547 |
547 |
0.00 |
0.95 |
0.75 |
0.86 |
| 6 |
Πu |
482 |
482 |
31.08 |
0.00 |
0.00 |
0.00 |
| 6 |
Πu |
482 |
482 |
31.08 |
0.00 |
0.00 |
0.00 |
| 7 |
Πu |
23 |
23 |
0.07 |
0.00 |
0.00 |
0.00 |
| 7 |
Πu |
23 |
23 |
0.07 |
0.00 |
0.00 |
0.00 |
Unscaled Zero Point Vibrational Energy (zpe) 4503.1 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 4503.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/daug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
2.093 |
|
|
|
| 2 |
C |
-0.201 |
|
|
|
| 3 |
C |
-0.201 |
|
|
|
| 4 |
O |
-0.845 |
|
|
|
| 5 |
O |
-0.845 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.115 |
0.000 |
0.000 |
| y |
0.000 |
-26.115 |
0.000 |
| z |
0.000 |
0.000 |
-33.316 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.601 |
0.000 |
0.000 |
| y |
0.000 |
3.601 |
0.000 |
| z |
0.000 |
0.000 |
-7.201 |
|
| Polar |
|---|
| 3z2-r2 | -14.402 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.804 |
0.000 |
0.000 |
| y |
0.000 |
3.804 |
0.000 |
| z |
0.000 |
0.000 |
13.144 |
<r2> (average value of r
2) Å
2
| <r2> |
135.062 |
| (<r2>)1/2 |
11.622 |