Jump to
S1C2
Vibrational Frequencies calculated at PBEPBE/daug-cc-pVDZ
Geometric Data calculated at PBEPBE/daug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at PBEPBE/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -194.125411 |
| Energy at 298.15K | |
| HF Energy | -194.125411 |
| Nuclear repulsion energy | 131.577728 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at PBEPBE/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3719 |
3719 |
21.89 |
126.28 |
0.22 |
0.36 |
| 2 |
A |
3063 |
3063 |
23.44 |
41.69 |
0.75 |
0.85 |
| 3 |
A |
3043 |
3043 |
43.96 |
58.58 |
0.56 |
0.71 |
| 4 |
A |
3011 |
3011 |
17.93 |
93.82 |
0.71 |
0.83 |
| 5 |
A |
2972 |
2972 |
59.34 |
97.75 |
0.09 |
0.16 |
| 6 |
A |
2967 |
2967 |
18.28 |
246.72 |
0.01 |
0.01 |
| 7 |
A |
2942 |
2942 |
38.31 |
106.62 |
0.75 |
0.85 |
| 8 |
A |
2903 |
2903 |
68.96 |
155.20 |
0.07 |
0.13 |
| 9 |
A |
1440 |
1440 |
2.13 |
3.73 |
0.66 |
0.80 |
| 10 |
A |
1431 |
1431 |
8.50 |
1.93 |
0.73 |
0.85 |
| 11 |
A |
1418 |
1418 |
7.57 |
7.34 |
0.74 |
0.85 |
| 12 |
A |
1398 |
1398 |
3.25 |
7.62 |
0.75 |
0.86 |
| 13 |
A |
1376 |
1376 |
3.71 |
0.49 |
0.50 |
0.67 |
| 14 |
A |
1338 |
1338 |
4.48 |
0.38 |
0.69 |
0.82 |
| 15 |
A |
1305 |
1305 |
0.78 |
0.86 |
0.69 |
0.82 |
| 16 |
A |
1268 |
1268 |
18.88 |
4.77 |
0.75 |
0.86 |
| 17 |
A |
1211 |
1211 |
0.62 |
3.52 |
0.70 |
0.82 |
| 18 |
A |
1198 |
1198 |
26.23 |
3.05 |
0.16 |
0.28 |
| 19 |
A |
1109 |
1109 |
3.07 |
1.71 |
0.18 |
0.31 |
| 20 |
A |
1076 |
1076 |
23.11 |
3.37 |
0.51 |
0.67 |
| 21 |
A |
1045 |
1045 |
23.69 |
2.73 |
0.72 |
0.84 |
| 22 |
A |
946 |
946 |
58.05 |
2.62 |
0.73 |
0.85 |
| 23 |
A |
883 |
883 |
1.75 |
0.35 |
0.41 |
0.58 |
| 24 |
A |
852 |
852 |
1.12 |
10.84 |
0.09 |
0.16 |
| 25 |
A |
735 |
735 |
0.84 |
0.22 |
0.54 |
0.70 |
| 26 |
A |
459 |
459 |
7.31 |
0.27 |
0.40 |
0.57 |
| 27 |
A |
316 |
316 |
8.29 |
0.51 |
0.40 |
0.57 |
| 28 |
A |
248 |
248 |
91.86 |
1.89 |
0.72 |
0.83 |
| 29 |
A |
208 |
208 |
1.86 |
0.09 |
0.36 |
0.53 |
| 30 |
A |
130 |
130 |
8.13 |
0.14 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 23004.6 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 23004.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at PBEPBE/daug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.554 |
-0.515 |
0.130 |
| C2 |
-0.636 |
0.636 |
-0.295 |
| C3 |
0.766 |
0.548 |
0.296 |
| O4 |
1.401 |
-0.638 |
-0.218 |
| H5 |
-2.558 |
-0.410 |
-0.321 |
| H6 |
-1.679 |
-0.537 |
1.230 |
| H7 |
-1.137 |
-1.488 |
-0.184 |
| H8 |
-0.548 |
0.666 |
-1.399 |
| H9 |
-1.071 |
1.607 |
0.015 |
| H10 |
0.705 |
0.505 |
1.407 |
| H11 |
1.350 |
1.453 |
0.019 |
| H12 |
2.281 |
-0.711 |
0.188 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5325 | 2.5579 | 2.9779 | 1.1052 | 1.1071 | 1.1041 | 2.1780 | 2.1793 | 2.7888 | 3.5096 | 3.8408 |
C2 | 1.5325 | | 1.5247 | 2.4035 | 2.1884 | 2.1885 | 2.1849 | 1.1079 | 1.1082 | 2.1712 | 2.1701 | 3.2493 | C3 | 2.5579 | 1.5247 | | 1.4393 | 3.5143 | 2.8333 | 2.8277 | 2.1485 | 2.1394 | 1.1134 | 1.1118 | 1.9723 | O4 | 2.9779 | 2.4035 | 1.4393 | | 3.9665 | 3.4041 | 2.6763 | 2.6250 | 3.3469 | 2.1049 | 2.1041 | 0.9721 | H5 | 1.1052 | 2.1884 | 3.5143 | 3.9665 | | 1.7870 | 1.7886 | 2.5220 | 2.5287 | 3.8043 | 4.3425 | 4.8750 | H6 | 1.1071 | 2.1885 | 2.8333 | 3.4041 | 1.7870 | | 1.7874 | 3.1042 | 2.5383 | 2.6080 | 3.8208 | 4.0982 | H7 | 1.1041 | 2.1849 | 2.8277 | 2.6763 | 1.7886 | 1.7874 | | 2.5416 | 3.1019 | 3.1459 | 3.8562 | 3.5247 | H8 | 2.1780 | 1.1079 | 2.1485 | 2.6250 | 2.5220 | 3.1042 | 2.5416 | | 1.7779 | 3.0776 | 2.4965 | 3.5237 | H9 | 2.1793 | 1.1082 | 2.1394 | 3.3469 | 2.5287 | 2.5383 | 3.1019 | 1.7779 | | 2.5111 | 2.4257 | 4.0790 | H10 | 2.7888 | 2.1712 | 1.1134 | 2.1049 | 3.8043 | 2.6080 | 3.1459 | 3.0776 | 2.5111 | | 1.8001 | 2.3343 | H11 | 3.5096 | 2.1701 | 1.1118 | 2.1041 | 4.3425 | 3.8208 | 3.8562 | 2.4965 | 2.4257 | 1.8001 | | 2.3614 | H12 | 3.8408 | 3.2493 | 1.9723 | 0.9721 | 4.8750 | 4.0982 | 3.5247 | 3.5237 | 4.0790 | 2.3343 | 2.3614 | |
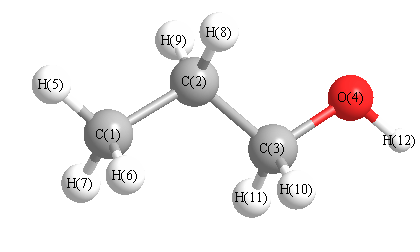 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.578 |
|
C1 |
C2 |
H8 |
110.116 |
| C1 |
C2 |
H9 |
110.198 |
|
C2 |
C1 |
H5 |
111.097 |
| C2 |
C1 |
H6 |
110.986 |
|
C2 |
C1 |
H7 |
110.881 |
| C2 |
C3 |
O4 |
108.332 |
|
C2 |
C3 |
H10 |
109.796 |
| C2 |
C3 |
H11 |
109.803 |
|
C3 |
C2 |
H8 |
108.354 |
| C3 |
C2 |
H9 |
107.635 |
|
C3 |
O4 |
H12 |
108.185 |
| O4 |
C3 |
H10 |
110.441 |
|
O4 |
C3 |
H11 |
110.478 |
| H5 |
C1 |
H6 |
107.756 |
|
H5 |
C1 |
H7 |
108.113 |
| H6 |
C1 |
H7 |
107.869 |
|
H8 |
C2 |
H9 |
106.694 |
| H10 |
C3 |
H11 |
107.990 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at PBEPBE/daug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.842 |
|
|
|
| 2 |
C |
0.302 |
|
|
|
| 3 |
C |
2.190 |
|
|
|
| 4 |
O |
-0.714 |
|
|
|
| 5 |
H |
-0.353 |
|
|
|
| 6 |
H |
-0.217 |
|
|
|
| 7 |
H |
-0.147 |
|
|
|
| 8 |
H |
-0.457 |
|
|
|
| 9 |
H |
-0.688 |
|
|
|
| 10 |
H |
-0.555 |
|
|
|
| 11 |
H |
-0.646 |
|
|
|
| 12 |
H |
0.443 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.570 |
0.934 |
0.915 |
1.426 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.687 |
-0.148 |
1.994 |
| y |
-0.148 |
-27.852 |
-0.537 |
| z |
1.994 |
-0.537 |
-27.482 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.980 |
-0.148 |
1.994 |
| y |
-0.148 |
-2.267 |
-0.537 |
| z |
1.994 |
-0.537 |
-1.713 |
|
| Polar |
|---|
| 3z2-r2 | -3.426 |
|---|
| x2-y2 | 4.165 |
|---|
| xy | -0.148 |
|---|
| xz | 1.994 |
|---|
| yz | -0.537 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.888 |
0.098 |
0.073 |
| y |
0.098 |
7.132 |
-0.010 |
| z |
0.073 |
-0.010 |
6.696 |
<r2> (average value of r
2) Å
2
| <r2> |
97.008 |
| (<r2>)1/2 |
9.849 |
