Jump to
S1C2
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -194.179210 |
| Energy at 298.15K | |
| HF Energy | -193.959013 |
| Nuclear repulsion energy | 132.216612 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3842 |
3842 |
31.29 |
112.71 |
0.21 |
0.35 |
| 2 |
A |
3146 |
3146 |
25.42 |
35.61 |
0.75 |
0.86 |
| 3 |
A |
3122 |
3122 |
49.73 |
54.92 |
0.55 |
0.71 |
| 4 |
A |
3096 |
3096 |
20.12 |
81.24 |
0.72 |
0.84 |
| 5 |
A |
3061 |
3061 |
76.07 |
123.72 |
0.15 |
0.26 |
| 6 |
A |
3049 |
3049 |
29.64 |
201.33 |
0.02 |
0.04 |
| 7 |
A |
3046 |
3046 |
5.62 |
93.13 |
0.45 |
0.62 |
| 8 |
A |
3010 |
3010 |
60.63 |
126.70 |
0.08 |
0.15 |
| 9 |
A |
1508 |
1508 |
1.87 |
3.24 |
0.73 |
0.84 |
| 10 |
A |
1496 |
1496 |
7.80 |
1.60 |
0.75 |
0.86 |
| 11 |
A |
1482 |
1482 |
6.90 |
6.52 |
0.75 |
0.85 |
| 12 |
A |
1465 |
1465 |
2.23 |
7.54 |
0.73 |
0.85 |
| 13 |
A |
1437 |
1437 |
4.22 |
0.75 |
0.25 |
0.40 |
| 14 |
A |
1402 |
1402 |
3.49 |
0.33 |
0.75 |
0.86 |
| 15 |
A |
1367 |
1367 |
1.18 |
0.79 |
0.71 |
0.83 |
| 16 |
A |
1317 |
1317 |
17.42 |
4.86 |
0.72 |
0.84 |
| 17 |
A |
1264 |
1264 |
0.28 |
2.90 |
0.72 |
0.83 |
| 18 |
A |
1242 |
1242 |
34.41 |
2.02 |
0.23 |
0.38 |
| 19 |
A |
1157 |
1157 |
4.34 |
1.60 |
0.11 |
0.19 |
| 20 |
A |
1110 |
1110 |
15.27 |
3.92 |
0.48 |
0.65 |
| 21 |
A |
1077 |
1077 |
32.73 |
3.03 |
0.65 |
0.79 |
| 22 |
A |
977 |
977 |
53.30 |
3.60 |
0.69 |
0.81 |
| 23 |
A |
918 |
918 |
2.05 |
0.34 |
0.55 |
0.71 |
| 24 |
A |
874 |
874 |
0.73 |
13.18 |
0.08 |
0.14 |
| 25 |
A |
765 |
765 |
0.97 |
0.32 |
0.29 |
0.46 |
| 26 |
A |
475 |
475 |
7.87 |
0.26 |
0.49 |
0.66 |
| 27 |
A |
325 |
325 |
8.47 |
0.39 |
0.33 |
0.50 |
| 28 |
A |
257 |
257 |
96.45 |
1.47 |
0.71 |
0.83 |
| 29 |
A |
221 |
221 |
4.25 |
0.07 |
0.44 |
0.61 |
| 30 |
A |
135 |
135 |
8.55 |
0.14 |
0.73 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 23820.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 23820.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.542 |
-0.518 |
0.129 |
| C2 |
-0.635 |
0.641 |
-0.293 |
| C3 |
0.765 |
0.551 |
0.295 |
| O4 |
1.386 |
-0.639 |
-0.215 |
| H5 |
-2.543 |
-0.413 |
-0.308 |
| H6 |
-1.654 |
-0.549 |
1.222 |
| H7 |
-1.124 |
-1.477 |
-0.196 |
| H8 |
-0.551 |
0.675 |
-1.389 |
| H9 |
-1.072 |
1.599 |
0.026 |
| H10 |
0.709 |
0.511 |
1.396 |
| H11 |
1.351 |
1.439 |
0.010 |
| H12 |
2.267 |
-0.716 |
0.169 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5304 | 2.5472 | 2.9509 | 1.0976 | 1.0993 | 1.0960 | 2.1697 | 2.1707 | 2.7803 | 3.4943 | 3.8139 |
C2 | 1.5304 | | 1.5209 | 2.3940 | 2.1792 | 2.1791 | 2.1758 | 1.0995 | 1.1003 | 2.1626 | 2.1619 | 3.2368 | C3 | 2.5472 | 1.5209 | | 1.4355 | 3.4973 | 2.8137 | 2.8143 | 2.1399 | 2.1322 | 1.1032 | 1.1017 | 1.9691 | O4 | 2.9509 | 2.3940 | 1.4355 | | 3.9371 | 3.3637 | 2.6469 | 2.6184 | 3.3334 | 2.0911 | 2.0901 | 0.9631 | H5 | 1.0976 | 2.1792 | 3.4973 | 3.9371 | | 1.7754 | 1.7774 | 2.5136 | 2.5141 | 3.7860 | 4.3234 | 4.8428 | H6 | 1.0993 | 2.1791 | 2.8137 | 3.3637 | 1.7754 | | 1.7762 | 3.0871 | 2.5265 | 2.5955 | 3.8009 | 4.0629 | H7 | 1.0960 | 2.1758 | 2.8143 | 2.6469 | 1.7774 | 1.7762 | | 2.5259 | 3.0847 | 3.1382 | 3.8304 | 3.4942 | H8 | 2.1697 | 1.0995 | 2.1399 | 2.6184 | 2.5136 | 3.0871 | 2.5259 | | 1.7684 | 3.0606 | 2.4815 | 3.5068 | H9 | 2.1707 | 1.1003 | 2.1322 | 3.3334 | 2.5141 | 2.5265 | 3.0847 | 1.7684 | | 2.4972 | 2.4286 | 4.0659 | H10 | 2.7803 | 2.1626 | 1.1032 | 2.0911 | 3.7860 | 2.5955 | 3.1382 | 3.0606 | 2.4972 | | 1.7867 | 2.3317 | H11 | 3.4943 | 2.1619 | 1.1017 | 2.0901 | 4.3234 | 3.8009 | 3.8304 | 2.4815 | 2.4286 | 1.7867 | | 2.3472 | H12 | 3.8139 | 3.2368 | 1.9691 | 0.9631 | 4.8428 | 4.0629 | 3.4942 | 3.5068 | 4.0659 | 2.3317 | 2.3472 | |
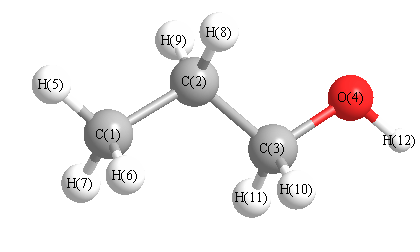 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
113.186 |
|
C1 |
C2 |
H8 |
110.099 |
| C1 |
C2 |
H9 |
110.124 |
|
C2 |
C1 |
H5 |
110.968 |
| C2 |
C1 |
H6 |
110.855 |
|
C2 |
C1 |
H7 |
110.793 |
| C2 |
C3 |
O4 |
108.114 |
|
C2 |
C3 |
H10 |
109.980 |
| C2 |
C3 |
H11 |
110.019 |
|
C3 |
C2 |
H8 |
108.423 |
| C3 |
C2 |
H9 |
107.782 |
|
C3 |
O4 |
H12 |
108.765 |
| O4 |
C3 |
H10 |
110.228 |
|
O4 |
C3 |
H11 |
110.243 |
| H5 |
C1 |
H6 |
107.838 |
|
H5 |
C1 |
H7 |
108.249 |
| H6 |
C1 |
H7 |
108.018 |
|
H8 |
C2 |
H9 |
107.007 |
| H10 |
C3 |
H11 |
108.257 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP=FULLultrafine/daug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.937 |
|
|
|
| 2 |
C |
0.314 |
|
|
|
| 3 |
C |
2.256 |
|
|
|
| 4 |
O |
-0.799 |
|
|
|
| 5 |
H |
-0.365 |
|
|
|
| 6 |
H |
-0.282 |
|
|
|
| 7 |
H |
-0.186 |
|
|
|
| 8 |
H |
-0.469 |
|
|
|
| 9 |
H |
-0.657 |
|
|
|
| 10 |
H |
-0.540 |
|
|
|
| 11 |
H |
-0.638 |
|
|
|
| 12 |
H |
0.427 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.603 |
1.017 |
0.949 |
1.516 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.685 |
-0.072 |
1.964 |
| y |
-0.072 |
-27.914 |
-0.518 |
| z |
1.964 |
-0.518 |
-27.473 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.008 |
-0.072 |
1.964 |
| y |
-0.072 |
-2.334 |
-0.518 |
| z |
1.964 |
-0.518 |
-1.674 |
|
| Polar |
|---|
| 3z2-r2 | -3.347 |
|---|
| x2-y2 | 4.228 |
|---|
| xy | -0.072 |
|---|
| xz | 1.964 |
|---|
| yz | -0.518 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.420 |
0.075 |
0.061 |
| y |
0.075 |
6.746 |
-0.004 |
| z |
0.061 |
-0.004 |
6.319 |
<r2> (average value of r
2) Å
2
| <r2> |
96.122 |
| (<r2>)1/2 |
9.804 |
