Jump to
S1C2
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -477.794553 |
| Energy at 298.15K | |
| HF Energy | -477.639615 |
| Nuclear repulsion energy | 106.771138 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3135 |
3135 |
23.46 |
|
|
|
| 2 |
A' |
3083 |
3083 |
21.34 |
|
|
|
| 3 |
A' |
3055 |
3055 |
20.84 |
|
|
|
| 4 |
A' |
2702 |
2702 |
3.78 |
|
|
|
| 5 |
A' |
1490 |
1490 |
2.20 |
|
|
|
| 6 |
A' |
1474 |
1474 |
2.46 |
|
|
|
| 7 |
A' |
1400 |
1400 |
3.63 |
|
|
|
| 8 |
A' |
1289 |
1289 |
32.42 |
|
|
|
| 9 |
A' |
1111 |
1111 |
0.98 |
|
|
|
| 10 |
A' |
999 |
999 |
3.83 |
|
|
|
| 11 |
A' |
855 |
855 |
0.87 |
|
|
|
| 12 |
A' |
670 |
670 |
1.97 |
|
|
|
| 13 |
A' |
305 |
305 |
2.11 |
|
|
|
| 14 |
A" |
3150 |
3150 |
24.57 |
|
|
|
| 15 |
A" |
3128 |
3128 |
1.44 |
|
|
|
| 16 |
A" |
1480 |
1480 |
8.59 |
|
|
|
| 17 |
A" |
1263 |
1263 |
0.36 |
|
|
|
| 18 |
A" |
1041 |
1041 |
0.46 |
|
|
|
| 19 |
A" |
786 |
786 |
2.86 |
|
|
|
| 20 |
A" |
253 |
253 |
0.83 |
|
|
|
| 21 |
A" |
170 |
170 |
13.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16418.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16418.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
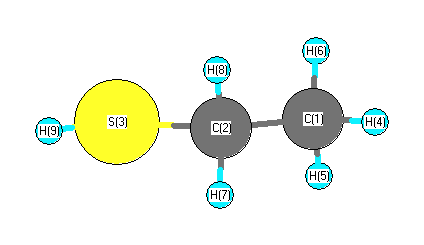
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.520 |
0.699 |
0.000 |
| C2 |
0.000 |
0.838 |
0.000 |
| S3 |
-0.757 |
-0.846 |
0.000 |
| H4 |
1.987 |
1.692 |
0.000 |
| H5 |
1.867 |
0.157 |
0.889 |
| H6 |
1.867 |
0.157 |
-0.889 |
| H7 |
-0.337 |
1.379 |
0.892 |
| H8 |
-0.337 |
1.379 |
-0.892 |
| H9 |
-2.050 |
-0.449 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5265 | 2.7520 | 1.0979 | 1.0974 | 1.0974 | 2.1697 | 2.1697 | 3.7501 |
C2 | 1.5265 | | 1.8467 | 2.1630 | 2.1770 | 2.1770 | 1.0959 | 1.0959 | 2.4206 | S3 | 2.7520 | 1.8467 | | 3.7384 | 2.9469 | 2.9469 | 2.4333 | 2.4333 | 1.3521 | H4 | 1.0979 | 2.1630 | 3.7384 | | 1.7781 | 1.7781 | 2.5093 | 2.5093 | 4.5699 | H5 | 1.0974 | 2.1770 | 2.9469 | 1.7781 | | 1.7790 | 2.5197 | 3.0857 | 4.0619 | H6 | 1.0974 | 2.1770 | 2.9469 | 1.7781 | 1.7790 | | 3.0857 | 2.5197 | 4.0619 | H7 | 2.1697 | 1.0959 | 2.4333 | 2.5093 | 2.5197 | 3.0857 | | 1.7836 | 2.6588 | H8 | 2.1697 | 1.0959 | 2.4333 | 2.5093 | 3.0857 | 2.5197 | 1.7836 | | 2.6588 | H9 | 3.7501 | 2.4206 | 1.3521 | 4.5699 | 4.0619 | 4.0619 | 2.6588 | 2.6588 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.972 |
|
C1 |
C2 |
H7 |
110.578 |
| C1 |
C2 |
H8 |
110.578 |
|
C2 |
C1 |
H4 |
109.934 |
| C2 |
C1 |
H5 |
111.071 |
|
C2 |
C1 |
H6 |
111.071 |
| C2 |
S3 |
H9 |
97.138 |
|
S3 |
C2 |
H7 |
108.877 |
| S3 |
C2 |
H8 |
108.877 |
|
H4 |
C1 |
H5 |
108.186 |
| H4 |
C1 |
H6 |
108.186 |
|
H5 |
C1 |
H6 |
108.293 |
| H7 |
C2 |
H8 |
108.922 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -477.795329 |
| Energy at 298.15K | |
| HF Energy | -477.640296 |
| Nuclear repulsion energy | 106.513686 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3151 |
3151 |
26.56 |
10.52 |
0.75 |
0.85 |
| 2 |
A |
3130 |
3130 |
5.24 |
85.08 |
0.75 |
0.86 |
| 3 |
A |
3121 |
3121 |
25.50 |
94.26 |
0.66 |
0.80 |
| 4 |
A |
3083 |
3083 |
14.15 |
130.26 |
0.09 |
0.16 |
| 5 |
A |
3047 |
3047 |
26.44 |
197.40 |
0.03 |
0.07 |
| 6 |
A |
2694 |
2694 |
3.63 |
120.75 |
0.19 |
0.33 |
| 7 |
A |
1484 |
1484 |
2.26 |
2.72 |
0.75 |
0.86 |
| 8 |
A |
1478 |
1478 |
9.37 |
5.21 |
0.74 |
0.85 |
| 9 |
A |
1460 |
1460 |
0.94 |
7.58 |
0.70 |
0.82 |
| 10 |
A |
1398 |
1398 |
3.97 |
0.56 |
0.74 |
0.85 |
| 11 |
A |
1297 |
1297 |
20.49 |
2.70 |
0.30 |
0.46 |
| 12 |
A |
1275 |
1275 |
3.51 |
1.61 |
0.73 |
0.85 |
| 13 |
A |
1117 |
1117 |
5.49 |
3.74 |
0.27 |
0.42 |
| 14 |
A |
1069 |
1069 |
0.57 |
4.43 |
0.19 |
0.32 |
| 15 |
A |
990 |
990 |
6.75 |
3.85 |
0.74 |
0.85 |
| 16 |
A |
867 |
867 |
5.03 |
1.74 |
0.26 |
0.42 |
| 17 |
A |
732 |
732 |
1.37 |
2.87 |
0.20 |
0.34 |
| 18 |
A |
657 |
657 |
4.11 |
15.51 |
0.19 |
0.32 |
| 19 |
A |
328 |
328 |
0.98 |
1.55 |
0.20 |
0.33 |
| 20 |
A |
256 |
256 |
1.01 |
0.12 |
0.34 |
0.50 |
| 21 |
A |
203 |
203 |
13.20 |
0.62 |
0.52 |
0.69 |
Unscaled Zero Point Vibrational Energy (zpe) 16418.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16418.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
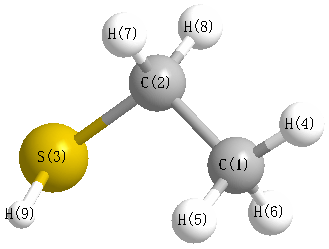
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.646 |
-0.354 |
-0.053 |
| C2 |
0.505 |
0.647 |
0.091 |
| S3 |
-1.171 |
-0.100 |
-0.082 |
| H4 |
2.614 |
0.162 |
0.017 |
| H5 |
1.607 |
-1.111 |
0.741 |
| H6 |
1.594 |
-0.872 |
-1.019 |
| H7 |
0.557 |
1.178 |
1.048 |
| H8 |
0.545 |
1.401 |
-0.706 |
| H9 |
-1.093 |
-0.911 |
0.998 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5250 | 2.8281 | 1.0996 | 1.0978 | 1.0965 | 2.1789 | 2.1727 | 2.9859 |
C2 | 1.5250 | | 1.8428 | 2.1658 | 2.1746 | 2.1731 | 1.0962 | 1.0974 | 2.4094 | S3 | 2.8281 | 1.8428 | | 3.7953 | 3.0686 | 3.0189 | 2.4282 | 2.3633 | 1.3529 | H4 | 1.0996 | 2.1658 | 3.7953 | | 1.7773 | 1.7836 | 2.5160 | 2.5184 | 3.9822 | H5 | 1.0978 | 2.1746 | 3.0686 | 1.7773 | | 1.7763 | 2.5370 | 3.0877 | 2.7195 | H6 | 1.0965 | 2.1731 | 3.0189 | 1.7836 | 1.7763 | | 3.0900 | 2.5228 | 3.3593 | H7 | 2.1789 | 1.0962 | 2.4282 | 2.5160 | 2.5370 | 3.0900 | | 1.7682 | 2.6628 | H8 | 2.1727 | 1.0974 | 2.3633 | 2.5184 | 3.0877 | 2.5228 | 1.7682 | | 3.3063 | H9 | 2.9859 | 2.4094 | 1.3529 | 3.9822 | 2.7195 | 3.3593 | 2.6628 | 3.3063 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
113.900 |
|
C1 |
C2 |
H7 |
111.405 |
| C1 |
C2 |
H8 |
110.841 |
|
C2 |
C1 |
H4 |
110.168 |
| C2 |
C1 |
H5 |
110.972 |
|
C2 |
C1 |
H6 |
110.930 |
| C2 |
S3 |
H9 |
96.668 |
|
S3 |
C2 |
H7 |
108.748 |
| S3 |
C2 |
H8 |
104.096 |
|
H4 |
C1 |
H5 |
107.964 |
| H4 |
C1 |
H6 |
108.618 |
|
H5 |
C1 |
H6 |
108.093 |
| H7 |
C2 |
H8 |
107.421 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
