Jump to
S1C2
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -216.967753 |
| Energy at 298.15K | |
| HF Energy | -216.757305 |
| Nuclear repulsion energy | 115.713218 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3280 |
3280 |
4.37 |
51.75 |
0.72 |
0.83 |
| 2 |
A' |
3186 |
3186 |
5.23 |
186.61 |
0.11 |
0.20 |
| 3 |
A' |
3174 |
3174 |
8.78 |
25.80 |
0.74 |
0.85 |
| 4 |
A' |
3055 |
3055 |
33.74 |
187.18 |
0.09 |
0.16 |
| 5 |
A' |
1710 |
1710 |
5.07 |
35.54 |
0.04 |
0.07 |
| 6 |
A' |
1483 |
1483 |
2.85 |
12.34 |
0.53 |
0.70 |
| 7 |
A' |
1447 |
1447 |
6.73 |
7.70 |
0.36 |
0.53 |
| 8 |
A' |
1399 |
1399 |
10.75 |
2.58 |
0.65 |
0.78 |
| 9 |
A' |
1311 |
1311 |
0.19 |
20.46 |
0.21 |
0.35 |
| 10 |
A' |
1117 |
1117 |
47.61 |
1.44 |
0.68 |
0.81 |
| 11 |
A' |
996 |
996 |
43.83 |
4.76 |
0.64 |
0.78 |
| 12 |
A' |
912 |
912 |
5.87 |
7.28 |
0.04 |
0.08 |
| 13 |
A' |
602 |
602 |
7.61 |
1.61 |
0.69 |
0.82 |
| 14 |
A' |
260 |
260 |
2.48 |
1.29 |
0.42 |
0.59 |
| 15 |
A" |
3102 |
3102 |
18.91 |
82.96 |
0.75 |
0.86 |
| 16 |
A" |
1252 |
1252 |
0.01 |
3.63 |
0.75 |
0.86 |
| 17 |
A" |
1041 |
1041 |
12.35 |
0.92 |
0.75 |
0.86 |
| 18 |
A" |
1013 |
1013 |
6.43 |
0.17 |
0.75 |
0.86 |
| 19 |
A" |
974 |
974 |
45.76 |
2.11 |
0.75 |
0.86 |
| 20 |
A" |
563 |
563 |
11.15 |
1.25 |
0.75 |
0.86 |
| 21 |
A" |
169 |
169 |
3.24 |
0.92 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16022.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16022.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.951 |
-0.200 |
0.000 |
| C2 |
0.000 |
0.956 |
0.000 |
| C3 |
1.335 |
0.853 |
0.000 |
| F4 |
-0.270 |
-1.427 |
0.000 |
| H5 |
1.962 |
1.743 |
0.000 |
| H6 |
1.827 |
-0.117 |
0.000 |
| H7 |
-0.481 |
1.937 |
0.000 |
| H8 |
-1.593 |
-0.186 |
0.893 |
| H9 |
-1.593 |
-0.186 |
-0.893 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4967 | 2.5166 | 1.4027 | 3.5011 | 2.7787 | 2.1883 | 1.0998 | 1.0998 |
C2 | 1.4967 | | 1.3387 | 2.3977 | 2.1138 | 2.1186 | 1.0929 | 2.1536 | 2.1536 | C3 | 2.5166 | 1.3387 | | 2.7881 | 1.0881 | 1.0882 | 2.1146 | 3.2325 | 3.2325 | F4 | 1.4027 | 2.3977 | 2.7881 | | 3.8762 | 2.4719 | 3.3704 | 2.0216 | 2.0216 | H5 | 3.5011 | 2.1138 | 1.0881 | 3.8762 | | 1.8649 | 2.4505 | 4.1416 | 4.1416 | H6 | 2.7787 | 2.1186 | 1.0882 | 2.4719 | 1.8649 | | 3.0897 | 3.5351 | 3.5351 | H7 | 2.1883 | 1.0929 | 2.1146 | 3.3704 | 2.4505 | 3.0897 | | 2.5575 | 2.5575 | H8 | 1.0998 | 2.1536 | 3.2325 | 2.0216 | 4.1416 | 3.5351 | 2.5575 | | 1.7853 | H9 | 1.0998 | 2.1536 | 3.2325 | 2.0216 | 4.1416 | 3.5351 | 2.5575 | 1.7853 | |
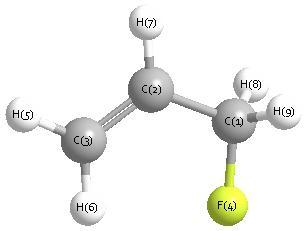 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.046 |
|
C1 |
C2 |
H7 |
114.456 |
| C2 |
C1 |
F4 |
111.529 |
|
C2 |
C1 |
H8 |
111.158 |
| C2 |
C1 |
H9 |
111.158 |
|
C2 |
C3 |
H5 |
120.797 |
| C2 |
C3 |
H6 |
121.266 |
|
C3 |
C2 |
H7 |
120.499 |
| F4 |
C1 |
H8 |
107.147 |
|
F4 |
C1 |
H9 |
107.147 |
| H5 |
C3 |
H6 |
117.936 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -216.968017 |
| Energy at 298.15K | |
| HF Energy | -216.757294 |
| Nuclear repulsion energy | 113.495121 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3263 |
3263 |
9.96 |
60.87 |
0.60 |
0.75 |
| 2 |
A |
3194 |
3194 |
6.30 |
107.86 |
0.22 |
0.36 |
| 3 |
A |
3164 |
3164 |
7.48 |
93.71 |
0.09 |
0.17 |
| 4 |
A |
3135 |
3135 |
16.99 |
63.77 |
0.74 |
0.85 |
| 5 |
A |
3073 |
3073 |
30.31 |
137.82 |
0.07 |
0.12 |
| 6 |
A |
1705 |
1705 |
0.32 |
37.57 |
0.06 |
0.11 |
| 7 |
A |
1492 |
1492 |
5.26 |
3.19 |
0.62 |
0.76 |
| 8 |
A |
1457 |
1457 |
13.27 |
10.45 |
0.48 |
0.65 |
| 9 |
A |
1371 |
1371 |
13.33 |
4.09 |
0.22 |
0.36 |
| 10 |
A |
1316 |
1316 |
0.09 |
15.10 |
0.29 |
0.44 |
| 11 |
A |
1254 |
1254 |
4.67 |
7.19 |
0.67 |
0.81 |
| 12 |
A |
1181 |
1181 |
2.96 |
1.20 |
0.39 |
0.56 |
| 13 |
A |
1029 |
1029 |
10.01 |
0.75 |
0.70 |
0.82 |
| 14 |
A |
1007 |
1007 |
126.34 |
6.84 |
0.66 |
0.80 |
| 15 |
A |
987 |
987 |
28.52 |
2.07 |
0.53 |
0.69 |
| 16 |
A |
982 |
982 |
38.20 |
3.43 |
0.04 |
0.08 |
| 17 |
A |
925 |
925 |
6.99 |
4.40 |
0.13 |
0.23 |
| 18 |
A |
660 |
660 |
7.15 |
1.77 |
0.23 |
0.38 |
| 19 |
A |
430 |
430 |
2.09 |
3.59 |
0.30 |
0.46 |
| 20 |
A |
336 |
336 |
8.31 |
1.22 |
0.74 |
0.85 |
| 21 |
A |
116 |
116 |
1.08 |
2.55 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16039.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16039.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.607 |
0.428 |
0.345 |
| C2 |
0.648 |
-0.380 |
0.265 |
| C3 |
1.799 |
0.096 |
-0.231 |
| F4 |
-1.640 |
-0.224 |
-0.370 |
| H5 |
2.703 |
-0.509 |
-0.251 |
| H6 |
1.870 |
1.110 |
-0.626 |
| H7 |
0.591 |
-1.400 |
0.650 |
| H8 |
-0.965 |
0.526 |
1.378 |
| H9 |
-0.478 |
1.422 |
-0.100 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4940 | 2.4954 | 1.4158 | 3.4912 | 2.7461 | 2.2061 | 1.0983 | 1.0970 |
C2 | 1.4940 | | 1.3404 | 2.3799 | 2.1231 | 2.1229 | 1.0915 | 2.1593 | 2.1559 | C3 | 2.4954 | 1.3404 | | 3.4569 | 1.0886 | 1.0906 | 2.1144 | 3.2271 | 2.6381 | F4 | 1.4158 | 2.3799 | 3.4569 | | 4.3547 | 3.7637 | 2.7208 | 2.0185 | 2.0332 | H5 | 3.4912 | 2.1231 | 1.0886 | 4.3547 | | 1.8594 | 2.4627 | 4.1455 | 3.7248 | H6 | 2.7461 | 2.1229 | 1.0906 | 3.7637 | 1.8594 | | 3.0921 | 3.5211 | 2.4263 | H7 | 2.2061 | 1.0915 | 2.1144 | 2.7208 | 2.4627 | 3.0921 | | 2.5810 | 3.1093 | H8 | 1.0983 | 2.1593 | 3.2271 | 2.0185 | 4.1455 | 3.5211 | 2.5810 | | 1.7959 | H9 | 1.0970 | 2.1559 | 2.6381 | 2.0332 | 3.7248 | 2.4263 | 3.1093 | 1.7959 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.290 |
|
C1 |
C2 |
H7 |
116.260 |
| C2 |
C1 |
F4 |
109.721 |
|
C2 |
C1 |
H8 |
111.903 |
| C2 |
C1 |
H9 |
111.706 |
|
C2 |
C3 |
H5 |
121.525 |
| C2 |
C3 |
H6 |
121.336 |
|
C3 |
C2 |
H7 |
120.446 |
| F4 |
C1 |
H8 |
106.126 |
|
F4 |
C1 |
H9 |
107.344 |
| H5 |
C3 |
H6 |
117.138 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP=FULLultrafine/daug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
1.436 |
|
|
|
| 2 |
C |
1.658 |
|
|
|
| 3 |
C |
1.118 |
|
|
|
| 4 |
F |
-0.470 |
|
|
|
| 5 |
H |
-0.926 |
|
|
|
| 6 |
H |
-0.814 |
|
|
|
| 7 |
H |
-0.933 |
|
|
|
| 8 |
H |
-0.327 |
|
|
|
| 9 |
H |
-0.741 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.635 |
0.774 |
0.942 |
2.040 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.375 |
-1.224 |
-2.203 |
| y |
-1.224 |
-23.024 |
-1.058 |
| z |
-2.203 |
-1.058 |
-25.284 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.221 |
-1.224 |
-2.203 |
| y |
-1.224 |
2.805 |
-1.058 |
| z |
-2.203 |
-1.058 |
-0.584 |
|
| Polar |
|---|
| 3z2-r2 | -1.169 |
|---|
| x2-y2 | -3.351 |
|---|
| xy | -1.224 |
|---|
| xz | -2.203 |
|---|
| yz | -1.058 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.607 |
0.241 |
-0.619 |
| y |
0.241 |
5.598 |
-0.221 |
| z |
-0.619 |
-0.221 |
5.202 |
<r2> (average value of r
2) Å
2
| <r2> |
89.913 |
| (<r2>)1/2 |
9.482 |
