Jump to
S1C2
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -343.086466 |
| Energy at 298.15K | |
| HF Energy | -342.725134 |
| Nuclear repulsion energy | 270.342355 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3306 |
3306 |
0.09 |
158.53 |
0.12 |
0.21 |
| 2 |
A' |
3285 |
3285 |
0.33 |
64.89 |
0.30 |
0.46 |
| 3 |
A' |
3273 |
3273 |
1.02 |
78.58 |
0.69 |
0.81 |
| 4 |
A' |
2948 |
2948 |
90.85 |
148.95 |
0.30 |
0.46 |
| 5 |
A' |
1727 |
1727 |
356.76 |
200.31 |
0.31 |
0.47 |
| 6 |
A' |
1594 |
1594 |
9.62 |
3.61 |
0.54 |
0.70 |
| 7 |
A' |
1506 |
1506 |
42.90 |
129.03 |
0.24 |
0.38 |
| 8 |
A' |
1426 |
1426 |
34.64 |
43.98 |
0.43 |
0.60 |
| 9 |
A' |
1383 |
1383 |
0.12 |
31.16 |
0.11 |
0.20 |
| 10 |
A' |
1304 |
1304 |
42.29 |
7.54 |
0.52 |
0.68 |
| 11 |
A' |
1239 |
1239 |
1.75 |
11.31 |
0.25 |
0.40 |
| 12 |
A' |
1182 |
1182 |
9.34 |
6.19 |
0.09 |
0.17 |
| 13 |
A' |
1101 |
1101 |
7.13 |
14.34 |
0.23 |
0.37 |
| 14 |
A' |
1038 |
1038 |
42.48 |
5.42 |
0.39 |
0.56 |
| 15 |
A' |
939 |
939 |
17.56 |
8.77 |
0.08 |
0.14 |
| 16 |
A' |
893 |
893 |
10.34 |
6.06 |
0.73 |
0.84 |
| 17 |
A' |
761 |
761 |
62.83 |
1.60 |
0.67 |
0.80 |
| 18 |
A' |
498 |
498 |
0.99 |
9.03 |
0.20 |
0.33 |
| 19 |
A' |
202 |
202 |
5.84 |
0.94 |
0.65 |
0.79 |
| 20 |
A" |
999 |
999 |
0.12 |
1.11 |
0.75 |
0.86 |
| 21 |
A" |
891 |
891 |
1.02 |
0.48 |
0.75 |
0.86 |
| 22 |
A" |
836 |
836 |
4.39 |
1.12 |
0.75 |
0.86 |
| 23 |
A" |
769 |
769 |
68.51 |
1.45 |
0.75 |
0.86 |
| 24 |
A" |
651 |
651 |
0.40 |
0.82 |
0.75 |
0.86 |
| 25 |
A" |
605 |
605 |
7.52 |
0.17 |
0.75 |
0.86 |
| 26 |
A" |
294 |
294 |
13.72 |
0.37 |
0.75 |
0.86 |
| 27 |
A" |
134 |
134 |
1.42 |
1.21 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17391.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17391.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.520 |
-0.899 |
0.000 |
| C2 |
-0.532 |
-1.757 |
0.000 |
| C3 |
-1.725 |
-1.075 |
0.000 |
| C4 |
0.000 |
0.369 |
0.000 |
| C5 |
-1.380 |
0.310 |
0.000 |
| C6 |
0.899 |
1.516 |
0.000 |
| O7 |
2.120 |
1.460 |
0.000 |
| H8 |
-0.283 |
-2.810 |
0.000 |
| H9 |
-2.716 |
-1.514 |
0.000 |
| H10 |
-2.051 |
1.162 |
0.000 |
| H11 |
0.361 |
2.490 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3574 | 2.2516 | 1.3701 | 2.2512 | 2.4449 | 2.8502 | 2.0731 | 3.2934 | 3.2944 | 3.3924 |
C2 | 1.3574 | | 1.3739 | 2.1914 | 2.2338 | 3.5725 | 4.1690 | 1.0828 | 2.1970 | 3.2901 | 4.3396 | C3 | 2.2516 | 1.3739 | | 2.2496 | 1.4275 | 3.6881 | 4.6052 | 2.2567 | 1.0836 | 2.2605 | 4.1304 | C4 | 1.3701 | 2.1914 | 2.2496 | | 1.3808 | 1.4578 | 2.3840 | 3.1920 | 3.3046 | 2.1987 | 2.1514 | C5 | 2.2512 | 2.2338 | 1.4275 | 1.3808 | | 2.5784 | 3.6833 | 3.3077 | 2.2610 | 1.0846 | 2.7896 | C6 | 2.4449 | 3.5725 | 3.6881 | 1.4578 | 2.5784 | | 1.2218 | 4.4853 | 4.7171 | 2.9712 | 1.1123 | O7 | 2.8502 | 4.1690 | 4.6052 | 2.3840 | 3.6833 | 1.2218 | | 4.8997 | 5.6768 | 4.1811 | 2.0380 | H8 | 2.0731 | 1.0828 | 2.2567 | 3.1920 | 3.3077 | 4.4853 | 4.8997 | | 2.7572 | 4.3481 | 5.3393 | H9 | 3.2934 | 2.1970 | 1.0836 | 3.3046 | 2.2610 | 4.7171 | 5.6768 | 2.7572 | | 2.7572 | 5.0495 | H10 | 3.2944 | 3.2901 | 2.2605 | 2.1987 | 1.0846 | 2.9712 | 4.1811 | 4.3481 | 2.7572 | | 2.7533 | H11 | 3.3924 | 4.3396 | 4.1304 | 2.1514 | 2.7896 | 1.1123 | 2.0380 | 5.3393 | 5.0495 | 2.7533 | |
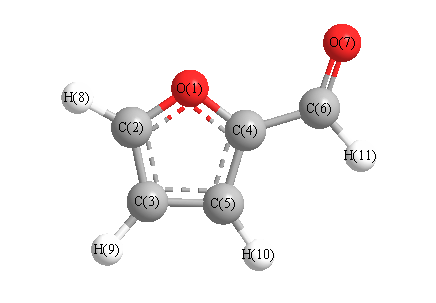 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.044 |
|
O1 |
C2 |
H8 |
115.870 |
| O1 |
C4 |
C5 |
109.841 |
|
O1 |
C4 |
C6 |
119.626 |
| C2 |
O1 |
C4 |
106.915 |
|
C2 |
C3 |
C5 |
105.744 |
| C2 |
C3 |
H9 |
126.358 |
|
C3 |
C2 |
H8 |
133.086 |
| C3 |
C5 |
C4 |
106.455 |
|
C3 |
C5 |
H10 |
127.756 |
| C4 |
C5 |
H10 |
125.789 |
|
C4 |
C6 |
O7 |
125.434 |
| C4 |
C6 |
H11 |
112.983 |
|
C5 |
C3 |
H9 |
127.898 |
| C5 |
C4 |
C6 |
130.532 |
|
O7 |
C6 |
H11 |
121.583 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -343.087294 |
| Energy at 298.15K | |
| HF Energy | -342.725862 |
| Nuclear repulsion energy | 269.500235 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3307 |
3307 |
0.04 |
161.88 |
0.13 |
0.22 |
| 2 |
A' |
3292 |
3292 |
1.33 |
43.15 |
0.23 |
0.37 |
| 3 |
A' |
3277 |
3277 |
0.63 |
82.73 |
0.70 |
0.82 |
| 4 |
A' |
2970 |
2970 |
81.26 |
155.27 |
0.30 |
0.46 |
| 5 |
A' |
1718 |
1718 |
280.73 |
153.70 |
0.33 |
0.50 |
| 6 |
A' |
1605 |
1605 |
76.92 |
31.04 |
0.35 |
0.51 |
| 7 |
A' |
1499 |
1499 |
103.20 |
235.65 |
0.25 |
0.39 |
| 8 |
A' |
1434 |
1434 |
1.08 |
5.61 |
0.51 |
0.67 |
| 9 |
A' |
1391 |
1391 |
12.76 |
37.85 |
0.36 |
0.53 |
| 10 |
A' |
1265 |
1265 |
25.93 |
5.29 |
0.29 |
0.44 |
| 11 |
A' |
1221 |
1221 |
2.80 |
3.85 |
0.47 |
0.64 |
| 12 |
A' |
1172 |
1172 |
10.91 |
12.69 |
0.09 |
0.17 |
| 13 |
A' |
1105 |
1105 |
22.55 |
16.41 |
0.27 |
0.43 |
| 14 |
A' |
1032 |
1032 |
38.99 |
3.81 |
0.35 |
0.51 |
| 15 |
A' |
953 |
953 |
8.32 |
10.19 |
0.10 |
0.19 |
| 16 |
A' |
893 |
893 |
6.50 |
4.91 |
0.73 |
0.84 |
| 17 |
A' |
753 |
753 |
72.71 |
2.98 |
0.56 |
0.72 |
| 18 |
A' |
493 |
493 |
0.91 |
6.65 |
0.23 |
0.37 |
| 19 |
A' |
204 |
204 |
6.93 |
0.37 |
0.25 |
0.40 |
| 20 |
A" |
1007 |
1007 |
0.09 |
1.85 |
0.75 |
0.86 |
| 21 |
A" |
898 |
898 |
1.20 |
0.59 |
0.75 |
0.86 |
| 22 |
A" |
838 |
838 |
8.22 |
0.56 |
0.75 |
0.86 |
| 23 |
A" |
765 |
765 |
62.22 |
1.99 |
0.75 |
0.86 |
| 24 |
A" |
638 |
638 |
0.71 |
0.38 |
0.75 |
0.86 |
| 25 |
A" |
605 |
605 |
9.57 |
0.23 |
0.75 |
0.86 |
| 26 |
A" |
247 |
247 |
14.11 |
1.63 |
0.75 |
0.86 |
| 27 |
A" |
154 |
154 |
2.92 |
0.91 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17367.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 17367.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP=FULLultrafine/daug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.248 |
-0.295 |
0.000 |
| C2 |
1.054 |
-1.638 |
0.000 |
| C3 |
-0.287 |
-1.938 |
0.000 |
| C4 |
0.000 |
0.290 |
0.000 |
| C5 |
-0.973 |
-0.687 |
0.000 |
| C6 |
-0.041 |
1.748 |
0.000 |
| O7 |
-1.080 |
2.395 |
0.000 |
| H8 |
1.950 |
-2.247 |
0.000 |
| H9 |
-0.721 |
-2.931 |
0.000 |
| H10 |
-2.041 |
-0.506 |
0.000 |
| H11 |
0.954 |
2.241 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3572 | 2.2485 | 1.3779 | 2.2550 | 2.4153 | 3.5571 | 2.0747 | 3.2902 | 3.2958 | 2.5530 |
C2 | 1.3572 | | 1.3748 | 2.1975 | 2.2394 | 3.5588 | 4.5631 | 1.0827 | 2.1961 | 3.2965 | 3.8807 | C3 | 2.2485 | 1.3748 | | 2.2460 | 1.4263 | 3.6937 | 4.4046 | 2.2583 | 1.0838 | 2.2644 | 4.3593 | C4 | 1.3779 | 2.1975 | 2.2460 | | 1.3787 | 1.4586 | 2.3660 | 3.1995 | 3.3006 | 2.1909 | 2.1720 | C5 | 2.2550 | 2.2394 | 1.4263 | 1.3787 | | 2.6069 | 3.0838 | 3.3129 | 2.2582 | 1.0837 | 3.5051 | C6 | 2.4153 | 3.5588 | 3.6937 | 1.4586 | 2.6069 | | 1.2238 | 4.4635 | 4.7280 | 3.0129 | 1.1106 | O7 | 3.5571 | 4.5631 | 4.4046 | 2.3660 | 3.0838 | 1.2238 | | 5.5432 | 5.3382 | 3.0557 | 2.0394 | H8 | 2.0747 | 1.0827 | 2.2583 | 3.1995 | 3.3129 | 4.4635 | 5.5432 | | 2.7566 | 4.3544 | 4.5975 | H9 | 3.2902 | 2.1961 | 1.0838 | 3.3006 | 2.2582 | 4.7280 | 5.3382 | 2.7566 | | 2.7617 | 5.4366 | H10 | 3.2958 | 3.2965 | 2.2644 | 2.1909 | 1.0837 | 3.0129 | 3.0557 | 4.3544 | 2.7617 | | 4.0639 | H11 | 2.5530 | 3.8807 | 4.3593 | 2.1720 | 3.5051 | 1.1106 | 2.0394 | 4.5975 | 5.4366 | 4.0639 | |
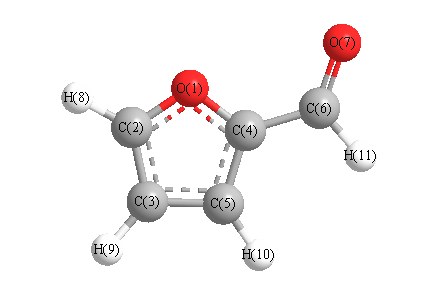 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
110.772 |
|
O1 |
C2 |
H8 |
116.039 |
| O1 |
C4 |
C5 |
109.783 |
|
O1 |
C4 |
C6 |
116.728 |
| C2 |
O1 |
C4 |
106.917 |
|
C2 |
C3 |
C5 |
106.145 |
| C2 |
C3 |
H9 |
126.155 |
|
C3 |
C2 |
H8 |
133.190 |
| C3 |
C5 |
C4 |
106.384 |
|
C3 |
C5 |
H10 |
128.365 |
| C4 |
C5 |
H10 |
125.251 |
|
C4 |
C6 |
O7 |
123.551 |
| C4 |
C6 |
H11 |
114.753 |
|
C5 |
C3 |
H9 |
127.700 |
| C5 |
C4 |
C6 |
133.489 |
|
O7 |
C6 |
H11 |
121.696 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
