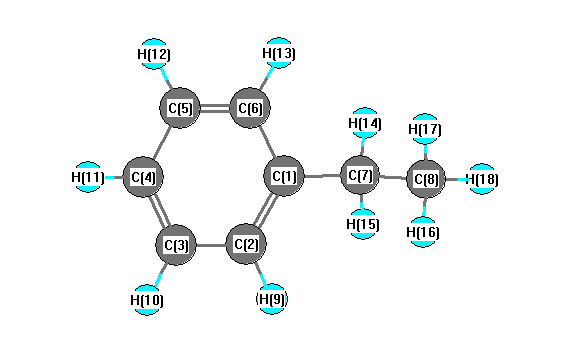
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3346 |
3033 |
18.14 |
|
|
|
| 2 |
A' |
3323 |
3012 |
9.95 |
|
|
|
| 3 |
A' |
3306 |
2997 |
9.64 |
|
|
|
| 4 |
A' |
3218 |
2917 |
66.66 |
|
|
|
| 5 |
A' |
3171 |
2874 |
27.35 |
|
|
|
| 6 |
A' |
3158 |
2863 |
42.10 |
|
|
|
| 7 |
A' |
1788 |
1621 |
8.79 |
|
|
|
| 8 |
A' |
1655 |
1500 |
14.59 |
|
|
|
| 9 |
A' |
1627 |
1475 |
2.86 |
|
|
|
| 10 |
A' |
1608 |
1457 |
0.45 |
|
|
|
| 11 |
A' |
1533 |
1389 |
1.78 |
|
|
|
| 12 |
A' |
1479 |
1340 |
4.94 |
|
|
|
| 13 |
A' |
1310 |
1187 |
0.23 |
|
|
|
| 14 |
A' |
1289 |
1169 |
0.48 |
|
|
|
| 15 |
A' |
1173 |
1064 |
5.56 |
|
|
|
| 16 |
A' |
1121 |
1016 |
2.43 |
|
|
|
| 17 |
A' |
1113 |
1008 |
0.08 |
|
|
|
| 18 |
A' |
1085 |
984 |
0.00 |
|
|
|
| 19 |
A' |
1031 |
934 |
1.14 |
|
|
|
| 20 |
A' |
1026 |
930 |
2.12 |
|
|
|
| 21 |
A' |
851 |
772 |
34.52 |
|
|
|
| 22 |
A' |
822 |
745 |
12.52 |
|
|
|
| 23 |
A' |
777 |
705 |
40.81 |
|
|
|
| 24 |
A' |
611 |
554 |
9.86 |
|
|
|
| 25 |
A' |
537 |
486 |
5.18 |
|
|
|
| 26 |
A' |
337 |
306 |
0.23 |
|
|
|
| 27 |
A' |
148 |
135 |
0.92 |
|
|
|
| 28 |
A" |
3332 |
3021 |
44.35 |
|
|
|
| 29 |
A" |
3309 |
3000 |
4.69 |
|
|
|
| 30 |
A" |
3228 |
2926 |
54.07 |
|
|
|
| 31 |
A" |
3196 |
2897 |
3.80 |
|
|
|
| 32 |
A" |
1762 |
1597 |
1.78 |
|
|
|
| 33 |
A" |
1611 |
1461 |
7.04 |
|
|
|
| 34 |
A" |
1604 |
1454 |
8.09 |
|
|
|
| 35 |
A" |
1477 |
1339 |
0.37 |
|
|
|
| 36 |
A" |
1391 |
1261 |
0.06 |
|
|
|
| 37 |
A" |
1324 |
1200 |
0.00 |
|
|
|
| 38 |
A" |
1228 |
1113 |
0.16 |
|
|
|
| 39 |
A" |
1165 |
1056 |
3.82 |
|
|
|
| 40 |
A" |
1124 |
1018 |
0.46 |
|
|
|
| 41 |
A" |
1101 |
998 |
0.01 |
|
|
|
| 42 |
A" |
951 |
862 |
0.01 |
|
|
|
| 43 |
A" |
847 |
767 |
0.37 |
|
|
|
| 44 |
A" |
680 |
616 |
0.03 |
|
|
|
| 45 |
A" |
455 |
412 |
0.00 |
|
|
|
| 46 |
A" |
381 |
346 |
0.16 |
|
|
|
| 47 |
A" |
236 |
214 |
0.04 |
|
|
|
| 48 |
A" |
43 |
39 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36443.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 33032.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.153 |
|
|
|
| 2 |
C |
-0.178 |
|
|
|
| 3 |
C |
-0.068 |
|
|
|
| 4 |
C |
-0.131 |
|
|
|
| 5 |
C |
-0.068 |
|
|
|
| 6 |
C |
-0.178 |
|
|
|
| 7 |
C |
-0.069 |
|
|
|
| 8 |
C |
-0.172 |
|
|
|
| 9 |
H |
0.082 |
|
|
|
| 10 |
H |
0.093 |
|
|
|
| 11 |
H |
0.090 |
|
|
|
| 12 |
H |
0.093 |
|
|
|
| 13 |
H |
0.082 |
|
|
|
| 14 |
H |
0.053 |
|
|
|
| 15 |
H |
0.053 |
|
|
|
| 16 |
H |
0.054 |
|
|
|
| 17 |
H |
0.054 |
|
|
|
| 18 |
H |
0.059 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.043 |
0.382 |
0.000 |
0.384 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-53.783 |
-0.163 |
0.000 |
| y |
-0.163 |
-45.457 |
0.000 |
| z |
0.000 |
0.000 |
-45.336 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.387 |
-0.163 |
0.000 |
| y |
-0.163 |
4.103 |
0.000 |
| z |
0.000 |
0.000 |
4.284 |
|
| Polar |
|---|
| 3z2-r2 | 8.568 |
|---|
| x2-y2 | -8.327 |
|---|
| xy | -0.163 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.006 |
0.692 |
0.000 |
| y |
0.692 |
16.133 |
0.000 |
| z |
0.000 |
0.000 |
13.500 |
<r2> (average value of r
2) Å
2
| <r2> |
289.162 |
| (<r2>)1/2 |
17.005 |