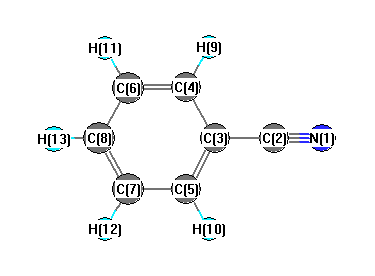
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3363 |
3048 |
4.26 |
|
|
|
| 2 |
A1 |
3349 |
3035 |
11.96 |
|
|
|
| 3 |
A1 |
3328 |
3017 |
0.08 |
|
|
|
| 4 |
A1 |
2579 |
2338 |
55.26 |
|
|
|
| 5 |
A1 |
1785 |
1618 |
0.89 |
|
|
|
| 6 |
A1 |
1651 |
1497 |
16.13 |
|
|
|
| 7 |
A1 |
1293 |
1172 |
0.37 |
|
|
|
| 8 |
A1 |
1285 |
1165 |
0.98 |
|
|
|
| 9 |
A1 |
1118 |
1014 |
2.66 |
|
|
|
| 10 |
A1 |
1085 |
984 |
0.02 |
|
|
|
| 11 |
A1 |
817 |
741 |
3.41 |
|
|
|
| 12 |
A1 |
493 |
447 |
0.04 |
|
|
|
| 13 |
A2 |
1111 |
1007 |
0.00 |
|
|
|
| 14 |
A2 |
955 |
866 |
0.00 |
|
|
|
| 15 |
A2 |
448 |
406 |
0.00 |
|
|
|
| 16 |
B1 |
1130 |
1024 |
0.00 |
|
|
|
| 17 |
B1 |
1059 |
960 |
3.29 |
|
|
|
| 18 |
B1 |
857 |
777 |
49.38 |
|
|
|
| 19 |
B1 |
763 |
692 |
39.52 |
|
|
|
| 20 |
B1 |
627 |
568 |
21.77 |
|
|
|
| 21 |
B1 |
433 |
392 |
0.74 |
|
|
|
| 22 |
B1 |
162 |
147 |
2.13 |
|
|
|
| 23 |
B2 |
3359 |
3044 |
11.24 |
|
|
|
| 24 |
B2 |
3341 |
3028 |
3.73 |
|
|
|
| 25 |
B2 |
1758 |
1593 |
0.52 |
|
|
|
| 26 |
B2 |
1598 |
1448 |
10.11 |
|
|
|
| 27 |
B2 |
1464 |
1327 |
0.68 |
|
|
|
| 28 |
B2 |
1324 |
1200 |
2.26 |
|
|
|
| 29 |
B2 |
1201 |
1088 |
8.50 |
|
|
|
| 30 |
B2 |
1166 |
1057 |
0.03 |
|
|
|
| 31 |
B2 |
686 |
621 |
0.30 |
|
|
|
| 32 |
B2 |
620 |
562 |
0.89 |
|
|
|
| 33 |
B2 |
187 |
169 |
5.99 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 23197.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 21026.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.125 |
|
|
|
| 2 |
C |
-0.075 |
|
|
|
| 3 |
C |
0.263 |
|
|
|
| 4 |
C |
-0.181 |
|
|
|
| 5 |
C |
-0.181 |
|
|
|
| 6 |
C |
-0.077 |
|
|
|
| 7 |
C |
-0.077 |
|
|
|
| 8 |
C |
-0.073 |
|
|
|
| 9 |
H |
0.109 |
|
|
|
| 10 |
H |
0.109 |
|
|
|
| 11 |
H |
0.103 |
|
|
|
| 12 |
H |
0.103 |
|
|
|
| 13 |
H |
0.103 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.913 |
4.913 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.727 |
0.000 |
0.000 |
| y |
0.000 |
-39.566 |
0.000 |
| z |
0.000 |
0.000 |
-57.358 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.265 |
0.000 |
0.000 |
| y |
0.000 |
13.977 |
0.000 |
| z |
0.000 |
0.000 |
-12.712 |
|
| Polar |
|---|
| 3z2-r2 | -25.424 |
|---|
| x2-y2 | -10.161 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.657 |
0.000 |
0.000 |
| y |
0.000 |
11.751 |
0.000 |
| z |
0.000 |
0.000 |
16.246 |
<r2> (average value of r
2) Å
2
| <r2> |
253.531 |
| (<r2>)1/2 |
15.923 |