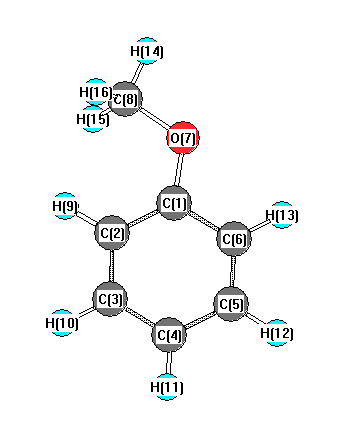
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3368 |
3053 |
8.10 |
|
|
|
| 2 |
A' |
3353 |
3039 |
11.83 |
|
|
|
| 3 |
A' |
3345 |
3031 |
26.46 |
|
|
|
| 4 |
A' |
3326 |
3015 |
13.68 |
|
|
|
| 5 |
A' |
3317 |
3007 |
1.63 |
|
|
|
| 6 |
A' |
3271 |
2965 |
39.45 |
|
|
|
| 7 |
A' |
3159 |
2863 |
50.57 |
|
|
|
| 8 |
A' |
1785 |
1618 |
101.57 |
|
|
|
| 9 |
A' |
1766 |
1600 |
28.55 |
|
|
|
| 10 |
A' |
1660 |
1505 |
110.35 |
|
|
|
| 11 |
A' |
1632 |
1479 |
19.79 |
|
|
|
| 12 |
A' |
1610 |
1459 |
18.08 |
|
|
|
| 13 |
A' |
1606 |
1456 |
7.47 |
|
|
|
| 14 |
A' |
1467 |
1330 |
5.37 |
|
|
|
| 15 |
A' |
1417 |
1284 |
297.27 |
|
|
|
| 16 |
A' |
1329 |
1205 |
37.94 |
|
|
|
| 17 |
A' |
1307 |
1185 |
3.22 |
|
|
|
| 18 |
A' |
1283 |
1163 |
13.53 |
|
|
|
| 19 |
A' |
1200 |
1087 |
4.15 |
|
|
|
| 20 |
A' |
1180 |
1069 |
56.80 |
|
|
|
| 21 |
A' |
1170 |
1061 |
14.97 |
|
|
|
| 22 |
A' |
1113 |
1009 |
2.05 |
|
|
|
| 23 |
A' |
1077 |
976 |
3.17 |
|
|
|
| 24 |
A' |
856 |
776 |
16.08 |
|
|
|
| 25 |
A' |
673 |
610 |
0.62 |
|
|
|
| 26 |
A' |
600 |
544 |
6.96 |
|
|
|
| 27 |
A' |
478 |
433 |
0.80 |
|
|
|
| 28 |
A' |
274 |
248 |
2.69 |
|
|
|
| 29 |
A" |
3214 |
2914 |
48.14 |
|
|
|
| 30 |
A" |
1622 |
1470 |
6.80 |
|
|
|
| 31 |
A" |
1285 |
1164 |
2.67 |
|
|
|
| 32 |
A" |
1103 |
1000 |
0.32 |
|
|
|
| 33 |
A" |
1094 |
991 |
0.00 |
|
|
|
| 34 |
A" |
1002 |
909 |
10.61 |
|
|
|
| 35 |
A" |
927 |
840 |
0.09 |
|
|
|
| 36 |
A" |
848 |
768 |
77.48 |
|
|
|
| 37 |
A" |
769 |
697 |
25.64 |
|
|
|
| 38 |
A" |
572 |
518 |
11.41 |
|
|
|
| 39 |
A" |
463 |
420 |
0.00 |
|
|
|
| 40 |
A" |
293 |
266 |
0.48 |
|
|
|
| 41 |
A" |
228 |
206 |
0.03 |
|
|
|
| 42 |
A" |
85 |
77 |
4.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31062.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 28154.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.338 |
|
|
|
| 2 |
C |
-0.167 |
|
|
|
| 3 |
C |
-0.103 |
|
|
|
| 4 |
C |
-0.126 |
|
|
|
| 5 |
C |
-0.046 |
|
|
|
| 6 |
C |
-0.173 |
|
|
|
| 7 |
O |
-0.378 |
|
|
|
| 8 |
C |
0.005 |
|
|
|
| 9 |
H |
0.075 |
|
|
|
| 10 |
H |
0.091 |
|
|
|
| 11 |
H |
0.090 |
|
|
|
| 12 |
H |
0.091 |
|
|
|
| 13 |
H |
0.089 |
|
|
|
| 14 |
H |
0.085 |
|
|
|
| 15 |
H |
0.064 |
|
|
|
| 16 |
H |
0.064 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.293 |
0.064 |
0.000 |
1.294 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.972 |
3.065 |
0.000 |
| y |
3.065 |
-44.296 |
0.000 |
| z |
0.000 |
0.000 |
-51.814 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
7.083 |
3.065 |
0.000 |
| y |
3.065 |
2.097 |
0.000 |
| z |
0.000 |
0.000 |
-9.180 |
|
| Polar |
|---|
| 3z2-r2 | -18.361 |
|---|
| x2-y2 | 3.324 |
|---|
| xy | 3.065 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.262 |
1.208 |
0.000 |
| y |
1.208 |
14.297 |
0.000 |
| z |
0.000 |
0.000 |
7.541 |
<r2> (average value of r
2) Å
2
| <r2> |
270.860 |
| (<r2>)1/2 |
16.458 |