Jump to
S1C2
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -5226.125843 |
| Energy at 298.15K | |
| HF Energy | -5225.835343 |
| Nuclear repulsion energy | 417.216415 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3129 |
3129 |
0.00 |
158.25 |
0.04 |
0.07 |
| 2 |
Ag |
1499 |
1499 |
0.00 |
8.60 |
0.73 |
0.84 |
| 3 |
Ag |
1292 |
1292 |
0.00 |
38.01 |
0.42 |
0.59 |
| 4 |
Ag |
1074 |
1074 |
0.00 |
11.03 |
0.72 |
0.84 |
| 5 |
Ag |
677 |
677 |
0.00 |
90.71 |
0.28 |
0.44 |
| 6 |
Ag |
191 |
191 |
0.00 |
4.58 |
0.28 |
0.44 |
| 7 |
Au |
3214 |
3214 |
0.12 |
0.00 |
0.69 |
0.81 |
| 8 |
Au |
1112 |
1112 |
3.10 |
0.00 |
0.73 |
0.84 |
| 9 |
Au |
763 |
763 |
3.35 |
0.00 |
0.73 |
0.84 |
| 10 |
Au |
103 |
103 |
4.36 |
0.00 |
0.00 |
0.00 |
| 11 |
Bg |
3191 |
3191 |
0.00 |
76.10 |
0.75 |
0.86 |
| 12 |
Bg |
1298 |
1298 |
0.00 |
2.57 |
0.75 |
0.86 |
| 13 |
Bg |
955 |
955 |
0.00 |
2.27 |
0.75 |
0.86 |
| 14 |
Bu |
3138 |
3138 |
5.75 |
0.00 |
0.50 |
0.67 |
| 15 |
Bu |
1496 |
1496 |
7.21 |
0.00 |
0.00 |
0.00 |
| 16 |
Bu |
1219 |
1219 |
54.59 |
0.00 |
0.00 |
0.00 |
| 17 |
Bu |
605 |
605 |
67.11 |
0.00 |
0.00 |
0.00 |
| 18 |
Bu |
180 |
180 |
6.39 |
0.00 |
0.49 |
0.66 |
Unscaled Zero Point Vibrational Energy (zpe) 12568.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12568.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.493 |
0.569 |
0.000 |
| C2 |
-0.493 |
-0.569 |
0.000 |
| Br3 |
-0.493 |
2.265 |
0.000 |
| Br4 |
0.493 |
-2.265 |
0.000 |
| H5 |
1.112 |
0.575 |
0.889 |
| H6 |
1.112 |
0.575 |
-0.889 |
| H7 |
-1.112 |
-0.575 |
0.889 |
| H8 |
-1.112 |
-0.575 |
-0.889 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5053 | 1.9616 | 2.8345 | 1.0831 | 1.0831 | 2.1618 | 2.1618 |
C2 | 1.5053 | | 2.8345 | 1.9616 | 2.1618 | 2.1618 | 1.0831 | 1.0831 | Br3 | 1.9616 | 2.8345 | | 4.6366 | 2.4940 | 2.4940 | 3.0402 | 3.0402 | Br4 | 2.8345 | 1.9616 | 4.6366 | | 3.0402 | 3.0402 | 2.4940 | 2.4940 | H5 | 1.0831 | 2.1618 | 2.4940 | 3.0402 | | 1.7773 | 2.5036 | 3.0703 | H6 | 1.0831 | 2.1618 | 2.4940 | 3.0402 | 1.7773 | | 3.0703 | 2.5036 | H7 | 2.1618 | 1.0831 | 3.0402 | 2.4940 | 2.5036 | 3.0703 | | 1.7773 | H8 | 2.1618 | 1.0831 | 3.0402 | 2.4940 | 3.0703 | 2.5036 | 1.7773 | |
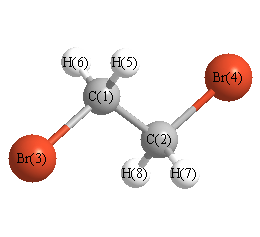 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
108.977 |
|
C1 |
C2 |
H7 |
112.240 |
| C1 |
C2 |
H8 |
112.240 |
|
C2 |
C1 |
Br3 |
108.977 |
| C2 |
C1 |
H5 |
112.240 |
|
C2 |
C1 |
H6 |
112.240 |
| Br3 |
C1 |
H5 |
106.387 |
|
Br3 |
C1 |
H6 |
106.387 |
| Br4 |
C2 |
H7 |
106.387 |
|
Br4 |
C2 |
H8 |
106.387 |
| H5 |
C1 |
H6 |
110.259 |
|
H7 |
C2 |
H8 |
110.259 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.149 |
|
|
|
| 2 |
C |
-0.149 |
|
|
|
| 3 |
Br |
-0.128 |
|
|
|
| 4 |
Br |
-0.128 |
|
|
|
| 5 |
H |
0.138 |
|
|
|
| 6 |
H |
0.138 |
|
|
|
| 7 |
H |
0.138 |
|
|
|
| 8 |
H |
0.138 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.234 |
1.373 |
0.000 |
| y |
1.373 |
-56.292 |
0.000 |
| z |
0.000 |
0.000 |
-49.786 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.805 |
1.373 |
0.000 |
| y |
1.373 |
-6.782 |
0.000 |
| z |
0.000 |
0.000 |
2.977 |
|
| Polar |
|---|
| 3z2-r2 | 5.954 |
|---|
| x2-y2 | 7.058 |
|---|
| xy | 1.373 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.130 |
-1.882 |
0.000 |
| y |
-1.882 |
13.789 |
0.000 |
| z |
0.000 |
0.000 |
7.098 |
<r2> (average value of r
2) Å
2
| <r2> |
424.778 |
| (<r2>)1/2 |
20.610 |
Jump to
S1C1
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -5226.122354 |
| Energy at 298.15K | |
| HF Energy | -5225.831435 |
| Nuclear repulsion energy | 451.118311 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3171 |
3171 |
0.01 |
86.98 |
0.75 |
0.86 |
| 2 |
A |
3111 |
3111 |
14.80 |
242.96 |
0.02 |
0.03 |
| 3 |
A |
1478 |
1478 |
0.46 |
3.57 |
0.72 |
0.84 |
| 4 |
A |
1320 |
1320 |
19.30 |
2.61 |
0.40 |
0.57 |
| 5 |
A |
1201 |
1201 |
1.63 |
7.33 |
0.72 |
0.84 |
| 6 |
A |
1044 |
1044 |
0.88 |
1.65 |
0.70 |
0.82 |
| 7 |
A |
917 |
917 |
7.11 |
4.63 |
0.29 |
0.44 |
| 8 |
A |
566 |
566 |
7.41 |
18.63 |
0.07 |
0.13 |
| 9 |
A |
229 |
229 |
1.14 |
1.45 |
0.37 |
0.54 |
| 10 |
A |
79 |
79 |
0.23 |
1.04 |
0.68 |
0.81 |
| 11 |
B |
3184 |
3184 |
0.87 |
25.43 |
0.75 |
0.86 |
| 12 |
B |
3103 |
3103 |
2.10 |
43.12 |
0.75 |
0.86 |
| 13 |
B |
1476 |
1476 |
10.93 |
10.15 |
0.75 |
0.86 |
| 14 |
B |
1289 |
1289 |
64.22 |
3.34 |
0.75 |
0.86 |
| 15 |
B |
1133 |
1133 |
1.94 |
2.39 |
0.75 |
0.86 |
| 16 |
B |
856 |
856 |
17.23 |
1.23 |
0.75 |
0.86 |
| 17 |
B |
600 |
600 |
14.21 |
9.76 |
0.75 |
0.86 |
| 18 |
B |
358 |
358 |
5.86 |
2.19 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12557.2 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12557.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.311 |
0.686 |
1.179 |
| C2 |
-0.311 |
-0.686 |
1.179 |
| Br3 |
-0.311 |
1.805 |
-0.293 |
| Br4 |
0.311 |
-1.805 |
-0.293 |
| H5 |
0.031 |
1.213 |
2.087 |
| H6 |
1.391 |
0.643 |
1.098 |
| H7 |
-0.031 |
-1.213 |
2.087 |
| H8 |
-1.391 |
-0.643 |
1.098 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Br4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5069 | 1.9508 | 2.8937 | 1.0866 | 1.0840 | 2.1330 | 2.1614 |
C2 | 1.5069 | | 2.8937 | 1.9508 | 2.1330 | 2.1614 | 1.0866 | 1.0840 | Br3 | 1.9508 | 2.8937 | | 3.6630 | 2.4763 | 2.4865 | 3.8541 | 3.0162 | Br4 | 2.8937 | 1.9508 | 3.6630 | | 3.8541 | 3.0162 | 2.4763 | 2.4865 | H5 | 1.0866 | 2.1330 | 2.4763 | 3.8541 | | 1.7756 | 2.4275 | 2.5390 | H6 | 1.0840 | 2.1614 | 2.4865 | 3.0162 | 1.7756 | | 2.5390 | 3.0654 | H7 | 2.1330 | 1.0866 | 3.8541 | 2.4763 | 2.4275 | 2.5390 | | 1.7756 | H8 | 2.1614 | 1.0840 | 3.0162 | 2.4865 | 2.5390 | 3.0654 | 1.7756 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Br4 |
112.999 |
|
C1 |
C2 |
H7 |
109.595 |
| C1 |
C2 |
H8 |
112.041 |
|
C2 |
C1 |
Br3 |
112.999 |
| C2 |
C1 |
H5 |
109.595 |
|
C2 |
C1 |
H6 |
112.041 |
| Br3 |
C1 |
H5 |
105.676 |
|
Br3 |
C1 |
H6 |
106.514 |
| Br4 |
C2 |
H7 |
105.676 |
|
Br4 |
C2 |
H8 |
106.514 |
| H5 |
C1 |
H6 |
109.774 |
|
H7 |
C2 |
H8 |
109.774 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.148 |
|
|
|
| 2 |
C |
-0.148 |
|
|
|
| 3 |
Br |
-0.114 |
|
|
|
| 4 |
Br |
-0.114 |
|
|
|
| 5 |
H |
0.131 |
|
|
|
| 6 |
H |
0.131 |
|
|
|
| 7 |
H |
0.131 |
|
|
|
| 8 |
H |
0.131 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.599 |
2.599 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-49.588 |
0.935 |
0.000 |
| y |
0.935 |
-54.275 |
0.000 |
| z |
0.000 |
0.000 |
-45.456 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.278 |
0.935 |
0.000 |
| y |
0.935 |
-6.754 |
0.000 |
| z |
0.000 |
0.000 |
6.476 |
|
| Polar |
|---|
| 3z2-r2 | 12.951 |
|---|
| x2-y2 | 4.688 |
|---|
| xy | 0.935 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.475 |
-0.745 |
0.000 |
| y |
-0.745 |
10.767 |
0.000 |
| z |
0.000 |
0.000 |
9.363 |
<r2> (average value of r
2) Å
2
| <r2> |
314.178 |
| (<r2>)1/2 |
17.725 |
