All results from a given calculation for CH2CHCH2CH3 (1-Butene)
using model chemistry: QCISD(T)=FULL/Def2TZVPP
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C1 |
1A |
Energy calculated at QCISD(T)=FULL/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -157.003812 |
| Energy at 298.15K | |
| HF Energy | -156.168192 |
| Nuclear repulsion energy | 117.283552 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/Def2TZVPP
Geometric Data calculated at QCISD(T)=FULL/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.716 |
-0.248 |
-0.295 |
| C2 |
0.540 |
0.525 |
0.308 |
| C3 |
-0.714 |
-0.299 |
0.344 |
| C4 |
-1.853 |
0.019 |
-0.280 |
| H5 |
-0.664 |
-1.223 |
0.913 |
| H6 |
1.500 |
-0.535 |
-1.324 |
| H7 |
2.625 |
0.354 |
-0.291 |
| H8 |
1.912 |
-1.159 |
0.272 |
| H9 |
0.798 |
0.833 |
1.324 |
| H10 |
0.356 |
1.436 |
-0.265 |
| H11 |
-1.938 |
0.930 |
-0.857 |
| H12 |
-2.724 |
-0.619 |
-0.228 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
| C1 | | 1.5312 | 2.5133 | 3.5791 | 2.8417 | 1.0899 | 1.0897 | 1.0908 | 2.1528 | 2.1643 | 3.8800 | 4.4559 |
C2 | 1.5312 | | 1.5008 | 2.5152 | 2.2067 | 2.1698 | 2.1756 | 2.1722 | 1.0933 | 1.0916 | 2.7677 | 3.4995 | C3 | 2.5133 | 1.5008 | | 1.3368 | 1.0857 | 2.7816 | 3.4605 | 2.7639 | 2.1283 | 2.1276 | 2.1100 | 2.1140 | C4 | 3.5791 | 2.5152 | 1.3368 | | 2.0920 | 3.5549 | 4.4900 | 3.9831 | 3.2036 | 2.6246 | 1.0824 | 1.0806 | H5 | 2.8417 | 2.2067 | 1.0857 | 2.0920 | | 3.1870 | 3.8405 | 2.6552 | 2.5559 | 3.0816 | 3.0644 | 2.4309 | H6 | 1.0899 | 2.1698 | 2.7816 | 3.5549 | 3.1870 | | 1.7676 | 1.7623 | 3.0625 | 2.5125 | 3.7656 | 4.3640 | H7 | 1.0897 | 2.1756 | 3.4605 | 4.4900 | 3.8405 | 1.7676 | | 1.7643 | 2.4848 | 2.5131 | 4.6333 | 5.4365 | H8 | 1.0908 | 2.1722 | 2.7639 | 3.9831 | 2.6552 | 1.7623 | 1.7643 | | 2.5130 | 3.0723 | 4.5231 | 4.6937 | H9 | 2.1528 | 1.0933 | 2.1283 | 3.2036 | 2.5559 | 3.0625 | 2.4848 | 2.5130 | | 1.7567 | 3.5006 | 4.1136 | H10 | 2.1643 | 1.0916 | 2.1276 | 2.6246 | 3.0816 | 2.5125 | 2.5131 | 3.0723 | 1.7567 | | 2.4225 | 3.7026 | H11 | 3.8800 | 2.7677 | 2.1100 | 1.0824 | 3.0644 | 3.7656 | 4.6333 | 4.5231 | 3.5006 | 2.4225 | | 1.8476 | H12 | 4.4559 | 3.4995 | 2.1140 | 1.0806 | 2.4309 | 4.3640 | 5.4365 | 4.6937 | 4.1136 | 3.7026 | 1.8476 | |
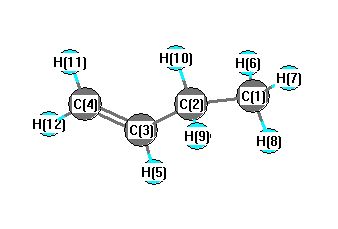 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
111.972 |
|
C1 |
C2 |
H9 |
109.087 |
| C1 |
C2 |
H10 |
110.084 |
|
C2 |
C1 |
H6 |
110.622 |
| C2 |
C1 |
H7 |
111.103 |
|
C2 |
C1 |
H8 |
110.763 |
| C2 |
C3 |
C4 |
124.738 |
|
C2 |
C3 |
H5 |
116.183 |
| C3 |
C2 |
H9 |
109.247 |
|
C3 |
C2 |
H10 |
109.290 |
| C3 |
C4 |
H11 |
121.067 |
|
C3 |
C4 |
H12 |
121.601 |
| C4 |
C3 |
H5 |
119.073 |
|
H6 |
C1 |
H7 |
108.386 |
| H6 |
C1 |
H8 |
107.828 |
|
H7 |
C1 |
H8 |
108.023 |
| H9 |
C2 |
H10 |
107.031 |
|
H11 |
C4 |
H12 |
117.332 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
