Jump to
S1C2
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -191.836698 |
| Energy at 298.15K | |
| HF Energy | -191.603930 |
| Nuclear repulsion energy | 103.231159 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3258 |
3258 |
3.88 |
62.69 |
0.52 |
0.69 |
| 2 |
A' |
3205 |
3205 |
1.14 |
98.12 |
0.29 |
0.45 |
| 3 |
A' |
3161 |
3161 |
4.44 |
82.50 |
0.11 |
0.19 |
| 4 |
A' |
2918 |
2918 |
88.83 |
105.54 |
0.26 |
0.42 |
| 5 |
A' |
1754 |
1754 |
218.56 |
75.02 |
0.37 |
0.54 |
| 6 |
A' |
1675 |
1675 |
3.04 |
25.07 |
0.12 |
0.22 |
| 7 |
A' |
1466 |
1466 |
10.52 |
6.08 |
0.51 |
0.67 |
| 8 |
A' |
1398 |
1398 |
5.34 |
21.59 |
0.31 |
0.47 |
| 9 |
A' |
1307 |
1307 |
2.90 |
9.50 |
0.20 |
0.33 |
| 10 |
A' |
1180 |
1180 |
38.12 |
13.02 |
0.58 |
0.74 |
| 11 |
A' |
929 |
929 |
20.57 |
1.23 |
0.02 |
0.03 |
| 12 |
A' |
573 |
573 |
4.31 |
5.85 |
0.26 |
0.41 |
| 13 |
A' |
322 |
322 |
10.25 |
0.91 |
0.31 |
0.48 |
| 14 |
A" |
1042 |
1042 |
13.39 |
0.60 |
0.75 |
0.86 |
| 15 |
A" |
1029 |
1029 |
5.77 |
2.78 |
0.75 |
0.86 |
| 16 |
A" |
1001 |
1001 |
38.95 |
1.67 |
0.75 |
0.86 |
| 17 |
A" |
618 |
618 |
10.29 |
1.09 |
0.75 |
0.86 |
| 18 |
A" |
168 |
168 |
4.32 |
0.78 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13501.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13501.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.149 |
-0.742 |
0.000 |
| C2 |
0.000 |
0.720 |
0.000 |
| C3 |
1.211 |
1.280 |
0.000 |
| O4 |
-1.214 |
-1.321 |
0.000 |
| H5 |
0.804 |
-1.305 |
0.000 |
| H6 |
-0.910 |
1.305 |
0.000 |
| H7 |
1.349 |
2.351 |
0.000 |
| H8 |
2.103 |
0.667 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4701 | 2.4374 | 1.2118 | 1.1069 | 2.1841 | 3.4372 | 2.6565 |
C2 | 1.4701 | | 1.3341 | 2.3746 | 2.1787 | 1.0816 | 2.1166 | 2.1032 | C3 | 2.4374 | 1.3341 | | 3.5560 | 2.6169 | 2.1207 | 1.0796 | 1.0824 | O4 | 1.2118 | 2.3746 | 3.5560 | | 2.0185 | 2.6431 | 4.4778 | 3.8667 | H5 | 1.1069 | 2.1787 | 2.6169 | 2.0185 | | 3.1223 | 3.6962 | 2.3608 | H6 | 2.1841 | 1.0816 | 2.1207 | 2.6431 | 3.1223 | | 2.4894 | 3.0792 | H7 | 3.4372 | 2.1166 | 1.0796 | 4.4778 | 3.6962 | 2.4894 | | 1.8450 | H8 | 2.6565 | 2.1032 | 1.0824 | 3.8667 | 2.3608 | 3.0792 | 1.8450 | |
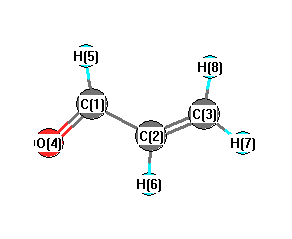 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.658 |
|
C1 |
C2 |
H6 |
116.909 |
| C2 |
C1 |
O4 |
124.322 |
|
C2 |
C1 |
H5 |
114.705 |
| C2 |
C3 |
H7 |
122.197 |
|
C2 |
C3 |
H8 |
120.648 |
| C3 |
C2 |
H6 |
122.433 |
|
O4 |
C1 |
H5 |
120.973 |
| H7 |
C3 |
H8 |
117.155 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.157 |
|
|
|
| 2 |
C |
-0.102 |
|
|
|
| 3 |
C |
-0.156 |
|
|
|
| 4 |
O |
-0.281 |
|
|
|
| 5 |
H |
0.050 |
|
|
|
| 6 |
H |
0.119 |
|
|
|
| 7 |
H |
0.113 |
|
|
|
| 8 |
H |
0.099 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.490 |
2.104 |
0.000 |
3.260 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.829 |
-2.679 |
-0.005 |
| y |
-2.679 |
-24.291 |
-0.006 |
| z |
-0.005 |
-0.006 |
-25.122 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.122 |
-2.679 |
-0.005 |
| y |
-2.679 |
0.684 |
-0.006 |
| z |
-0.005 |
-0.006 |
-0.562 |
|
| Polar |
|---|
| 3z2-r2 | -1.124 |
|---|
| x2-y2 | -0.538 |
|---|
| xy | -2.679 |
|---|
| xz | -0.005 |
|---|
| yz | -0.006 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.364 |
1.748 |
0.001 |
| y |
1.748 |
6.806 |
0.001 |
| z |
0.001 |
0.001 |
3.579 |
<r2> (average value of r
2) Å
2
| <r2> |
83.477 |
| (<r2>)1/2 |
9.137 |
Jump to
S1C1
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -191.833217 |
| Energy at 298.15K | |
| HF Energy | -191.600296 |
| Nuclear repulsion energy | 104.719363 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3268 |
3268 |
1.37 |
52.06 |
0.61 |
0.76 |
| 2 |
A' |
3193 |
3193 |
9.71 |
127.17 |
0.22 |
0.36 |
| 3 |
A' |
3170 |
3170 |
4.47 |
60.83 |
0.13 |
0.24 |
| 4 |
A' |
2944 |
2944 |
131.48 |
169.05 |
0.29 |
0.45 |
| 5 |
A' |
1760 |
1760 |
90.56 |
12.94 |
0.65 |
0.79 |
| 6 |
A' |
1665 |
1665 |
72.32 |
56.18 |
0.17 |
0.28 |
| 7 |
A' |
1449 |
1449 |
42.74 |
7.91 |
0.50 |
0.67 |
| 8 |
A' |
1436 |
1436 |
1.12 |
10.46 |
0.21 |
0.34 |
| 9 |
A' |
1322 |
1322 |
2.53 |
17.41 |
0.31 |
0.47 |
| 10 |
A' |
1078 |
1078 |
4.10 |
4.09 |
0.69 |
0.81 |
| 11 |
A' |
937 |
937 |
61.81 |
5.05 |
0.13 |
0.22 |
| 12 |
A' |
685 |
685 |
11.45 |
1.02 |
0.72 |
0.84 |
| 13 |
A' |
286 |
286 |
6.84 |
3.43 |
0.42 |
0.59 |
| 14 |
A" |
1040 |
1040 |
9.81 |
5.25 |
0.75 |
0.86 |
| 15 |
A" |
1031 |
1031 |
27.62 |
1.09 |
0.75 |
0.86 |
| 16 |
A" |
1013 |
1013 |
13.05 |
0.83 |
0.75 |
0.86 |
| 17 |
A" |
562 |
562 |
10.47 |
1.73 |
0.75 |
0.86 |
| 18 |
A" |
151 |
151 |
6.36 |
0.12 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 13493.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13493.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.886 |
-0.292 |
0.000 |
| C2 |
0.000 |
0.895 |
0.000 |
| C3 |
1.328 |
0.778 |
0.000 |
| O4 |
-0.495 |
-1.440 |
0.000 |
| H5 |
-1.969 |
-0.073 |
0.000 |
| H6 |
-0.480 |
1.865 |
0.000 |
| H7 |
1.974 |
1.643 |
0.000 |
| H8 |
1.785 |
-0.201 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4815 | 2.4598 | 1.2123 | 1.1042 | 2.1950 | 3.4538 | 2.6730 |
C2 | 1.4815 | | 1.3335 | 2.3867 | 2.1938 | 1.0823 | 2.1114 | 2.0950 | C3 | 2.4598 | 1.3335 | | 2.8716 | 3.4052 | 2.1098 | 1.0796 | 1.0811 | O4 | 1.2123 | 2.3867 | 2.8716 | | 2.0093 | 3.3047 | 3.9503 | 2.5952 | H5 | 1.1042 | 2.1938 | 3.4052 | 2.0093 | | 2.4438 | 4.3004 | 3.7560 | H6 | 2.1950 | 1.0823 | 2.1098 | 3.3047 | 2.4438 | | 2.4644 | 3.0662 | H7 | 3.4538 | 2.1114 | 1.0796 | 3.9503 | 4.3004 | 2.4644 | | 1.8545 | H8 | 2.6730 | 2.0950 | 1.0811 | 2.5952 | 3.7560 | 3.0662 | 1.8545 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.727 |
|
C1 |
C2 |
H6 |
116.921 |
| C2 |
C1 |
O4 |
124.450 |
|
C2 |
C1 |
H5 |
115.313 |
| C2 |
C3 |
H7 |
121.732 |
|
C2 |
C3 |
H8 |
120.016 |
| C3 |
C2 |
H6 |
121.353 |
|
O4 |
C1 |
H5 |
120.237 |
| H7 |
C3 |
H8 |
118.252 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.167 |
|
|
|
| 2 |
C |
-0.134 |
|
|
|
| 3 |
C |
-0.147 |
|
|
|
| 4 |
O |
-0.290 |
|
|
|
| 5 |
H |
0.062 |
|
|
|
| 6 |
H |
0.107 |
|
|
|
| 7 |
H |
0.115 |
|
|
|
| 8 |
H |
0.120 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.201 |
2.701 |
0.000 |
2.708 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.510 |
-0.676 |
-0.003 |
| y |
-0.676 |
-26.576 |
-0.006 |
| z |
-0.003 |
-0.006 |
-25.096 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.326 |
-0.676 |
-0.003 |
| y |
-0.676 |
-3.773 |
-0.006 |
| z |
-0.003 |
-0.006 |
-1.553 |
|
| Polar |
|---|
| 3z2-r2 | -3.106 |
|---|
| x2-y2 | 6.066 |
|---|
| xy | -0.676 |
|---|
| xz | -0.003 |
|---|
| yz | -0.006 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.501 |
0.530 |
0.000 |
| y |
0.530 |
6.194 |
0.001 |
| z |
0.000 |
0.001 |
3.568 |
<r2> (average value of r
2) Å
2
| <r2> |
75.243 |
| (<r2>)1/2 |
8.674 |
