Jump to
S1C2
Energy calculated at QCISD(T)=FULL/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -191.680636 |
| Energy at 298.15K | |
| HF Energy | -190.840350 |
| Nuclear repulsion energy | 103.013220 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/Def2TZVPP
Geometric Data calculated at QCISD(T)=FULL/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.154 |
-0.743 |
0.000 |
| C2 |
0.000 |
0.725 |
0.000 |
| C3 |
1.222 |
1.274 |
0.000 |
| O4 |
-1.223 |
-1.317 |
0.000 |
| H5 |
0.798 |
-1.306 |
0.000 |
| H6 |
-0.905 |
1.316 |
0.000 |
| H7 |
1.373 |
2.343 |
0.000 |
| H8 |
2.105 |
0.648 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4763 | 2.4420 | 1.2131 | 1.1056 | 2.1923 | 3.4436 | 2.6530 |
C2 | 1.4763 | | 1.3398 | 2.3803 | 2.1820 | 1.0812 | 2.1222 | 2.1067 | C3 | 2.4420 | 1.3398 | | 3.5626 | 2.6147 | 2.1278 | 1.0798 | 1.0829 | O4 | 1.2131 | 2.3803 | 3.5626 | | 2.0204 | 2.6526 | 4.4874 | 3.8647 | H5 | 1.1056 | 2.1820 | 2.6147 | 2.0204 | | 3.1267 | 3.6943 | 2.3506 | H6 | 2.1923 | 1.0812 | 2.1278 | 2.6526 | 3.1267 | | 2.4989 | 3.0840 | H7 | 3.4436 | 2.1222 | 1.0798 | 4.4874 | 3.6943 | 2.4989 | | 1.8473 | H8 | 2.6530 | 2.1067 | 1.0829 | 3.8647 | 2.3506 | 3.0840 | 1.8473 | |
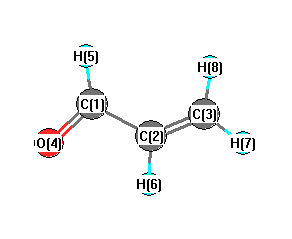 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.178 |
|
C1 |
C2 |
H6 |
117.165 |
| C2 |
C1 |
O4 |
124.227 |
|
C2 |
C1 |
H5 |
114.606 |
| C2 |
C3 |
H7 |
122.214 |
|
C2 |
C3 |
H8 |
120.450 |
| C3 |
C2 |
H6 |
122.656 |
|
O4 |
C1 |
H5 |
121.167 |
| H7 |
C3 |
H8 |
117.335 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)=FULL/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -191.677545 |
| Energy at 298.15K | |
| HF Energy | -190.836533 |
| Nuclear repulsion energy | 104.571164 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3252 |
3252 |
|
|
|
|
| 2 |
A' |
3183 |
3183 |
|
|
|
|
| 3 |
A' |
3155 |
3155 |
|
|
|
|
| 4 |
A' |
2948 |
2948 |
|
|
|
|
| 5 |
A' |
1756 |
1756 |
|
|
|
|
| 6 |
A' |
1658 |
1658 |
|
|
|
|
| 7 |
A' |
1444 |
1444 |
|
|
|
|
| 8 |
A' |
1431 |
1431 |
|
|
|
|
| 9 |
A' |
1312 |
1312 |
|
|
|
|
| 10 |
A' |
1069 |
1069 |
|
|
|
|
| 11 |
A' |
933 |
933 |
|
|
|
|
| 12 |
A' |
678 |
678 |
|
|
|
|
| 13 |
A' |
285 |
285 |
|
|
|
|
| 14 |
A" |
1023 |
1023 |
|
|
|
|
| 15 |
A" |
1015 |
1015 |
|
|
|
|
| 16 |
A" |
984 |
984 |
|
|
|
|
| 17 |
A" |
562 |
562 |
|
|
|
|
| 18 |
A" |
152 |
152 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13419.4 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 13419.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)=FULL/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.892 |
-0.289 |
0.000 |
| C2 |
0.000 |
0.901 |
0.000 |
| C3 |
1.333 |
0.768 |
0.000 |
| O4 |
-0.495 |
-1.436 |
0.000 |
| H5 |
-1.974 |
-0.073 |
0.000 |
| H6 |
-0.472 |
1.875 |
0.000 |
| H7 |
1.989 |
1.625 |
0.000 |
| H8 |
1.774 |
-0.219 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4874 | 2.4633 | 1.2139 | 1.1026 | 2.2039 | 3.4594 | 2.6676 |
C2 | 1.4874 | | 1.3395 | 2.3888 | 2.2011 | 1.0820 | 2.1170 | 2.0984 | C3 | 2.4633 | 1.3395 | | 2.8632 | 3.4119 | 2.1174 | 1.0798 | 1.0814 | O4 | 1.2139 | 2.3888 | 2.8632 | | 2.0107 | 3.3106 | 3.9424 | 2.5749 | H5 | 1.1026 | 2.2011 | 3.4119 | 2.0107 | | 2.4594 | 4.3117 | 3.7508 | H6 | 2.2039 | 1.0820 | 2.1174 | 3.3106 | 2.4594 | | 2.4743 | 3.0711 | H7 | 3.4594 | 2.1170 | 1.0798 | 3.9424 | 4.3117 | 2.4743 | | 1.8569 | H8 | 2.6676 | 2.0984 | 1.0814 | 2.5749 | 3.7508 | 3.0711 | 1.8569 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.158 |
|
C1 |
C2 |
H6 |
117.254 |
| C2 |
C1 |
O4 |
124.031 |
|
C2 |
C1 |
H5 |
115.595 |
| C2 |
C3 |
H7 |
121.737 |
|
C2 |
C3 |
H8 |
119.803 |
| C3 |
C2 |
H6 |
121.587 |
|
O4 |
C1 |
H5 |
120.374 |
| H7 |
C3 |
H8 |
118.460 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
