Jump to
S1C2
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -3112.435787 |
| Energy at 298.15K | |
| HF Energy | -3112.171662 |
| Nuclear repulsion energy | 288.682304 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3134 |
3134 |
7.55 |
30.42 |
0.05 |
0.09 |
| 2 |
A' |
3121 |
3121 |
2.36 |
139.28 |
0.04 |
0.09 |
| 3 |
A' |
1504 |
1504 |
3.96 |
4.57 |
0.72 |
0.84 |
| 4 |
A' |
1498 |
1498 |
3.25 |
5.24 |
0.74 |
0.85 |
| 5 |
A' |
1323 |
1323 |
1.59 |
15.30 |
0.51 |
0.68 |
| 6 |
A' |
1236 |
1236 |
41.84 |
2.94 |
0.38 |
0.55 |
| 7 |
A' |
1074 |
1074 |
0.98 |
6.23 |
0.74 |
0.85 |
| 8 |
A' |
746 |
746 |
25.03 |
47.28 |
0.29 |
0.45 |
| 9 |
A' |
649 |
649 |
56.69 |
13.29 |
0.25 |
0.40 |
| 10 |
A' |
249 |
249 |
0.81 |
4.75 |
0.30 |
0.46 |
| 11 |
A' |
195 |
195 |
6.64 |
0.24 |
0.25 |
0.40 |
| 12 |
A" |
3207 |
3207 |
0.77 |
7.24 |
0.75 |
0.86 |
| 13 |
A" |
3180 |
3180 |
0.40 |
77.09 |
0.75 |
0.86 |
| 14 |
A" |
1303 |
1303 |
0.01 |
3.72 |
0.75 |
0.86 |
| 15 |
A" |
1134 |
1134 |
2.18 |
0.04 |
0.75 |
0.86 |
| 16 |
A" |
983 |
983 |
0.19 |
2.41 |
0.75 |
0.86 |
| 17 |
A" |
772 |
772 |
2.77 |
0.02 |
0.75 |
0.86 |
| 18 |
A" |
111 |
111 |
5.37 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 12709.7 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12709.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.643 |
0.000 |
| C2 |
1.235 |
-0.223 |
0.000 |
| Br3 |
-1.588 |
-0.501 |
0.000 |
| Cl4 |
2.690 |
0.835 |
0.000 |
| H5 |
-0.062 |
1.260 |
0.888 |
| H6 |
-0.062 |
1.260 |
-0.888 |
| H7 |
1.288 |
-0.844 |
0.887 |
| H8 |
1.288 |
-0.844 |
-0.887 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5082 | 1.9575 | 2.6965 | 1.0833 | 1.0833 | 2.1584 | 2.1584 |
C2 | 1.5082 | | 2.8370 | 1.7984 | 2.1611 | 2.1611 | 1.0842 | 1.0842 | Br3 | 1.9575 | 2.8370 | | 4.4817 | 2.4937 | 2.4937 | 3.0296 | 3.0296 | Cl4 | 2.6965 | 1.7984 | 4.4817 | | 2.9231 | 2.9231 | 2.3600 | 2.3600 | H5 | 1.0833 | 2.1611 | 2.4937 | 2.9231 | | 1.7769 | 2.5005 | 3.0665 | H6 | 1.0833 | 2.1611 | 2.4937 | 2.9231 | 1.7769 | | 3.0665 | 2.5005 | H7 | 2.1584 | 1.0842 | 3.0296 | 2.3600 | 2.5005 | 3.0665 | | 1.7732 | H8 | 2.1584 | 1.0842 | 3.0296 | 2.3600 | 3.0665 | 2.5005 | 1.7732 | |
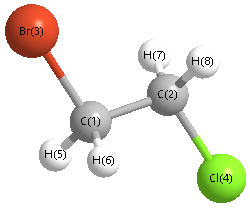 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
108.961 |
|
C1 |
C2 |
H7 |
111.683 |
| C1 |
C2 |
H8 |
111.683 |
|
C2 |
C1 |
Br3 |
109.200 |
| C2 |
C1 |
Cl4 |
39.103 |
|
C2 |
C1 |
H6 |
111.959 |
| Br3 |
C1 |
H5 |
106.619 |
|
Br3 |
C1 |
H6 |
106.619 |
| Cl4 |
C2 |
H7 |
107.301 |
|
Cl4 |
C2 |
H8 |
107.301 |
| H5 |
C1 |
H6 |
110.195 |
|
H7 |
C2 |
H8 |
109.714 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.147 |
|
|
|
| 2 |
C |
-0.092 |
|
|
|
| 3 |
Br |
-0.128 |
|
|
|
| 4 |
Cl |
-0.151 |
|
|
|
| 5 |
H |
0.134 |
|
|
|
| 6 |
H |
0.134 |
|
|
|
| 7 |
H |
0.125 |
|
|
|
| 8 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.004 |
0.006 |
0.000 |
0.007 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.272 |
-2.205 |
-0.010 |
| y |
-2.205 |
-43.677 |
-0.003 |
| z |
-0.010 |
-0.003 |
-43.762 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.552 |
-2.205 |
-0.010 |
| y |
-2.205 |
3.340 |
-0.003 |
| z |
-0.010 |
-0.003 |
3.212 |
|
| Polar |
|---|
| 3z2-r2 | 6.425 |
|---|
| x2-y2 | -6.595 |
|---|
| xy | -2.205 |
|---|
| xz | -0.010 |
|---|
| yz | -0.003 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.373 |
1.934 |
-0.001 |
| y |
1.934 |
7.593 |
-0.000 |
| z |
-0.001 |
-0.000 |
6.333 |
<r2> (average value of r
2) Å
2
| <r2> |
283.590 |
| (<r2>)1/2 |
16.840 |
Jump to
S1C1
Energy calculated at B2PLYP/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -3112.432918 |
| Energy at 298.15K | |
| HF Energy | -3112.168483 |
| Nuclear repulsion energy | 304.689762 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3183 |
3183 |
1.42 |
36.41 |
0.75 |
0.86 |
| 2 |
A |
3161 |
3161 |
1.27 |
77.46 |
0.75 |
0.86 |
| 3 |
A |
3112 |
3112 |
12.34 |
169.39 |
0.03 |
0.07 |
| 4 |
A |
3094 |
3094 |
9.09 |
108.48 |
0.17 |
0.30 |
| 5 |
A |
1484 |
1484 |
4.84 |
6.58 |
0.75 |
0.86 |
| 6 |
A |
1476 |
1476 |
6.78 |
7.21 |
0.73 |
0.85 |
| 7 |
A |
1342 |
1342 |
19.63 |
1.40 |
0.75 |
0.86 |
| 8 |
A |
1304 |
1304 |
47.88 |
2.93 |
0.44 |
0.61 |
| 9 |
A |
1218 |
1218 |
2.27 |
7.55 |
0.70 |
0.83 |
| 10 |
A |
1154 |
1154 |
1.28 |
2.56 |
0.72 |
0.84 |
| 11 |
A |
1049 |
1049 |
1.73 |
1.63 |
0.75 |
0.86 |
| 12 |
A |
942 |
942 |
8.25 |
3.68 |
0.29 |
0.46 |
| 13 |
A |
879 |
879 |
18.31 |
1.49 |
0.75 |
0.86 |
| 14 |
A |
682 |
682 |
19.56 |
8.84 |
0.39 |
0.56 |
| 15 |
A |
583 |
583 |
10.78 |
15.89 |
0.18 |
0.30 |
| 16 |
A |
388 |
388 |
6.94 |
2.11 |
0.70 |
0.82 |
| 17 |
A |
243 |
243 |
1.15 |
1.24 |
0.39 |
0.56 |
| 18 |
A |
98 |
98 |
0.53 |
1.33 |
0.70 |
0.82 |
Unscaled Zero Point Vibrational Energy (zpe) 12696.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 12696.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.013 |
1.099 |
-0.384 |
| C2 |
1.283 |
0.898 |
0.404 |
| Br3 |
-1.357 |
-0.224 |
0.036 |
| Cl4 |
2.171 |
-0.576 |
-0.086 |
| H5 |
-0.421 |
2.065 |
-0.141 |
| H6 |
0.189 |
1.033 |
-1.452 |
| H7 |
1.951 |
1.738 |
0.225 |
| H8 |
1.084 |
0.816 |
1.467 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5080 | 1.9510 | 2.7478 | 1.0861 | 1.0840 | 2.1286 | 2.1577 |
C2 | 1.5080 | | 2.8919 | 1.7887 | 2.1359 | 2.1588 | 1.0878 | 1.0849 | Br3 | 1.9510 | 2.8919 | | 3.5476 | 2.4790 | 2.4866 | 3.8502 | 3.0150 | Cl4 | 2.7478 | 1.7887 | 3.5476 | | 3.7003 | 2.8952 | 2.3444 | 2.3522 | H5 | 1.0861 | 2.1359 | 2.4790 | 3.7003 | | 1.7766 | 2.4218 | 2.5316 | H6 | 1.0840 | 2.1588 | 2.4866 | 2.8952 | 1.7766 | | 2.5323 | 3.0613 | H7 | 2.1286 | 1.0878 | 3.8502 | 2.3444 | 2.4218 | 2.5323 | | 1.7731 | H8 | 2.1577 | 1.0849 | 3.0150 | 2.3522 | 2.5316 | 3.0613 | 1.7731 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.644 |
|
C1 |
C2 |
H7 |
109.101 |
| C1 |
C2 |
H8 |
111.595 |
|
C2 |
C1 |
Br3 |
112.826 |
| C2 |
C1 |
Cl4 |
36.925 |
|
C2 |
C1 |
H6 |
111.742 |
| Br3 |
C1 |
H5 |
105.879 |
|
Br3 |
C1 |
H6 |
106.504 |
| Cl4 |
C2 |
H7 |
106.628 |
|
Cl4 |
C2 |
H8 |
107.332 |
| H5 |
C1 |
H6 |
109.902 |
|
H7 |
C2 |
H8 |
109.380 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.146 |
|
|
|
| 2 |
C |
-0.087 |
|
|
|
| 3 |
Br |
-0.116 |
|
|
|
| 4 |
Cl |
-0.141 |
|
|
|
| 5 |
H |
0.126 |
|
|
|
| 6 |
H |
0.128 |
|
|
|
| 7 |
H |
0.117 |
|
|
|
| 8 |
H |
0.119 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.405 |
2.610 |
0.095 |
2.643 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-47.887 |
2.598 |
0.418 |
| y |
2.598 |
-40.851 |
-0.079 |
| z |
0.418 |
-0.079 |
-43.437 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.743 |
2.598 |
0.418 |
| y |
2.598 |
4.811 |
-0.079 |
| z |
0.418 |
-0.079 |
0.932 |
|
| Polar |
|---|
| 3z2-r2 | 1.865 |
|---|
| x2-y2 | -7.036 |
|---|
| xy | 2.598 |
|---|
| xz | 0.418 |
|---|
| yz | -0.079 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.550 |
0.273 |
-0.220 |
| y |
0.273 |
8.319 |
0.033 |
| z |
-0.220 |
0.033 |
6.582 |
<r2> (average value of r
2) Å
2
| <r2> |
222.027 |
| (<r2>)1/2 |
14.901 |
