Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -997.122264 |
| Energy at 298.15K | -997.127112 |
| HF Energy | -997.122264 |
| Nuclear repulsion energy | 195.114235 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3252 |
2948 |
0.00 |
|
|
|
| 2 |
Ag |
1619 |
1468 |
0.00 |
|
|
|
| 3 |
Ag |
1466 |
1329 |
0.00 |
|
|
|
| 4 |
Ag |
1129 |
1023 |
0.00 |
|
|
|
| 5 |
Ag |
826 |
749 |
0.00 |
|
|
|
| 6 |
Ag |
322 |
292 |
0.00 |
|
|
|
| 7 |
Au |
3330 |
3018 |
5.81 |
|
|
|
| 8 |
Au |
1243 |
1126 |
1.01 |
|
|
|
| 9 |
Au |
833 |
755 |
1.70 |
|
|
|
| 10 |
Au |
126 |
114 |
8.21 |
|
|
|
| 11 |
Bg |
3306 |
2996 |
0.00 |
|
|
|
| 12 |
Bg |
1405 |
1274 |
0.00 |
|
|
|
| 13 |
Bg |
1099 |
996 |
0.00 |
|
|
|
| 14 |
Bu |
3259 |
2954 |
22.49 |
|
|
|
| 15 |
Bu |
1618 |
1467 |
8.35 |
|
|
|
| 16 |
Bu |
1371 |
1242 |
49.85 |
|
|
|
| 17 |
Bu |
770 |
698 |
133.99 |
|
|
|
| 18 |
Bu |
235 |
213 |
9.97 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13603.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 12330.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.471 |
0.592 |
0.000 |
| C2 |
-0.471 |
-0.592 |
0.000 |
| Cl3 |
-0.471 |
2.107 |
0.000 |
| Cl4 |
0.471 |
-2.107 |
0.000 |
| H5 |
1.093 |
0.599 |
0.878 |
| H6 |
1.093 |
0.599 |
-0.878 |
| H7 |
-1.093 |
-0.599 |
0.878 |
| H8 |
-1.093 |
-0.599 |
-0.878 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5130 | 1.7848 | 2.6991 | 1.0756 | 1.0756 | 2.1530 | 2.1530 |
C2 | 1.5130 | | 2.6991 | 1.7848 | 2.1530 | 2.1530 | 1.0756 | 1.0756 | Cl3 | 1.7848 | 2.6991 | | 4.3188 | 2.3432 | 2.3432 | 2.9126 | 2.9126 | Cl4 | 2.6991 | 1.7848 | 4.3188 | | 2.9126 | 2.9126 | 2.3432 | 2.3432 | H5 | 1.0756 | 2.1530 | 2.3432 | 2.9126 | | 1.7559 | 2.4924 | 3.0488 | H6 | 1.0756 | 2.1530 | 2.3432 | 2.9126 | 1.7559 | | 3.0488 | 2.4924 | H7 | 2.1530 | 1.0756 | 2.9126 | 2.3432 | 2.4924 | 3.0488 | | 1.7559 | H8 | 2.1530 | 1.0756 | 2.9126 | 2.3432 | 3.0488 | 2.4924 | 1.7559 | |
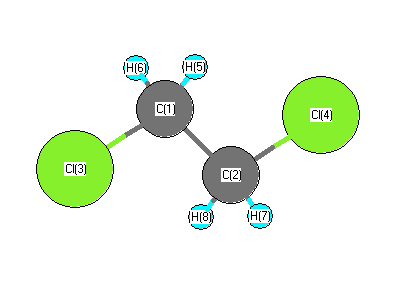 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.587 |
|
C1 |
C2 |
H7 |
111.432 |
| C1 |
C2 |
H8 |
111.432 |
|
C2 |
C1 |
Cl3 |
109.587 |
| C2 |
C1 |
H5 |
111.432 |
|
C2 |
C1 |
H6 |
111.432 |
| Cl3 |
C1 |
H5 |
107.400 |
|
Cl3 |
C1 |
H6 |
107.400 |
| Cl4 |
C2 |
H7 |
107.400 |
|
Cl4 |
C2 |
H8 |
107.400 |
| H5 |
C1 |
H6 |
109.425 |
|
H7 |
C2 |
H8 |
109.425 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.006 |
|
|
|
| 2 |
C |
0.006 |
|
|
|
| 3 |
Cl |
-0.181 |
|
|
|
| 4 |
Cl |
-0.181 |
|
|
|
| 5 |
H |
0.087 |
|
|
|
| 6 |
H |
0.087 |
|
|
|
| 7 |
H |
0.087 |
|
|
|
| 8 |
H |
0.087 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.451 |
2.293 |
0.000 |
| y |
2.293 |
-45.992 |
0.000 |
| z |
0.000 |
0.000 |
-37.676 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.383 |
2.293 |
0.000 |
| y |
2.293 |
-8.428 |
0.000 |
| z |
0.000 |
0.000 |
4.045 |
|
| Polar |
|---|
| 3z2-r2 | 8.091 |
|---|
| x2-y2 | 8.540 |
|---|
| xy | 2.293 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.201 |
-1.340 |
0.000 |
| y |
-1.340 |
9.287 |
0.000 |
| z |
0.000 |
0.000 |
5.349 |
<r2> (average value of r
2) Å
2
| <r2> |
200.120 |
| (<r2>)1/2 |
14.146 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -997.119329 |
| Energy at 298.15K | -997.124318 |
| HF Energy | -997.119329 |
| Nuclear repulsion energy | 202.365802 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3293 |
2985 |
1.28 |
|
|
|
| 2 |
A |
3241 |
2937 |
32.91 |
|
|
|
| 3 |
A |
1604 |
1454 |
0.21 |
|
|
|
| 4 |
A |
1467 |
1330 |
24.20 |
|
|
|
| 5 |
A |
1337 |
1212 |
0.33 |
|
|
|
| 6 |
A |
1125 |
1020 |
0.77 |
|
|
|
| 7 |
A |
1030 |
934 |
14.60 |
|
|
|
| 8 |
A |
708 |
642 |
28.36 |
|
|
|
| 9 |
A |
279 |
253 |
0.99 |
|
|
|
| 10 |
A |
122 |
110 |
1.19 |
|
|
|
| 11 |
B |
3306 |
2997 |
8.04 |
|
|
|
| 12 |
B |
3229 |
2927 |
4.09 |
|
|
|
| 13 |
B |
1600 |
1450 |
10.35 |
|
|
|
| 14 |
B |
1441 |
1306 |
51.79 |
|
|
|
| 15 |
B |
1262 |
1144 |
0.51 |
|
|
|
| 16 |
B |
976 |
884 |
17.83 |
|
|
|
| 17 |
B |
737 |
668 |
39.32 |
|
|
|
| 18 |
B |
439 |
398 |
9.75 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13597.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 12324.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.298 |
0.693 |
0.881 |
| C2 |
-0.298 |
-0.693 |
0.881 |
| Cl3 |
-0.298 |
1.699 |
-0.464 |
| Cl4 |
0.298 |
-1.699 |
-0.464 |
| H5 |
0.018 |
1.199 |
1.792 |
| H6 |
1.371 |
0.660 |
0.803 |
| H7 |
-0.018 |
-1.199 |
1.792 |
| H8 |
-1.371 |
-0.660 |
0.803 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5096 | 1.7822 | 2.7447 | 1.0788 | 1.0765 | 2.1235 | 2.1505 |
C2 | 1.5096 | | 2.7447 | 1.7822 | 2.1235 | 2.1505 | 1.0788 | 1.0765 | Cl3 | 1.7822 | 2.7447 | | 3.4506 | 2.3324 | 2.3398 | 3.6831 | 2.8848 | Cl4 | 2.7447 | 1.7822 | 3.4506 | | 3.6831 | 2.8848 | 2.3324 | 2.3398 | H5 | 1.0788 | 2.1235 | 2.3324 | 3.6831 | | 1.7606 | 2.3973 | 2.5222 | H6 | 1.0765 | 2.1505 | 2.3398 | 2.8848 | 1.7606 | | 2.5222 | 3.0437 | H7 | 2.1235 | 1.0788 | 3.6831 | 2.3324 | 2.3973 | 2.5222 | | 1.7606 | H8 | 2.1505 | 1.0765 | 2.8848 | 2.3398 | 2.5222 | 3.0437 | 1.7606 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.717 |
|
C1 |
C2 |
H7 |
109.115 |
| C1 |
C2 |
H8 |
111.425 |
|
C2 |
C1 |
Cl3 |
112.717 |
| C2 |
C1 |
H5 |
109.115 |
|
C2 |
C1 |
H6 |
111.425 |
| Cl3 |
C1 |
H5 |
106.617 |
|
Cl3 |
C1 |
H6 |
107.273 |
| Cl4 |
C2 |
H7 |
106.617 |
|
Cl4 |
C2 |
H8 |
107.273 |
| H5 |
C1 |
H6 |
109.546 |
|
H7 |
C2 |
H8 |
109.546 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.005 |
|
|
|
| 2 |
C |
0.005 |
|
|
|
| 3 |
Cl |
-0.169 |
|
|
|
| 4 |
Cl |
-0.169 |
|
|
|
| 5 |
H |
0.081 |
|
|
|
| 6 |
H |
0.084 |
|
|
|
| 7 |
H |
0.081 |
|
|
|
| 8 |
H |
0.084 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
2.957 |
2.957 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.591 |
1.354 |
0.000 |
| y |
1.354 |
-42.977 |
0.000 |
| z |
0.000 |
0.000 |
-35.505 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.651 |
1.354 |
0.000 |
| y |
1.354 |
-6.429 |
0.000 |
| z |
0.000 |
0.000 |
4.779 |
|
| Polar |
|---|
| 3z2-r2 | 9.557 |
|---|
| x2-y2 | 5.387 |
|---|
| xy | 1.354 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.684 |
-0.540 |
0.000 |
| y |
-0.540 |
7.626 |
0.000 |
| z |
0.000 |
0.000 |
7.091 |
<r2> (average value of r
2) Å
2
| <r2> |
164.376 |
| (<r2>)1/2 |
12.821 |
