Jump to
S1C2
Vibrational Frequencies calculated at QCISD/Def2TZVPP
Geometric Data calculated at QCISD/Def2TZVPP
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -207.615747 |
| Energy at 298.15K | -207.619107 |
| HF Energy | -206.856251 |
| Nuclear repulsion energy | 102.228486 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3886 |
3886 |
49.21 |
|
|
|
| 2 |
A |
3151 |
3151 |
3.29 |
|
|
|
| 3 |
A |
3076 |
3076 |
19.92 |
|
|
|
| 4 |
A |
2344 |
2344 |
1.16 |
|
|
|
| 5 |
A |
1517 |
1517 |
2.44 |
|
|
|
| 6 |
A |
1444 |
1444 |
46.13 |
|
|
|
| 7 |
A |
1401 |
1401 |
2.45 |
|
|
|
| 8 |
A |
1249 |
1249 |
16.01 |
|
|
|
| 9 |
A |
1116 |
1116 |
97.21 |
|
|
|
| 10 |
A |
1007 |
1007 |
21.46 |
|
|
|
| 11 |
A |
908 |
908 |
17.61 |
|
|
|
| 12 |
A |
585 |
585 |
1.53 |
|
|
|
| 13 |
A |
379 |
379 |
38.91 |
|
|
|
| 14 |
A |
302 |
302 |
101.53 |
|
|
|
| 15 |
A |
217 |
217 |
12.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11290.9 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 11290.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.574 |
0.590 |
0.038 |
| C2 |
0.825 |
0.110 |
-0.006 |
| O3 |
-1.508 |
-0.452 |
-0.110 |
| H4 |
-0.712 |
1.145 |
0.968 |
| H5 |
-0.730 |
1.275 |
-0.791 |
| H6 |
-1.397 |
-1.068 |
0.615 |
| N7 |
1.914 |
-0.277 |
-0.015 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
N7 |
| C1 | | 1.4797 | 1.4072 | 1.0918 | 1.0870 | 1.9391 | 2.6347 |
C2 | 1.4797 | | 2.4022 | 2.0935 | 2.0960 | 2.5905 | 1.1552 | O3 | 1.4072 | 2.4022 | | 2.0846 | 2.0127 | 0.9583 | 3.4277 | H4 | 1.0918 | 2.0935 | 2.0846 | | 1.7643 | 2.3437 | 3.1436 | H5 | 1.0870 | 2.0960 | 2.0127 | 1.7643 | | 2.8131 | 3.1624 | H6 | 1.9391 | 2.5905 | 0.9583 | 2.3437 | 2.8131 | | 3.4615 | N7 | 2.6347 | 1.1552 | 3.4277 | 3.1436 | 3.1624 | 3.4615 | |
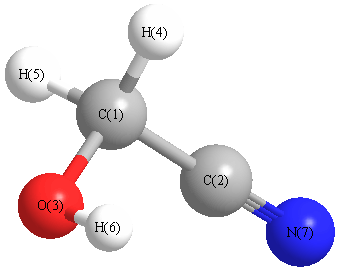 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N7 |
178.603 |
|
C1 |
O3 |
H6 |
108.639 |
| C2 |
C1 |
O3 |
112.611 |
|
C2 |
C1 |
H4 |
108.056 |
| C2 |
C1 |
H5 |
108.527 |
|
O3 |
C1 |
H4 |
112.454 |
| O3 |
C1 |
H5 |
106.897 |
|
H4 |
C1 |
H5 |
108.142 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
