Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -611.906765 |
| Energy at 298.15K | -611.910199 |
| HF Energy | -611.906765 |
| Nuclear repulsion energy | 144.238407 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3314 |
3004 |
1.42 |
|
|
|
| 2 |
A |
3232 |
2930 |
11.96 |
|
|
|
| 3 |
A |
3139 |
2845 |
50.88 |
|
|
|
| 4 |
A |
2005 |
1817 |
241.02 |
|
|
|
| 5 |
A |
1582 |
1434 |
9.29 |
|
|
|
| 6 |
A |
1529 |
1386 |
11.74 |
|
|
|
| 7 |
A |
1391 |
1261 |
27.44 |
|
|
|
| 8 |
A |
1305 |
1183 |
6.30 |
|
|
|
| 9 |
A |
1143 |
1036 |
13.33 |
|
|
|
| 10 |
A |
1120 |
1015 |
20.42 |
|
|
|
| 11 |
A |
882 |
800 |
28.11 |
|
|
|
| 12 |
A |
763 |
692 |
25.94 |
|
|
|
| 13 |
A |
497 |
451 |
15.09 |
|
|
|
| 14 |
A |
312 |
283 |
13.15 |
|
|
|
| 15 |
A |
65 |
59 |
20.90 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11140.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 10098.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.026 |
0.661 |
0.127 |
| C2 |
1.162 |
-0.336 |
0.166 |
| Cl3 |
-1.541 |
-0.162 |
-0.070 |
| O4 |
2.247 |
-0.056 |
-0.193 |
| H5 |
-0.005 |
1.201 |
1.061 |
| H6 |
0.168 |
1.347 |
-0.689 |
| H7 |
0.933 |
-1.307 |
0.609 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5113 | 1.7814 | 2.3556 | 1.0792 | 1.0755 | 2.2197 |
C2 | 1.5113 | | 2.7187 | 1.1776 | 2.1274 | 2.1331 | 1.0913 | Cl3 | 1.7814 | 2.7187 | | 3.7921 | 2.3446 | 2.3622 | 2.8103 | O4 | 2.3556 | 1.1776 | 3.7921 | | 2.8683 | 2.5565 | 1.9839 | H5 | 1.0792 | 2.1274 | 2.3446 | 2.8683 | | 1.7643 | 2.7159 | H6 | 1.0755 | 2.1331 | 2.3622 | 2.5565 | 1.7643 | | 3.0517 | H7 | 2.2197 | 1.0913 | 2.8103 | 1.9839 | 2.7159 | 3.0517 | |
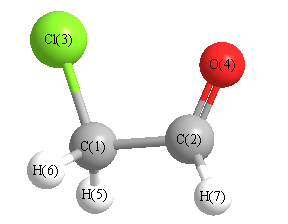 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
121.846 |
|
C1 |
C2 |
H7 |
116.121 |
| C2 |
C1 |
Cl3 |
111.050 |
|
C2 |
C1 |
H5 |
109.280 |
| C2 |
C1 |
H6 |
109.951 |
|
Cl3 |
C1 |
H5 |
107.546 |
| Cl3 |
C1 |
H6 |
109.040 |
|
O4 |
C2 |
H7 |
121.903 |
| H5 |
C1 |
H6 |
109.937 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.041 |
|
|
|
| 2 |
C |
0.263 |
|
|
|
| 3 |
Cl |
-0.162 |
|
|
|
| 4 |
O |
-0.371 |
|
|
|
| 5 |
H |
0.082 |
|
|
|
| 6 |
H |
0.107 |
|
|
|
| 7 |
H |
0.041 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.983 |
0.620 |
1.196 |
1.667 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.049 |
-0.357 |
1.728 |
| y |
-0.357 |
-28.275 |
-0.073 |
| z |
1.728 |
-0.073 |
-29.307 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-11.258 |
-0.357 |
1.728 |
| y |
-0.357 |
6.403 |
-0.073 |
| z |
1.728 |
-0.073 |
4.855 |
|
| Polar |
|---|
| 3z2-r2 | 9.710 |
|---|
| x2-y2 | -11.775 |
|---|
| xy | -0.357 |
|---|
| xz | 1.728 |
|---|
| yz | -0.073 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.998 |
0.523 |
-0.176 |
| y |
0.523 |
4.821 |
-0.160 |
| z |
-0.176 |
-0.160 |
4.206 |
<r2> (average value of r
2) Å
2
| <r2> |
120.793 |
| (<r2>)1/2 |
10.991 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -611.904124 |
| Energy at 298.15K | -611.907714 |
| HF Energy | -611.904124 |
| Nuclear repulsion energy | 147.963886 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3214 |
2913 |
26.09 |
|
|
|
| 2 |
A' |
3082 |
2794 |
87.48 |
|
|
|
| 3 |
A' |
2022 |
1833 |
205.38 |
|
|
|
| 4 |
A' |
1572 |
1425 |
21.37 |
|
|
|
| 5 |
A' |
1528 |
1385 |
26.47 |
|
|
|
| 6 |
A' |
1440 |
1305 |
15.69 |
|
|
|
| 7 |
A' |
1021 |
926 |
5.72 |
|
|
|
| 8 |
A' |
829 |
751 |
7.65 |
|
|
|
| 9 |
A' |
678 |
615 |
66.25 |
|
|
|
| 10 |
A' |
230 |
208 |
3.75 |
|
|
|
| 11 |
A" |
3261 |
2956 |
2.65 |
|
|
|
| 12 |
A" |
1328 |
1204 |
0.92 |
|
|
|
| 13 |
A" |
1144 |
1036 |
0.95 |
|
|
|
| 14 |
A" |
764 |
693 |
0.71 |
|
|
|
| 15 |
A" |
169 |
153 |
2.93 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 11140.4 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 10097.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.909 |
0.000 |
| C2 |
1.362 |
0.262 |
0.000 |
| Cl3 |
-1.345 |
-0.234 |
0.000 |
| O4 |
1.584 |
-0.891 |
0.000 |
| H5 |
-0.083 |
1.539 |
0.874 |
| H6 |
-0.083 |
1.539 |
-0.874 |
| H7 |
2.176 |
0.997 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
O4 |
H5 |
H6 |
H7 |
| C1 | | 1.5080 | 1.7644 | 2.3977 | 1.0808 | 1.0808 | 2.1777 |
C2 | 1.5080 | | 2.7520 | 1.1744 | 2.1172 | 2.1172 | 1.0959 | Cl3 | 1.7644 | 2.7520 | | 3.0016 | 2.3446 | 2.3446 | 3.7293 | O4 | 2.3977 | 1.1744 | 3.0016 | | 3.0735 | 3.0735 | 1.9781 | H5 | 1.0808 | 2.1172 | 2.3446 | 3.0735 | | 1.7490 | 2.4822 | H6 | 1.0808 | 2.1172 | 2.3446 | 3.0735 | 1.7490 | | 2.4822 | H7 | 2.1777 | 1.0959 | 3.7293 | 1.9781 | 2.4822 | 2.4822 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O4 |
126.274 |
|
C1 |
C2 |
H7 |
112.545 |
| C2 |
C1 |
Cl3 |
114.259 |
|
C2 |
C1 |
H5 |
108.611 |
| C2 |
C1 |
H6 |
108.611 |
|
Cl3 |
C1 |
H5 |
108.588 |
| Cl3 |
C1 |
H6 |
108.588 |
|
O4 |
C2 |
H7 |
121.181 |
| H5 |
C1 |
H6 |
108.016 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.038 |
|
|
|
| 2 |
C |
0.252 |
|
|
|
| 3 |
Cl |
-0.126 |
|
|
|
| 4 |
O |
-0.350 |
|
|
|
| 5 |
H |
0.080 |
|
|
|
| 6 |
H |
0.080 |
|
|
|
| 7 |
H |
0.025 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.600 |
3.779 |
0.000 |
3.826 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.672 |
4.278 |
0.000 |
| y |
4.278 |
-29.886 |
0.000 |
| z |
0.000 |
0.000 |
-29.367 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.046 |
4.278 |
0.000 |
| y |
4.278 |
1.134 |
0.000 |
| z |
0.000 |
0.000 |
1.912 |
|
| Polar |
|---|
| 3z2-r2 | 3.824 |
|---|
| x2-y2 | -2.787 |
|---|
| xy | 4.278 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.165 |
0.548 |
0.000 |
| y |
0.548 |
5.845 |
0.000 |
| z |
0.000 |
0.000 |
3.960 |
<r2> (average value of r
2) Å
2
| <r2> |
105.794 |
| (<r2>)1/2 |
10.286 |
