Jump to
S1C2
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -270.140386 |
| Energy at 298.15K | -270.151156 |
| HF Energy | -270.140386 |
| Nuclear repulsion energy | 237.239878 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3276 |
2970 |
17.54 |
|
|
|
| 2 |
A' |
3218 |
2917 |
59.89 |
|
|
|
| 3 |
A' |
3183 |
2885 |
15.10 |
|
|
|
| 4 |
A' |
3170 |
2873 |
12.74 |
|
|
|
| 5 |
A' |
3153 |
2857 |
38.09 |
|
|
|
| 6 |
A' |
3145 |
2850 |
13.37 |
|
|
|
| 7 |
A' |
1976 |
1791 |
224.65 |
|
|
|
| 8 |
A' |
1627 |
1474 |
6.09 |
|
|
|
| 9 |
A' |
1608 |
1458 |
1.51 |
|
|
|
| 10 |
A' |
1586 |
1437 |
20.74 |
|
|
|
| 11 |
A' |
1577 |
1429 |
0.71 |
|
|
|
| 12 |
A' |
1546 |
1401 |
30.50 |
|
|
|
| 13 |
A' |
1537 |
1393 |
12.52 |
|
|
|
| 14 |
A' |
1519 |
1377 |
12.86 |
|
|
|
| 15 |
A' |
1443 |
1308 |
7.94 |
|
|
|
| 16 |
A' |
1282 |
1162 |
68.44 |
|
|
|
| 17 |
A' |
1215 |
1102 |
0.24 |
|
|
|
| 18 |
A' |
1110 |
1006 |
0.33 |
|
|
|
| 19 |
A' |
1042 |
944 |
6.99 |
|
|
|
| 20 |
A' |
962 |
872 |
6.52 |
|
|
|
| 21 |
A' |
886 |
803 |
0.04 |
|
|
|
| 22 |
A' |
640 |
580 |
19.88 |
|
|
|
| 23 |
A' |
422 |
382 |
1.44 |
|
|
|
| 24 |
A' |
362 |
328 |
3.25 |
|
|
|
| 25 |
A' |
186 |
169 |
5.34 |
|
|
|
| 26 |
A" |
3227 |
2925 |
98.19 |
|
|
|
| 27 |
A" |
3222 |
2921 |
2.38 |
|
|
|
| 28 |
A" |
3193 |
2894 |
9.46 |
|
|
|
| 29 |
A" |
3167 |
2871 |
9.23 |
|
|
|
| 30 |
A" |
1616 |
1465 |
5.61 |
|
|
|
| 31 |
A" |
1598 |
1448 |
9.88 |
|
|
|
| 32 |
A" |
1435 |
1301 |
0.07 |
|
|
|
| 33 |
A" |
1363 |
1235 |
0.10 |
|
|
|
| 34 |
A" |
1250 |
1133 |
0.83 |
|
|
|
| 35 |
A" |
1064 |
964 |
0.94 |
|
|
|
| 36 |
A" |
904 |
820 |
0.06 |
|
|
|
| 37 |
A" |
783 |
710 |
2.99 |
|
|
|
| 38 |
A" |
520 |
471 |
1.29 |
|
|
|
| 39 |
A" |
263 |
238 |
0.01 |
|
|
|
| 40 |
A" |
112 |
101 |
0.11 |
|
|
|
| 41 |
A" |
104 |
95 |
1.50 |
|
|
|
| 42 |
A" |
20 |
19 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32755.3 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29689.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.412 |
1.348 |
0.000 |
| C2 |
-1.467 |
0.153 |
0.000 |
| C3 |
0.000 |
0.563 |
0.000 |
| C4 |
0.988 |
-0.581 |
0.000 |
| C5 |
2.448 |
-0.210 |
0.000 |
| O6 |
0.635 |
-1.714 |
0.000 |
| H7 |
-3.449 |
1.032 |
0.000 |
| H8 |
-2.263 |
1.972 |
0.875 |
| H9 |
-2.263 |
1.972 |
-0.875 |
| H10 |
-1.660 |
-0.473 |
0.862 |
| H11 |
-1.660 |
-0.473 |
-0.862 |
| H12 |
0.223 |
1.183 |
-0.864 |
| H13 |
0.223 |
1.183 |
0.864 |
| H14 |
3.066 |
-1.095 |
0.000 |
| H15 |
2.682 |
0.389 |
-0.873 |
| H16 |
2.682 |
0.389 |
0.873 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.5237 | 2.5369 | 3.9088 | 5.1037 | 4.3200 | 1.0834 | 1.0852 | 1.0852 | 2.1502 | 2.1502 | 2.7785 | 2.7785 | 5.9977 | 5.2565 | 5.2565 |
C2 | 1.5237 | | 1.5227 | 2.5616 | 3.9312 | 2.8113 | 2.1687 | 2.1703 | 2.1703 | 1.0829 | 1.0829 | 2.1592 | 2.1592 | 4.7007 | 4.2457 | 4.2457 | C3 | 2.5369 | 1.5227 | | 1.5108 | 2.5668 | 2.3639 | 3.4808 | 2.8063 | 2.8063 | 2.1379 | 2.1379 | 1.0866 | 1.0866 | 3.4848 | 2.8256 | 2.8256 | C4 | 3.9088 | 2.5616 | 1.5108 | | 1.5065 | 1.1875 | 4.7208 | 4.2252 | 4.2252 | 2.7867 | 2.7867 | 2.1070 | 2.1070 | 2.1404 | 2.1381 | 2.1381 | C5 | 5.1037 | 3.9312 | 2.5668 | 1.5065 | | 2.3560 | 6.0262 | 5.2652 | 5.2652 | 4.2055 | 4.2055 | 2.7630 | 2.7630 | 1.0792 | 1.0842 | 1.0842 | O6 | 4.3200 | 2.8113 | 2.3639 | 1.1875 | 2.3560 | | 4.9217 | 4.7705 | 4.7705 | 2.7481 | 2.7481 | 3.0510 | 3.0510 | 2.5083 | 3.0618 | 3.0618 | H7 | 1.0834 | 2.1687 | 3.4808 | 4.7208 | 6.0262 | 4.9217 | | 1.7477 | 1.7477 | 2.4920 | 2.4920 | 3.7754 | 3.7754 | 6.8529 | 6.2258 | 6.2258 | H8 | 1.0852 | 2.1703 | 2.8063 | 4.2252 | 5.2652 | 4.7705 | 1.7477 | | 1.7493 | 2.5185 | 3.0594 | 3.1352 | 2.6090 | 6.2103 | 5.4787 | 5.1923 | H9 | 1.0852 | 2.1703 | 2.8063 | 4.2252 | 5.2652 | 4.7705 | 1.7477 | 1.7493 | | 3.0594 | 2.5185 | 2.6090 | 3.1352 | 6.2103 | 5.1923 | 5.4787 | H10 | 2.1502 | 1.0829 | 2.1379 | 2.7867 | 4.2055 | 2.7481 | 2.4920 | 2.5185 | 3.0594 | | 1.7249 | 3.0441 | 2.5071 | 4.8435 | 4.7543 | 4.4261 | H11 | 2.1502 | 1.0829 | 2.1379 | 2.7867 | 4.2055 | 2.7481 | 2.4920 | 3.0594 | 2.5185 | 1.7249 | | 2.5071 | 3.0441 | 4.8435 | 4.4261 | 4.7543 | H12 | 2.7785 | 2.1592 | 1.0866 | 2.1070 | 2.7630 | 3.0510 | 3.7754 | 3.1352 | 2.6090 | 3.0441 | 2.5071 | | 1.7280 | 3.7431 | 2.5834 | 3.1133 | H13 | 2.7785 | 2.1592 | 1.0866 | 2.1070 | 2.7630 | 3.0510 | 3.7754 | 2.6090 | 3.1352 | 2.5071 | 3.0441 | 1.7280 | | 3.7431 | 3.1133 | 2.5834 | H14 | 5.9977 | 4.7007 | 3.4848 | 2.1404 | 1.0792 | 2.5083 | 6.8529 | 6.2103 | 6.2103 | 4.8435 | 4.8435 | 3.7431 | 3.7431 | | 1.7637 | 1.7637 | H15 | 5.2565 | 4.2457 | 2.8256 | 2.1381 | 1.0842 | 3.0618 | 6.2258 | 5.4787 | 5.1923 | 4.7543 | 4.4261 | 2.5834 | 3.1133 | 1.7637 | | 1.7468 | H16 | 5.2565 | 4.2457 | 2.8256 | 2.1381 | 1.0842 | 3.0618 | 6.2258 | 5.1923 | 5.4787 | 4.4261 | 4.7543 | 3.1133 | 2.5834 | 1.7637 | 1.7468 | |
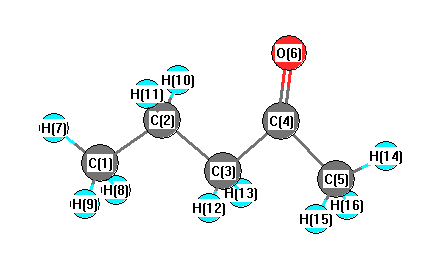 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.763 |
|
C1 |
C2 |
H10 |
110.010 |
| C1 |
C2 |
H11 |
110.010 |
|
C2 |
C1 |
H7 |
111.453 |
| C2 |
C1 |
H8 |
111.477 |
|
C2 |
C1 |
H9 |
111.477 |
| C2 |
C3 |
C4 |
115.226 |
|
C2 |
C3 |
H12 |
110.569 |
| C2 |
C3 |
H13 |
110.569 |
|
C3 |
C2 |
H10 |
109.106 |
| C3 |
C2 |
H11 |
109.106 |
|
C3 |
C4 |
C5 |
116.580 |
| C3 |
C4 |
O6 |
121.885 |
|
C4 |
C3 |
H12 |
107.302 |
| C4 |
C3 |
H13 |
107.302 |
|
C4 |
C5 |
H14 |
110.660 |
| C4 |
C5 |
H15 |
110.169 |
|
C4 |
C5 |
H16 |
110.169 |
| C5 |
C4 |
O6 |
121.535 |
|
H7 |
C1 |
H8 |
107.395 |
| H7 |
C1 |
H9 |
107.395 |
|
H8 |
C1 |
H9 |
107.414 |
| H10 |
C2 |
H11 |
105.589 |
|
H12 |
C3 |
H13 |
105.337 |
| H14 |
C5 |
H15 |
109.222 |
|
H14 |
C5 |
H16 |
109.222 |
| H15 |
C5 |
H16 |
107.328 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.179 |
|
|
|
| 2 |
C |
0.015 |
|
|
|
| 3 |
C |
-0.119 |
|
|
|
| 4 |
C |
0.344 |
|
|
|
| 5 |
C |
-0.212 |
|
|
|
| 6 |
O |
-0.423 |
|
|
|
| 7 |
H |
0.065 |
|
|
|
| 8 |
H |
0.045 |
|
|
|
| 9 |
H |
0.045 |
|
|
|
| 10 |
H |
0.039 |
|
|
|
| 11 |
H |
0.039 |
|
|
|
| 12 |
H |
0.053 |
|
|
|
| 13 |
H |
0.053 |
|
|
|
| 14 |
H |
0.105 |
|
|
|
| 15 |
H |
0.065 |
|
|
|
| 16 |
H |
0.065 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.760 |
2.871 |
0.000 |
2.970 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.528 |
1.589 |
0.000 |
| y |
1.589 |
-45.065 |
0.000 |
| z |
0.000 |
0.000 |
-37.214 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.612 |
1.589 |
0.000 |
| y |
1.589 |
-8.694 |
0.000 |
| z |
0.000 |
0.000 |
3.083 |
|
| Polar |
|---|
| 3z2-r2 | 6.165 |
|---|
| x2-y2 | 9.538 |
|---|
| xy | 1.589 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.961 |
-0.346 |
0.000 |
| y |
-0.346 |
9.027 |
0.000 |
| z |
0.000 |
0.000 |
7.346 |
<r2> (average value of r
2) Å
2
| <r2> |
227.136 |
| (<r2>)1/2 |
15.071 |
Jump to
S1C1
Energy calculated at HF/Def2TZVPP
| | hartrees |
|---|
| Energy at 0K | -270.140385 |
| Energy at 298.15K | -270.151155 |
| HF Energy | -270.140385 |
| Nuclear repulsion energy | 237.234811 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3276 |
2969 |
17.56 |
|
|
|
| 2 |
A' |
3218 |
2917 |
59.96 |
|
|
|
| 3 |
A' |
3183 |
2885 |
15.02 |
|
|
|
| 4 |
A' |
3170 |
2873 |
12.82 |
|
|
|
| 5 |
A' |
3153 |
2857 |
38.02 |
|
|
|
| 6 |
A' |
3145 |
2850 |
13.36 |
|
|
|
| 7 |
A' |
1976 |
1791 |
224.72 |
|
|
|
| 8 |
A' |
1627 |
1474 |
6.09 |
|
|
|
| 9 |
A' |
1608 |
1458 |
1.51 |
|
|
|
| 10 |
A' |
1586 |
1437 |
20.74 |
|
|
|
| 11 |
A' |
1577 |
1429 |
0.74 |
|
|
|
| 12 |
A' |
1546 |
1401 |
30.72 |
|
|
|
| 13 |
A' |
1537 |
1393 |
12.33 |
|
|
|
| 14 |
A' |
1519 |
1377 |
12.81 |
|
|
|
| 15 |
A' |
1443 |
1308 |
7.91 |
|
|
|
| 16 |
A' |
1282 |
1162 |
68.48 |
|
|
|
| 17 |
A' |
1215 |
1102 |
0.24 |
|
|
|
| 18 |
A' |
1110 |
1006 |
0.32 |
|
|
|
| 19 |
A' |
1042 |
944 |
6.99 |
|
|
|
| 20 |
A' |
962 |
872 |
6.54 |
|
|
|
| 21 |
A' |
886 |
803 |
0.04 |
|
|
|
| 22 |
A' |
640 |
580 |
19.86 |
|
|
|
| 23 |
A' |
422 |
382 |
1.44 |
|
|
|
| 24 |
A' |
362 |
328 |
3.25 |
|
|
|
| 25 |
A' |
186 |
169 |
5.34 |
|
|
|
| 26 |
A" |
3227 |
2925 |
97.52 |
|
|
|
| 27 |
A" |
3222 |
2920 |
2.90 |
|
|
|
| 28 |
A" |
3193 |
2894 |
9.60 |
|
|
|
| 29 |
A" |
3167 |
2871 |
9.23 |
|
|
|
| 30 |
A" |
1616 |
1465 |
5.61 |
|
|
|
| 31 |
A" |
1598 |
1448 |
9.88 |
|
|
|
| 32 |
A" |
1435 |
1301 |
0.07 |
|
|
|
| 33 |
A" |
1363 |
1235 |
0.10 |
|
|
|
| 34 |
A" |
1250 |
1133 |
0.83 |
|
|
|
| 35 |
A" |
1064 |
964 |
0.94 |
|
|
|
| 36 |
A" |
904 |
819 |
0.06 |
|
|
|
| 37 |
A" |
783 |
710 |
2.99 |
|
|
|
| 38 |
A" |
520 |
471 |
1.29 |
|
|
|
| 39 |
A" |
263 |
238 |
0.01 |
|
|
|
| 40 |
A" |
112 |
101 |
0.11 |
|
|
|
| 41 |
A" |
104 |
95 |
1.50 |
|
|
|
| 42 |
A" |
20 |
18 |
0.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32754.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 29689.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at HF/Def2TZVPP
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.408 |
1.352 |
0.000 |
| C2 |
-1.465 |
0.155 |
0.000 |
| C3 |
0.000 |
0.569 |
0.000 |
| C4 |
0.985 |
-0.576 |
0.000 |
| C5 |
2.449 |
-0.226 |
0.000 |
| O6 |
0.630 |
-1.711 |
0.000 |
| H7 |
-3.442 |
1.033 |
0.000 |
| H8 |
-2.256 |
1.975 |
0.876 |
| H9 |
-2.256 |
1.975 |
-0.876 |
| H10 |
-1.658 |
-0.468 |
0.864 |
| H11 |
-1.658 |
-0.468 |
-0.864 |
| H12 |
0.225 |
1.190 |
-0.864 |
| H13 |
0.225 |
1.190 |
0.864 |
| H14 |
3.045 |
-1.125 |
0.000 |
| H15 |
2.686 |
0.370 |
-0.873 |
| H16 |
2.686 |
0.370 |
0.873 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
O6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.5240 | 2.5319 | 3.9026 | 5.1064 | 4.3140 | 1.0828 | 1.0849 | 1.0849 | 2.1504 | 2.1504 | 2.7755 | 2.7755 | 5.9889 | 5.2601 | 5.2601 |
C2 | 1.5240 | | 1.5227 | 2.5569 | 3.9323 | 2.8054 | 2.1633 | 2.1691 | 2.1691 | 1.0825 | 1.0825 | 2.1620 | 2.1620 | 4.6881 | 4.2472 | 4.2472 | C3 | 2.5319 | 1.5227 | | 1.5107 | 2.5745 | 2.3652 | 3.4734 | 2.7988 | 2.7988 | 2.1379 | 2.1379 | 1.0873 | 1.0873 | 3.4843 | 2.8310 | 2.8310 | C4 | 3.9026 | 2.5569 | 1.5107 | | 1.5050 | 1.1883 | 4.7106 | 4.2167 | 4.2167 | 2.7823 | 2.7823 | 2.1079 | 2.1079 | 2.1317 | 2.1334 | 2.1334 | C5 | 5.1064 | 3.9323 | 2.5745 | 1.5050 | | 2.3474 | 6.0239 | 5.2674 | 5.2674 | 4.2030 | 4.2030 | 2.7742 | 2.7742 | 1.0785 | 1.0837 | 1.0837 | O6 | 4.3140 | 2.8054 | 2.3652 | 1.1883 | 2.3474 | | 4.9103 | 4.7623 | 4.7623 | 2.7429 | 2.7429 | 3.0532 | 3.0532 | 2.4848 | 3.0525 | 3.0525 | H7 | 1.0828 | 2.1633 | 3.4734 | 4.7106 | 6.0239 | 4.9103 | | 1.7495 | 1.7495 | 2.4871 | 2.4871 | 3.7708 | 3.7708 | 6.8366 | 6.2252 | 6.2252 | H8 | 1.0849 | 2.1691 | 2.7988 | 4.2167 | 5.2674 | 4.7623 | 1.7495 | | 1.7511 | 2.5155 | 3.0584 | 3.1303 | 2.6024 | 6.2029 | 5.4822 | 5.1958 | H9 | 1.0849 | 2.1691 | 2.7988 | 4.2167 | 5.2674 | 4.7623 | 1.7495 | 1.7511 | | 3.0584 | 2.5155 | 2.6024 | 3.1303 | 6.2029 | 5.1958 | 5.4822 | H10 | 2.1504 | 1.0825 | 2.1379 | 2.7823 | 4.2030 | 2.7429 | 2.4871 | 2.5155 | 3.0584 | | 1.7281 | 3.0462 | 2.5085 | 4.8260 | 4.7524 | 4.4235 | H11 | 2.1504 | 1.0825 | 2.1379 | 2.7823 | 4.2030 | 2.7429 | 2.4871 | 3.0584 | 2.5155 | 1.7281 | | 2.5085 | 3.0462 | 4.8260 | 4.4235 | 4.7524 | H12 | 2.7755 | 2.1620 | 1.0873 | 2.1079 | 2.7742 | 3.0532 | 3.7708 | 3.1303 | 2.6024 | 3.0462 | 2.5085 | | 1.7282 | 3.7491 | 2.5936 | 3.1217 | H13 | 2.7755 | 2.1620 | 1.0873 | 2.1079 | 2.7742 | 3.0532 | 3.7708 | 2.6024 | 3.1303 | 2.5085 | 3.0462 | 1.7282 | | 3.7491 | 3.1217 | 2.5936 | H14 | 5.9889 | 4.6881 | 3.4843 | 2.1317 | 1.0785 | 2.4848 | 6.8366 | 6.2029 | 6.2029 | 4.8260 | 4.8260 | 3.7491 | 3.7491 | | 1.7683 | 1.7683 | H15 | 5.2601 | 4.2472 | 2.8310 | 2.1334 | 1.0837 | 3.0525 | 6.2252 | 5.4822 | 5.1958 | 4.7524 | 4.4235 | 2.5936 | 3.1217 | 1.7683 | | 1.7463 | H16 | 5.2601 | 4.2472 | 2.8310 | 2.1334 | 1.0837 | 3.0525 | 6.2252 | 5.1958 | 5.4822 | 4.4235 | 4.7524 | 3.1217 | 2.5936 | 1.7683 | 1.7463 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
112.407 |
|
C1 |
C2 |
H10 |
110.028 |
| C1 |
C2 |
H11 |
110.028 |
|
C2 |
C1 |
H7 |
111.038 |
| C2 |
C1 |
H8 |
111.380 |
|
C2 |
C1 |
H9 |
111.380 |
| C2 |
C3 |
C4 |
114.897 |
|
C2 |
C3 |
H12 |
110.751 |
| C2 |
C3 |
H13 |
110.751 |
|
C3 |
C2 |
H10 |
109.123 |
| C3 |
C2 |
H11 |
109.123 |
|
C3 |
C4 |
C5 |
117.229 |
| C3 |
C4 |
O6 |
121.941 |
|
C4 |
C3 |
H12 |
107.332 |
| C4 |
C3 |
H13 |
107.332 |
|
C4 |
C5 |
H14 |
110.107 |
| C4 |
C5 |
H15 |
109.931 |
|
C4 |
C5 |
H16 |
109.931 |
| C5 |
C4 |
O6 |
120.829 |
|
H7 |
C1 |
H8 |
107.621 |
| H7 |
C1 |
H9 |
107.621 |
|
H8 |
C1 |
H9 |
107.615 |
| H10 |
C2 |
H11 |
105.918 |
|
H12 |
C3 |
H13 |
105.253 |
| H14 |
C5 |
H15 |
109.735 |
|
H14 |
C5 |
H16 |
109.735 |
| H15 |
C5 |
H16 |
107.359 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.179 |
|
|
|
| 2 |
C |
0.015 |
|
|
|
| 3 |
C |
-0.119 |
|
|
|
| 4 |
C |
0.344 |
|
|
|
| 5 |
C |
-0.212 |
|
|
|
| 6 |
O |
-0.423 |
|
|
|
| 7 |
H |
0.065 |
|
|
|
| 8 |
H |
0.045 |
|
|
|
| 9 |
H |
0.045 |
|
|
|
| 10 |
H |
0.039 |
|
|
|
| 11 |
H |
0.039 |
|
|
|
| 12 |
H |
0.053 |
|
|
|
| 13 |
H |
0.053 |
|
|
|
| 14 |
H |
0.105 |
|
|
|
| 15 |
H |
0.065 |
|
|
|
| 16 |
H |
0.065 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.758 |
2.872 |
0.000 |
2.970 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.529 |
1.588 |
0.000 |
| y |
1.588 |
-45.067 |
0.000 |
| z |
0.000 |
0.000 |
-37.214 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.612 |
1.588 |
0.000 |
| y |
1.588 |
-8.696 |
0.000 |
| z |
0.000 |
0.000 |
3.084 |
|
| Polar |
|---|
| 3z2-r2 | 6.168 |
|---|
| x2-y2 | 9.538 |
|---|
| xy | 1.588 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.961 |
-0.346 |
0.000 |
| y |
-0.346 |
9.027 |
0.000 |
| z |
0.000 |
0.000 |
7.346 |
<r2> (average value of r
2) Å
2
| <r2> |
227.154 |
| (<r2>)1/2 |
15.072 |
