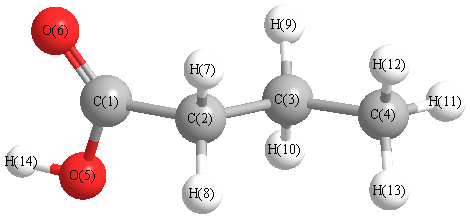Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4111 |
3726 |
118.48 |
|
|
|
| 2 |
A |
3257 |
2952 |
36.48 |
|
|
|
| 3 |
A |
3223 |
2921 |
49.12 |
|
|
|
| 4 |
A |
3220 |
2919 |
54.61 |
|
|
|
| 5 |
A |
3197 |
2898 |
3.75 |
|
|
|
| 6 |
A |
3194 |
2895 |
15.01 |
|
|
|
| 7 |
A |
3169 |
2872 |
13.35 |
|
|
|
| 8 |
A |
3157 |
2862 |
34.72 |
|
|
|
| 9 |
A |
1991 |
1805 |
455.88 |
|
|
|
| 10 |
A |
1630 |
1477 |
7.46 |
|
|
|
| 11 |
A |
1615 |
1464 |
7.92 |
|
|
|
| 12 |
A |
1612 |
1461 |
1.00 |
|
|
|
| 13 |
A |
1606 |
1456 |
8.36 |
|
|
|
| 14 |
A |
1541 |
1397 |
6.33 |
|
|
|
| 15 |
A |
1522 |
1380 |
30.39 |
|
|
|
| 16 |
A |
1490 |
1350 |
67.91 |
|
|
|
| 17 |
A |
1434 |
1300 |
2.00 |
|
|
|
| 18 |
A |
1411 |
1279 |
3.76 |
|
|
|
| 19 |
A |
1369 |
1241 |
38.01 |
|
|
|
| 20 |
A |
1323 |
1199 |
151.27 |
|
|
|
| 21 |
A |
1207 |
1094 |
9.61 |
|
|
|
| 22 |
A |
1179 |
1069 |
86.78 |
|
|
|
| 23 |
A |
1105 |
1001 |
1.15 |
|
|
|
| 24 |
A |
980 |
888 |
0.14 |
|
|
|
| 25 |
A |
968 |
877 |
2.73 |
|
|
|
| 26 |
A |
948 |
859 |
6.42 |
|
|
|
| 27 |
A |
813 |
736 |
24.81 |
|
|
|
| 28 |
A |
798 |
723 |
21.68 |
|
|
|
| 29 |
A |
665 |
602 |
71.90 |
|
|
|
| 30 |
A |
615 |
558 |
82.80 |
|
|
|
| 31 |
A |
465 |
421 |
3.85 |
|
|
|
| 32 |
A |
352 |
319 |
1.47 |
|
|
|
| 33 |
A |
262 |
237 |
0.02 |
|
|
|
| 34 |
A |
201 |
182 |
0.15 |
|
|
|
| 35 |
A |
96 |
87 |
0.50 |
|
|
|
| 36 |
A |
40 |
36 |
0.91 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 27881.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 25271.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.482 |
|
|
|
| 2 |
C |
-0.058 |
|
|
|
| 3 |
C |
-0.069 |
|
|
|
| 4 |
C |
-0.188 |
|
|
|
| 5 |
O |
-0.379 |
|
|
|
| 6 |
O |
-0.450 |
|
|
|
| 7 |
H |
0.084 |
|
|
|
| 8 |
H |
0.063 |
|
|
|
| 9 |
H |
0.053 |
|
|
|
| 10 |
H |
0.057 |
|
|
|
| 11 |
H |
0.065 |
|
|
|
| 12 |
H |
0.053 |
|
|
|
| 13 |
H |
0.051 |
|
|
|
| 14 |
H |
0.233 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.360 |
1.541 |
0.348 |
2.085 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.296 |
5.618 |
0.214 |
| y |
5.618 |
-39.988 |
-1.605 |
| z |
0.214 |
-1.605 |
-36.194 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.795 |
5.618 |
0.214 |
| y |
5.618 |
-3.742 |
-1.605 |
| z |
0.214 |
-1.605 |
1.948 |
|
| Polar |
|---|
| 3z2-r2 | 3.896 |
|---|
| x2-y2 | 3.691 |
|---|
| xy | 5.618 |
|---|
| xz | 0.214 |
|---|
| yz | -1.605 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.807 |
-0.181 |
-0.243 |
| y |
-0.181 |
7.328 |
0.068 |
| z |
-0.243 |
0.068 |
6.524 |
<r2> (average value of r
2) Å
2
| <r2> |
206.598 |
| (<r2>)1/2 |
14.374 |