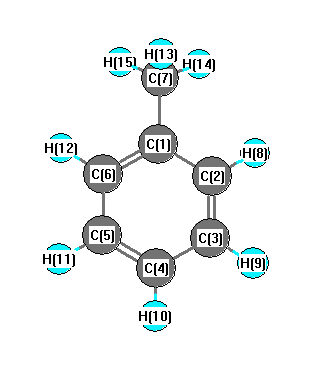
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3346 |
3033 |
16.40 |
|
|
|
| 2 |
A' |
3324 |
3013 |
11.71 |
|
|
|
| 3 |
A' |
3308 |
2998 |
8.16 |
|
|
|
| 4 |
A' |
3213 |
2913 |
26.87 |
|
|
|
| 5 |
A' |
3162 |
2866 |
35.90 |
|
|
|
| 6 |
A' |
1789 |
1621 |
10.96 |
|
|
|
| 7 |
A' |
1655 |
1500 |
16.71 |
|
|
|
| 8 |
A' |
1608 |
1457 |
5.53 |
|
|
|
| 9 |
A' |
1539 |
1395 |
0.06 |
|
|
|
| 10 |
A' |
1313 |
1190 |
0.79 |
|
|
|
| 11 |
A' |
1290 |
1169 |
0.48 |
|
|
|
| 12 |
A' |
1163 |
1054 |
5.93 |
|
|
|
| 13 |
A' |
1121 |
1016 |
2.13 |
|
|
|
| 14 |
A' |
1111 |
1007 |
0.04 |
|
|
|
| 15 |
A' |
1084 |
983 |
0.09 |
|
|
|
| 16 |
A' |
1012 |
918 |
0.95 |
|
|
|
| 17 |
A' |
846 |
767 |
1.75 |
|
|
|
| 18 |
A' |
818 |
742 |
73.32 |
|
|
|
| 19 |
A' |
769 |
697 |
16.76 |
|
|
|
| 20 |
A' |
561 |
509 |
0.82 |
|
|
|
| 21 |
A' |
520 |
471 |
10.02 |
|
|
|
| 22 |
A' |
228 |
207 |
2.55 |
|
|
|
| 23 |
A" |
3333 |
3021 |
46.24 |
|
|
|
| 24 |
A" |
3311 |
3001 |
4.13 |
|
|
|
| 25 |
A" |
3237 |
2934 |
23.45 |
|
|
|
| 26 |
A" |
1762 |
1597 |
1.14 |
|
|
|
| 27 |
A" |
1620 |
1468 |
14.58 |
|
|
|
| 28 |
A" |
1588 |
1439 |
0.03 |
|
|
|
| 29 |
A" |
1470 |
1332 |
0.03 |
|
|
|
| 30 |
A" |
1327 |
1203 |
0.00 |
|
|
|
| 31 |
A" |
1216 |
1102 |
0.30 |
|
|
|
| 32 |
A" |
1165 |
1056 |
3.83 |
|
|
|
| 33 |
A" |
1100 |
997 |
0.00 |
|
|
|
| 34 |
A" |
1071 |
970 |
0.01 |
|
|
|
| 35 |
A" |
951 |
862 |
0.02 |
|
|
|
| 36 |
A" |
680 |
616 |
0.07 |
|
|
|
| 37 |
A" |
455 |
412 |
0.00 |
|
|
|
| 38 |
A" |
367 |
333 |
0.37 |
|
|
|
| 39 |
A" |
23 |
21 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 29726.0 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 26943.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.079 |
|
|
|
| 2 |
C |
-0.149 |
|
|
|
| 3 |
C |
-0.066 |
|
|
|
| 4 |
C |
-0.127 |
|
|
|
| 5 |
C |
-0.066 |
|
|
|
| 6 |
C |
-0.149 |
|
|
|
| 7 |
C |
-0.172 |
|
|
|
| 8 |
H |
0.082 |
|
|
|
| 9 |
H |
0.093 |
|
|
|
| 10 |
H |
0.090 |
|
|
|
| 11 |
H |
0.093 |
|
|
|
| 12 |
H |
0.082 |
|
|
|
| 13 |
H |
0.074 |
|
|
|
| 14 |
H |
0.069 |
|
|
|
| 15 |
H |
0.069 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.032 |
0.368 |
0.000 |
0.370 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.760 |
-0.058 |
0.000 |
| y |
-0.058 |
-38.537 |
0.000 |
| z |
0.000 |
0.000 |
-38.775 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-8.104 |
-0.058 |
0.000 |
| y |
-0.058 |
4.230 |
0.000 |
| z |
0.000 |
0.000 |
3.874 |
|
| Polar |
|---|
| 3z2-r2 | 7.748 |
|---|
| x2-y2 | -8.223 |
|---|
| xy | -0.058 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.193 |
-0.082 |
0.000 |
| y |
-0.082 |
13.991 |
0.000 |
| z |
0.000 |
0.000 |
12.276 |
<r2> (average value of r
2) Å
2
| <r2> |
198.905 |
| (<r2>)1/2 |
14.103 |