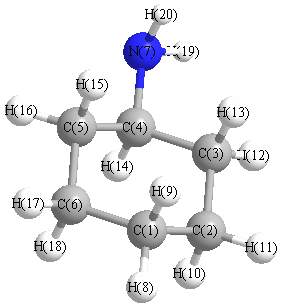
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3794 |
3439 |
0.76 |
|
|
|
| 2 |
A |
3716 |
3368 |
0.35 |
|
|
|
| 3 |
A |
3206 |
2906 |
66.43 |
|
|
|
| 4 |
A |
3191 |
2892 |
126.84 |
|
|
|
| 5 |
A |
3186 |
2888 |
84.67 |
|
|
|
| 6 |
A |
3183 |
2885 |
71.57 |
|
|
|
| 7 |
A |
3173 |
2876 |
81.32 |
|
|
|
| 8 |
A |
3145 |
2851 |
92.56 |
|
|
|
| 9 |
A |
3140 |
2846 |
9.76 |
|
|
|
| 10 |
A |
3138 |
2844 |
12.58 |
|
|
|
| 11 |
A |
3136 |
2842 |
6.55 |
|
|
|
| 12 |
A |
3126 |
2834 |
16.13 |
|
|
|
| 13 |
A |
3123 |
2830 |
10.64 |
|
|
|
| 14 |
A |
1790 |
1622 |
34.49 |
|
|
|
| 15 |
A |
1632 |
1480 |
1.52 |
|
|
|
| 16 |
A |
1616 |
1465 |
7.88 |
|
|
|
| 17 |
A |
1614 |
1463 |
4.42 |
|
|
|
| 18 |
A |
1610 |
1460 |
1.16 |
|
|
|
| 19 |
A |
1604 |
1454 |
0.15 |
|
|
|
| 20 |
A |
1553 |
1408 |
6.98 |
|
|
|
| 21 |
A |
1512 |
1370 |
4.13 |
|
|
|
| 22 |
A |
1510 |
1368 |
0.82 |
|
|
|
| 23 |
A |
1503 |
1362 |
1.32 |
|
|
|
| 24 |
A |
1491 |
1352 |
0.42 |
|
|
|
| 25 |
A |
1482 |
1343 |
1.69 |
|
|
|
| 26 |
A |
1452 |
1316 |
1.00 |
|
|
|
| 27 |
A |
1405 |
1274 |
0.83 |
|
|
|
| 28 |
A |
1389 |
1259 |
2.37 |
|
|
|
| 29 |
A |
1387 |
1257 |
0.88 |
|
|
|
| 30 |
A |
1326 |
1202 |
4.25 |
|
|
|
| 31 |
A |
1297 |
1176 |
1.81 |
|
|
|
| 32 |
A |
1218 |
1104 |
18.72 |
|
|
|
| 33 |
A |
1202 |
1089 |
5.75 |
|
|
|
| 34 |
A |
1163 |
1054 |
0.04 |
|
|
|
| 35 |
A |
1151 |
1043 |
1.74 |
|
|
|
| 36 |
A |
1120 |
1016 |
8.56 |
|
|
|
| 37 |
A |
1104 |
1001 |
1.04 |
|
|
|
| 38 |
A |
1059 |
960 |
2.78 |
|
|
|
| 39 |
A |
1002 |
908 |
6.41 |
|
|
|
| 40 |
A |
974 |
883 |
63.14 |
|
|
|
| 41 |
A |
949 |
860 |
1.87 |
|
|
|
| 42 |
A |
920 |
834 |
96.64 |
|
|
|
| 43 |
A |
901 |
816 |
10.42 |
|
|
|
| 44 |
A |
854 |
774 |
0.90 |
|
|
|
| 45 |
A |
834 |
756 |
4.06 |
|
|
|
| 46 |
A |
590 |
535 |
1.00 |
|
|
|
| 47 |
A |
490 |
445 |
0.98 |
|
|
|
| 48 |
A |
482 |
437 |
1.70 |
|
|
|
| 49 |
A |
436 |
395 |
0.23 |
|
|
|
| 50 |
A |
367 |
333 |
10.89 |
|
|
|
| 51 |
A |
354 |
321 |
0.76 |
|
|
|
| 52 |
A |
265 |
241 |
29.66 |
|
|
|
| 53 |
A |
238 |
215 |
10.20 |
|
|
|
| 54 |
A |
169 |
153 |
0.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 43634.9 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 39550.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.080 |
|
|
|
| 2 |
C |
-0.087 |
|
|
|
| 3 |
C |
-0.117 |
|
|
|
| 4 |
C |
0.218 |
|
|
|
| 5 |
C |
-0.125 |
|
|
|
| 6 |
C |
-0.089 |
|
|
|
| 7 |
N |
-0.408 |
|
|
|
| 8 |
H |
0.055 |
|
|
|
| 9 |
H |
0.031 |
|
|
|
| 10 |
H |
0.034 |
|
|
|
| 11 |
H |
0.051 |
|
|
|
| 12 |
H |
0.066 |
|
|
|
| 13 |
H |
0.024 |
|
|
|
| 14 |
H |
0.024 |
|
|
|
| 15 |
H |
0.025 |
|
|
|
| 16 |
H |
0.042 |
|
|
|
| 17 |
H |
0.051 |
|
|
|
| 18 |
H |
0.033 |
|
|
|
| 19 |
H |
0.127 |
|
|
|
| 20 |
H |
0.125 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.567 |
1.053 |
0.442 |
1.275 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.363 |
-3.822 |
-1.520 |
| y |
-3.822 |
-46.712 |
-0.727 |
| z |
-1.520 |
-0.727 |
-44.472 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.771 |
-3.822 |
-1.520 |
| y |
-3.822 |
-0.295 |
-0.727 |
| z |
-1.520 |
-0.727 |
3.066 |
|
| Polar |
|---|
| 3z2-r2 | 6.131 |
|---|
| x2-y2 | -1.651 |
|---|
| xy | -3.822 |
|---|
| xz | -1.520 |
|---|
| yz | -0.727 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.783 |
-0.099 |
-0.093 |
| y |
-0.099 |
10.867 |
-0.049 |
| z |
-0.093 |
-0.049 |
9.679 |
<r2> (average value of r
2) Å
2
| <r2> |
231.606 |
| (<r2>)1/2 |
15.219 |