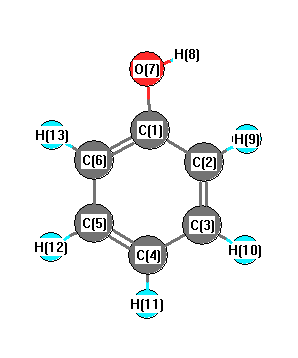
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
4182 |
3790 |
97.45 |
|
|
|
| 2 |
A' |
3354 |
3040 |
6.37 |
|
|
|
| 3 |
A' |
3347 |
3033 |
22.41 |
|
|
|
| 4 |
A' |
3331 |
3019 |
22.61 |
|
|
|
| 5 |
A' |
3321 |
3010 |
0.15 |
|
|
|
| 6 |
A' |
3305 |
2996 |
14.00 |
|
|
|
| 7 |
A' |
1787 |
1620 |
59.31 |
|
|
|
| 8 |
A' |
1777 |
1611 |
58.45 |
|
|
|
| 9 |
A' |
1657 |
1502 |
71.83 |
|
|
|
| 10 |
A' |
1624 |
1472 |
25.26 |
|
|
|
| 11 |
A' |
1487 |
1348 |
29.55 |
|
|
|
| 12 |
A' |
1387 |
1257 |
110.08 |
|
|
|
| 13 |
A' |
1361 |
1233 |
67.90 |
|
|
|
| 14 |
A' |
1285 |
1164 |
76.76 |
|
|
|
| 15 |
A' |
1277 |
1158 |
0.36 |
|
|
|
| 16 |
A' |
1176 |
1066 |
23.41 |
|
|
|
| 17 |
A' |
1166 |
1057 |
13.66 |
|
|
|
| 18 |
A' |
1114 |
1010 |
3.61 |
|
|
|
| 19 |
A' |
1081 |
979 |
4.64 |
|
|
|
| 20 |
A' |
887 |
804 |
19.89 |
|
|
|
| 21 |
A' |
677 |
614 |
0.36 |
|
|
|
| 22 |
A' |
573 |
520 |
2.24 |
|
|
|
| 23 |
A' |
441 |
399 |
10.20 |
|
|
|
| 24 |
A" |
1103 |
1000 |
0.31 |
|
|
|
| 25 |
A" |
1090 |
988 |
0.01 |
|
|
|
| 26 |
A" |
998 |
904 |
10.86 |
|
|
|
| 27 |
A" |
921 |
835 |
0.03 |
|
|
|
| 28 |
A" |
846 |
767 |
78.84 |
|
|
|
| 29 |
A" |
765 |
694 |
26.14 |
|
|
|
| 30 |
A" |
568 |
515 |
14.26 |
|
|
|
| 31 |
A" |
461 |
418 |
0.35 |
|
|
|
| 32 |
A" |
323 |
293 |
111.72 |
|
|
|
| 33 |
A" |
254 |
230 |
3.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24461.7 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 22172.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.286 |
|
|
|
| 2 |
C |
-0.146 |
|
|
|
| 3 |
C |
-0.081 |
|
|
|
| 4 |
C |
-0.131 |
|
|
|
| 5 |
C |
-0.054 |
|
|
|
| 6 |
C |
-0.122 |
|
|
|
| 7 |
O |
-0.392 |
|
|
|
| 8 |
H |
0.214 |
|
|
|
| 9 |
H |
0.059 |
|
|
|
| 10 |
H |
0.093 |
|
|
|
| 11 |
H |
0.090 |
|
|
|
| 12 |
H |
0.092 |
|
|
|
| 13 |
H |
0.092 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.335 |
-0.107 |
0.000 |
1.339 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-35.415 |
-4.364 |
0.000 |
| y |
-4.364 |
-37.854 |
0.000 |
| z |
0.000 |
0.000 |
-45.566 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.295 |
-4.364 |
0.000 |
| y |
-4.364 |
2.636 |
0.000 |
| z |
0.000 |
0.000 |
-8.931 |
|
| Polar |
|---|
| 3z2-r2 | -17.863 |
|---|
| x2-y2 | 2.439 |
|---|
| xy | -4.364 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.370 |
-0.174 |
0.000 |
| y |
-0.174 |
12.181 |
0.000 |
| z |
0.000 |
0.000 |
6.073 |
<r2> (average value of r
2) Å
2
| <r2> |
183.454 |
| (<r2>)1/2 |
13.545 |