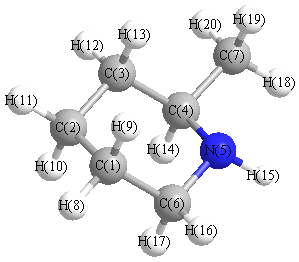
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3731 |
3382 |
0.26 |
|
|
|
| 2 |
A |
3233 |
2931 |
37.41 |
|
|
|
| 3 |
A |
3210 |
2909 |
92.12 |
|
|
|
| 4 |
A |
3201 |
2902 |
69.15 |
|
|
|
| 5 |
A |
3188 |
2889 |
96.06 |
|
|
|
| 6 |
A |
3184 |
2886 |
81.17 |
|
|
|
| 7 |
A |
3178 |
2880 |
64.86 |
|
|
|
| 8 |
A |
3153 |
2858 |
55.02 |
|
|
|
| 9 |
A |
3150 |
2855 |
38.32 |
|
|
|
| 10 |
A |
3139 |
2846 |
32.78 |
|
|
|
| 11 |
A |
3134 |
2841 |
12.12 |
|
|
|
| 12 |
A |
3133 |
2840 |
18.79 |
|
|
|
| 13 |
A |
3124 |
2832 |
8.54 |
|
|
|
| 14 |
A |
1637 |
1484 |
19.96 |
|
|
|
| 15 |
A |
1627 |
1475 |
1.34 |
|
|
|
| 16 |
A |
1617 |
1465 |
5.78 |
|
|
|
| 17 |
A |
1614 |
1463 |
3.56 |
|
|
|
| 18 |
A |
1609 |
1458 |
7.55 |
|
|
|
| 19 |
A |
1604 |
1454 |
0.79 |
|
|
|
| 20 |
A |
1600 |
1450 |
9.73 |
|
|
|
| 21 |
A |
1539 |
1395 |
7.30 |
|
|
|
| 22 |
A |
1521 |
1378 |
9.64 |
|
|
|
| 23 |
A |
1517 |
1375 |
1.50 |
|
|
|
| 24 |
A |
1504 |
1363 |
0.16 |
|
|
|
| 25 |
A |
1490 |
1350 |
1.42 |
|
|
|
| 26 |
A |
1482 |
1343 |
2.77 |
|
|
|
| 27 |
A |
1450 |
1314 |
0.37 |
|
|
|
| 28 |
A |
1411 |
1279 |
3.45 |
|
|
|
| 29 |
A |
1389 |
1259 |
5.99 |
|
|
|
| 30 |
A |
1350 |
1224 |
7.49 |
|
|
|
| 31 |
A |
1298 |
1176 |
4.20 |
|
|
|
| 32 |
A |
1274 |
1154 |
13.42 |
|
|
|
| 33 |
A |
1234 |
1119 |
28.69 |
|
|
|
| 34 |
A |
1192 |
1080 |
8.81 |
|
|
|
| 35 |
A |
1184 |
1073 |
2.61 |
|
|
|
| 36 |
A |
1132 |
1026 |
3.39 |
|
|
|
| 37 |
A |
1077 |
976 |
3.95 |
|
|
|
| 38 |
A |
1048 |
950 |
0.47 |
|
|
|
| 39 |
A |
1035 |
938 |
2.24 |
|
|
|
| 40 |
A |
985 |
893 |
14.07 |
|
|
|
| 41 |
A |
950 |
861 |
0.55 |
|
|
|
| 42 |
A |
913 |
828 |
19.39 |
|
|
|
| 43 |
A |
907 |
822 |
11.23 |
|
|
|
| 44 |
A |
863 |
783 |
22.46 |
|
|
|
| 45 |
A |
806 |
730 |
60.48 |
|
|
|
| 46 |
A |
604 |
547 |
1.35 |
|
|
|
| 47 |
A |
505 |
458 |
4.46 |
|
|
|
| 48 |
A |
491 |
445 |
1.15 |
|
|
|
| 49 |
A |
458 |
415 |
0.86 |
|
|
|
| 50 |
A |
353 |
320 |
1.00 |
|
|
|
| 51 |
A |
337 |
305 |
0.80 |
|
|
|
| 52 |
A |
265 |
240 |
1.91 |
|
|
|
| 53 |
A |
240 |
218 |
1.07 |
|
|
|
| 54 |
A |
161 |
146 |
0.80 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 43513.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 39440.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.082 |
|
|
|
| 2 |
C |
-0.059 |
|
|
|
| 3 |
C |
-0.128 |
|
|
|
| 4 |
C |
0.183 |
|
|
|
| 5 |
N |
-0.309 |
|
|
|
| 6 |
C |
0.009 |
|
|
|
| 7 |
C |
-0.215 |
|
|
|
| 8 |
H |
0.044 |
|
|
|
| 9 |
H |
0.024 |
|
|
|
| 10 |
H |
0.038 |
|
|
|
| 11 |
H |
0.049 |
|
|
|
| 12 |
H |
0.045 |
|
|
|
| 13 |
H |
0.028 |
|
|
|
| 14 |
H |
0.020 |
|
|
|
| 15 |
H |
0.097 |
|
|
|
| 16 |
H |
0.053 |
|
|
|
| 17 |
H |
0.035 |
|
|
|
| 18 |
H |
0.077 |
|
|
|
| 19 |
H |
0.039 |
|
|
|
| 20 |
H |
0.051 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.507 |
0.828 |
0.502 |
1.093 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.254 |
1.700 |
0.302 |
| y |
1.700 |
-48.583 |
-1.486 |
| z |
0.302 |
-1.486 |
-44.396 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.235 |
1.700 |
0.302 |
| y |
1.700 |
-3.258 |
-1.486 |
| z |
0.302 |
-1.486 |
3.023 |
|
| Polar |
|---|
| 3z2-r2 | 6.047 |
|---|
| x2-y2 | 2.329 |
|---|
| xy | 1.700 |
|---|
| xz | 0.302 |
|---|
| yz | -1.486 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.013 |
0.178 |
0.059 |
| y |
0.178 |
10.723 |
-0.071 |
| z |
0.059 |
-0.071 |
9.717 |
<r2> (average value of r
2) Å
2
| <r2> |
229.847 |
| (<r2>)1/2 |
15.161 |