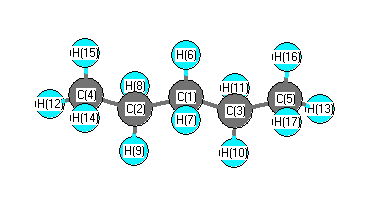
Vibrational Frequencies calculated at HF/Def2TZVPP
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3212 |
2911 |
82.80 |
|
|
|
| 2 |
A1 |
3154 |
2859 |
11.79 |
|
|
|
| 3 |
A1 |
3149 |
2854 |
117.96 |
|
|
|
| 4 |
A1 |
3133 |
2840 |
7.15 |
|
|
|
| 5 |
A1 |
1631 |
1478 |
6.75 |
|
|
|
| 6 |
A1 |
1613 |
1462 |
0.58 |
|
|
|
| 7 |
A1 |
1606 |
1456 |
0.20 |
|
|
|
| 8 |
A1 |
1539 |
1395 |
2.54 |
|
|
|
| 9 |
A1 |
1489 |
1350 |
0.04 |
|
|
|
| 10 |
A1 |
1254 |
1136 |
0.82 |
|
|
|
| 11 |
A1 |
1103 |
1000 |
0.31 |
|
|
|
| 12 |
A1 |
934 |
846 |
0.55 |
|
|
|
| 13 |
A1 |
421 |
382 |
0.00 |
|
|
|
| 14 |
A1 |
191 |
173 |
0.00 |
|
|
|
| 15 |
A2 |
3208 |
2908 |
0.00 |
|
|
|
| 16 |
A2 |
3161 |
2865 |
0.00 |
|
|
|
| 17 |
A2 |
1615 |
1463 |
0.00 |
|
|
|
| 18 |
A2 |
1439 |
1304 |
0.00 |
|
|
|
| 19 |
A2 |
1372 |
1243 |
0.00 |
|
|
|
| 20 |
A2 |
1065 |
966 |
0.00 |
|
|
|
| 21 |
A2 |
819 |
742 |
0.00 |
|
|
|
| 22 |
A2 |
268 |
243 |
0.00 |
|
|
|
| 23 |
A2 |
112 |
101 |
0.00 |
|
|
|
| 24 |
B1 |
3213 |
2912 |
211.76 |
|
|
|
| 25 |
B1 |
3181 |
2883 |
27.34 |
|
|
|
| 26 |
B1 |
3146 |
2852 |
0.11 |
|
|
|
| 27 |
B1 |
1614 |
1463 |
12.28 |
|
|
|
| 28 |
B1 |
1432 |
1298 |
0.75 |
|
|
|
| 29 |
B1 |
1311 |
1189 |
0.00 |
|
|
|
| 30 |
B1 |
929 |
842 |
0.57 |
|
|
|
| 31 |
B1 |
780 |
707 |
2.57 |
|
|
|
| 32 |
B1 |
257 |
233 |
0.00 |
|
|
|
| 33 |
B1 |
121 |
110 |
0.00 |
|
|
|
| 34 |
B2 |
3211 |
2911 |
46.92 |
|
|
|
| 35 |
B2 |
3152 |
2857 |
51.47 |
|
|
|
| 36 |
B2 |
3139 |
2846 |
3.67 |
|
|
|
| 37 |
B2 |
1622 |
1471 |
0.91 |
|
|
|
| 38 |
B2 |
1606 |
1456 |
0.79 |
|
|
|
| 39 |
B2 |
1540 |
1396 |
2.00 |
|
|
|
| 40 |
B2 |
1533 |
1389 |
0.20 |
|
|
|
| 41 |
B2 |
1402 |
1271 |
4.07 |
|
|
|
| 42 |
B2 |
1143 |
1036 |
0.69 |
|
|
|
| 43 |
B2 |
1112 |
1008 |
0.20 |
|
|
|
| 44 |
B2 |
981 |
889 |
1.60 |
|
|
|
| 45 |
B2 |
424 |
384 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37168.2 cm
-1
Scaled (by 0.9064) Zero Point Vibrational Energy (zpe) 33689.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/Def2TZVPP
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.054 |
|
|
|
| 2 |
C |
-0.022 |
|
|
|
| 3 |
C |
-0.022 |
|
|
|
| 4 |
C |
-0.183 |
|
|
|
| 5 |
C |
-0.183 |
|
|
|
| 6 |
H |
0.027 |
|
|
|
| 7 |
H |
0.027 |
|
|
|
| 8 |
H |
0.026 |
|
|
|
| 9 |
H |
0.026 |
|
|
|
| 10 |
H |
0.026 |
|
|
|
| 11 |
H |
0.026 |
|
|
|
| 12 |
H |
0.060 |
|
|
|
| 13 |
H |
0.060 |
|
|
|
| 14 |
H |
0.046 |
|
|
|
| 15 |
H |
0.046 |
|
|
|
| 16 |
H |
0.046 |
|
|
|
| 17 |
H |
0.046 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.088 |
0.088 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-34.598 |
0.000 |
0.000 |
| y |
0.000 |
-36.424 |
0.000 |
| z |
0.000 |
0.000 |
-35.723 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.475 |
0.000 |
0.000 |
| y |
0.000 |
-1.263 |
0.000 |
| z |
0.000 |
0.000 |
-0.212 |
|
| Polar |
|---|
| 3z2-r2 | -0.425 |
|---|
| x2-y2 | 1.825 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.962 |
0.000 |
0.000 |
| y |
0.000 |
10.333 |
0.000 |
| z |
0.000 |
0.000 |
8.350 |
<r2> (average value of r
2) Å
2
| <r2> |
200.553 |
| (<r2>)1/2 |
14.162 |